Upgrade to Pro — share decks privately, control downloads, hide ads and more …

"Mon make à moi", (tout sauf Galaxy)

Sponsored · Your Podcast. Everywhere. Effortlessly. Share. Educate. Inspire. Entertain. You do you. We'll handle the rest. →

"Mon make à moi", (tout sauf Galaxy)

Video available at: http://videos.rennes.inria.fr/ReNaBI-GO2013/indexPierreLindenbaum.html

11ème édition des rencontres de la plate-forme ReNaBI-Grand Ouest, la journée sera consacrée aux workflows en bio-informatique.*

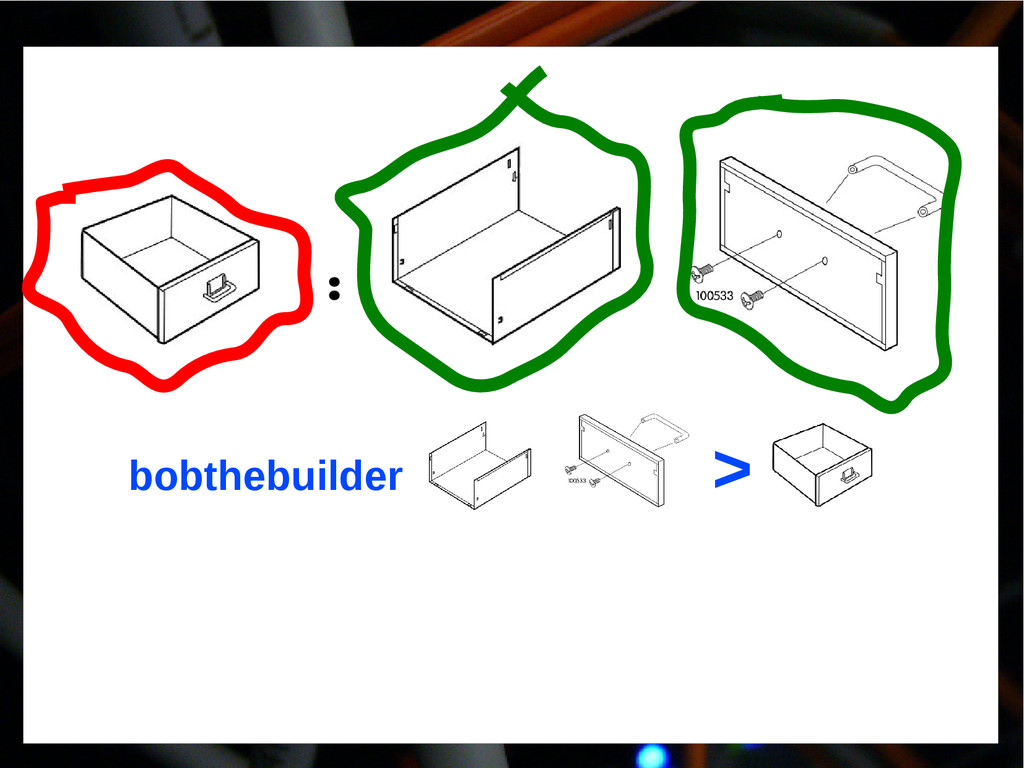
Avatar for Pierre Lindenbaum

Pierre Lindenbaum

November 29, 2013

Other Decks in Technology

Transcript

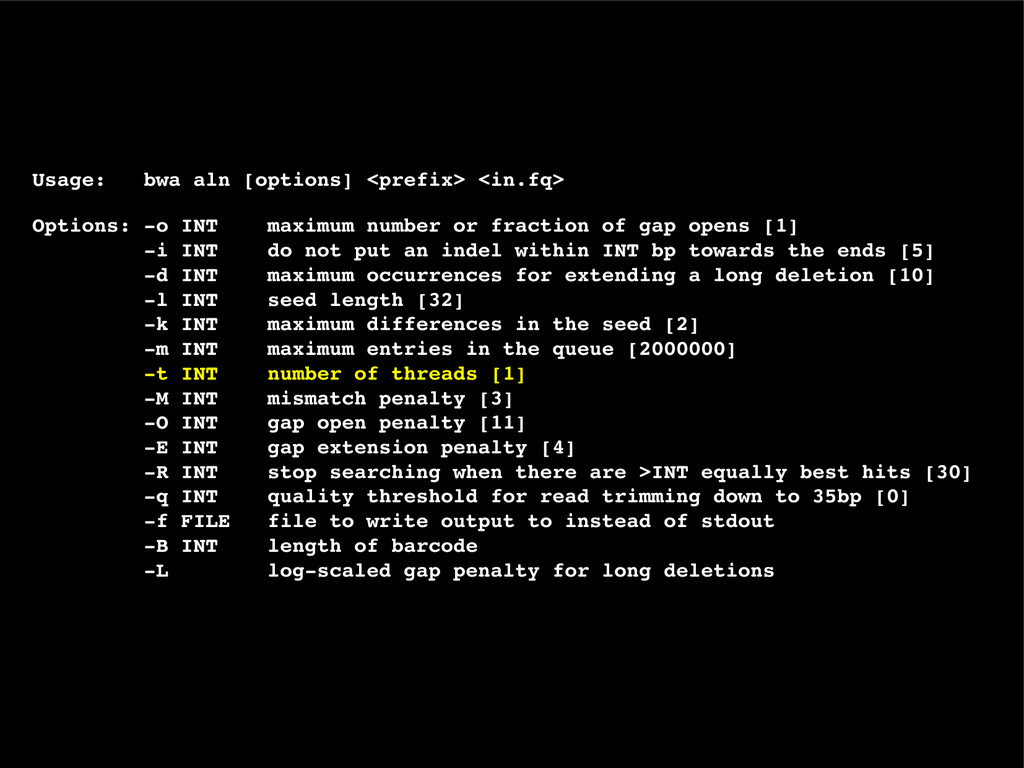
  1. Usage: bwa aln [options] <prefix> <in.fq> Options: ­o INT maximum

    number or fraction of gap opens [1] ­i INT do not put an indel within INT bp towards the ends [5] ­d INT maximum occurrences for extending a long deletion [10] ­l INT seed length [32] ­k INT maximum differences in the seed [2] ­m INT maximum entries in the queue [2000000] ­t INT number of threads [1] ­M INT mismatch penalty [3] ­O INT gap open penalty [11] ­E INT gap extension penalty [4] ­R INT stop searching when there are >INT equally best hits [30] ­q INT quality threshold for read trimming down to 35bp [0] ­f FILE file to write output to instead of stdout ­B INT length of barcode ­L log­scaled gap penalty for long deletions
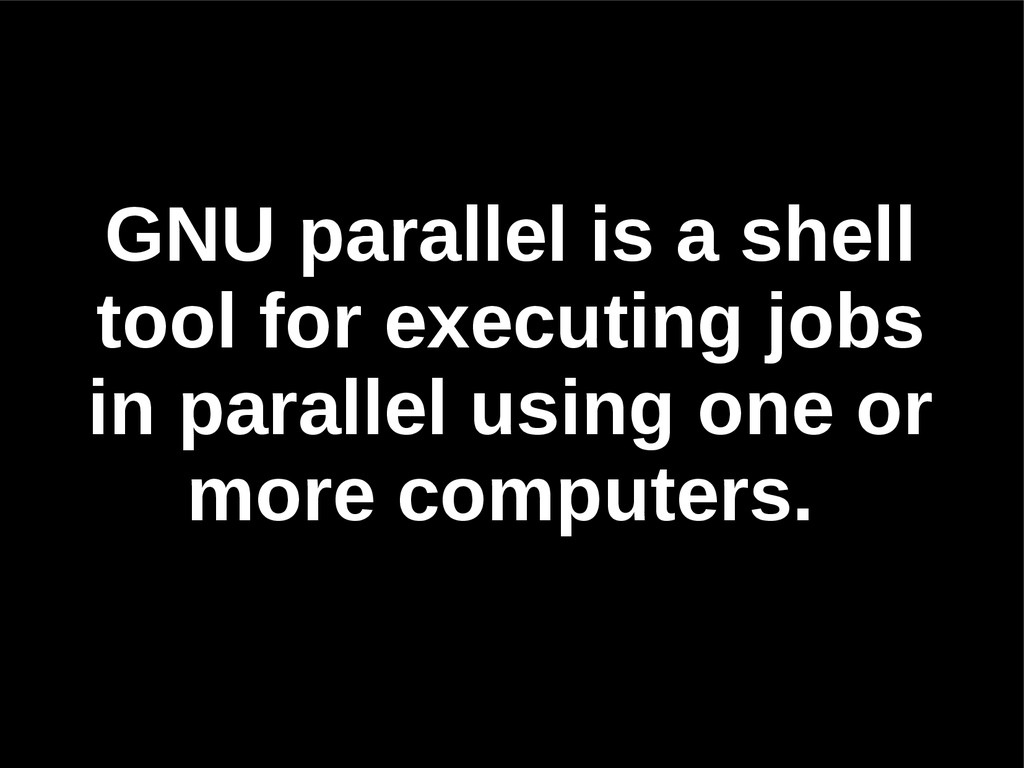
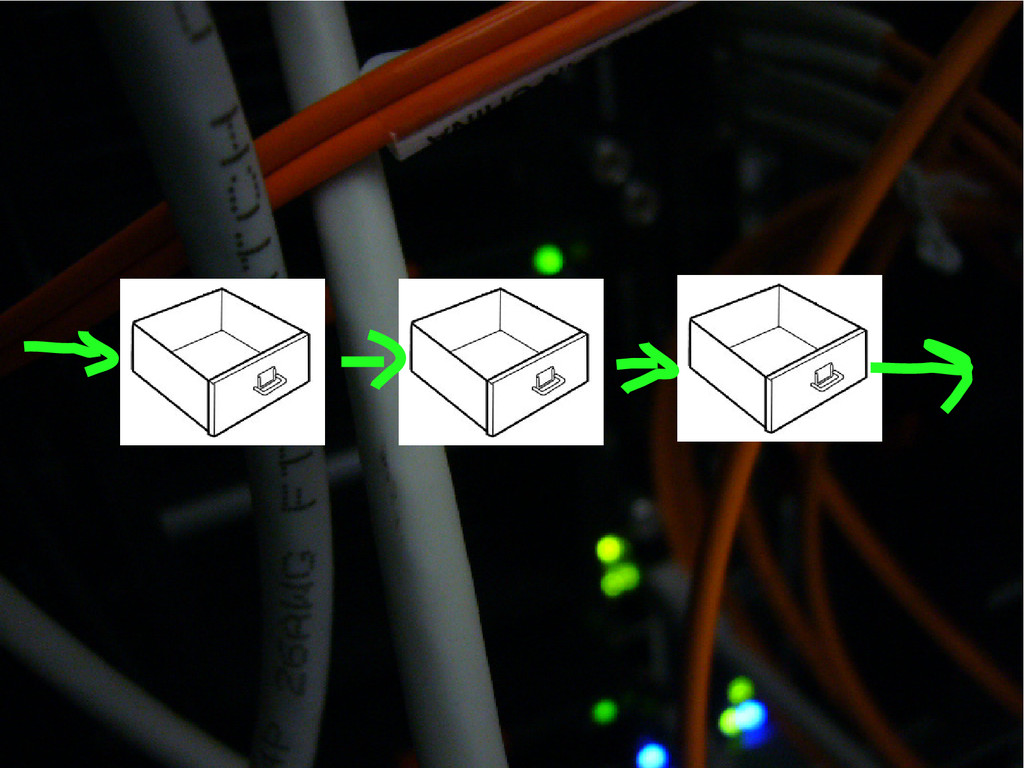
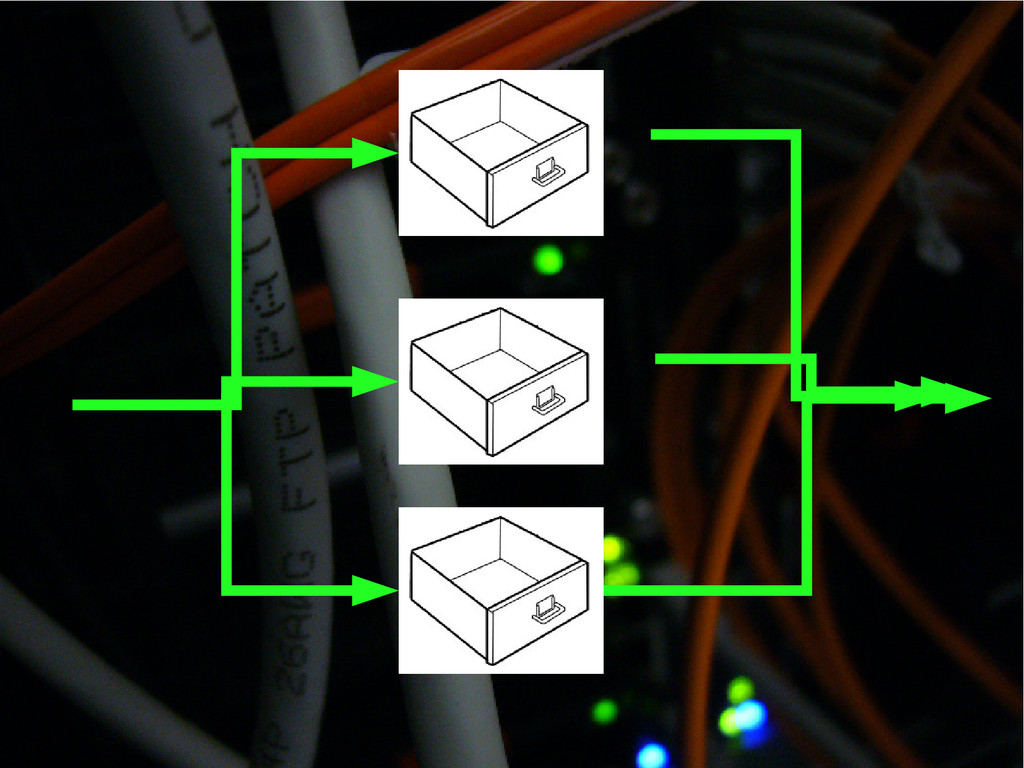
  2. GNU parallel is a shell tool for executing jobs in

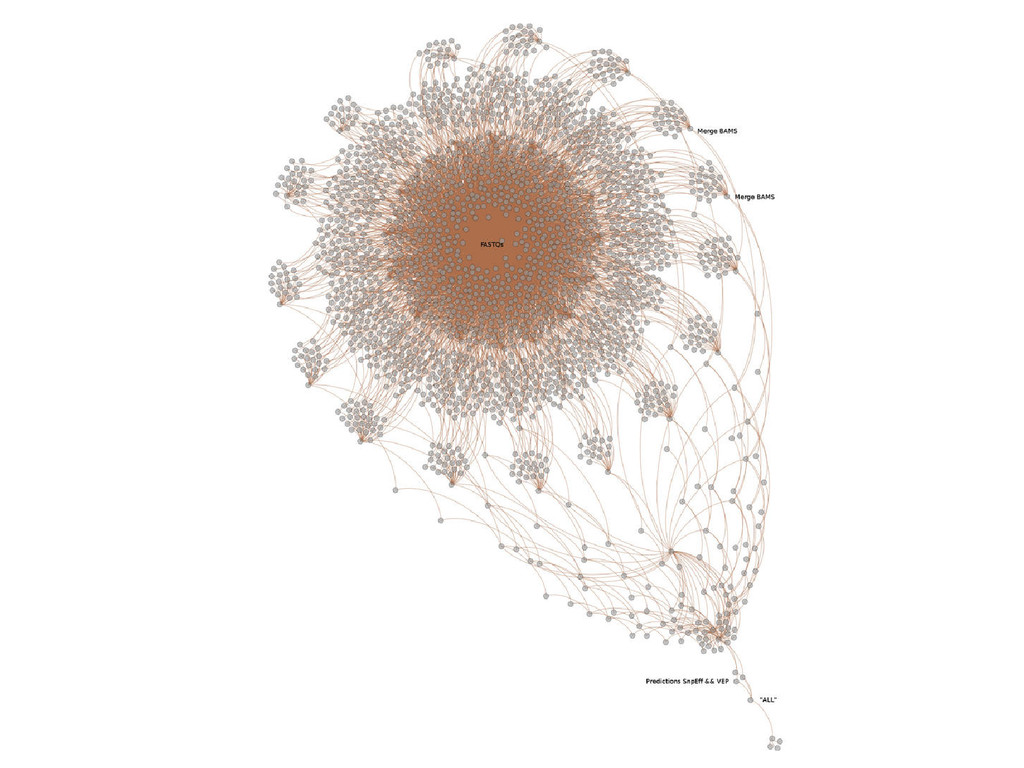
    parallel using one or more computers.
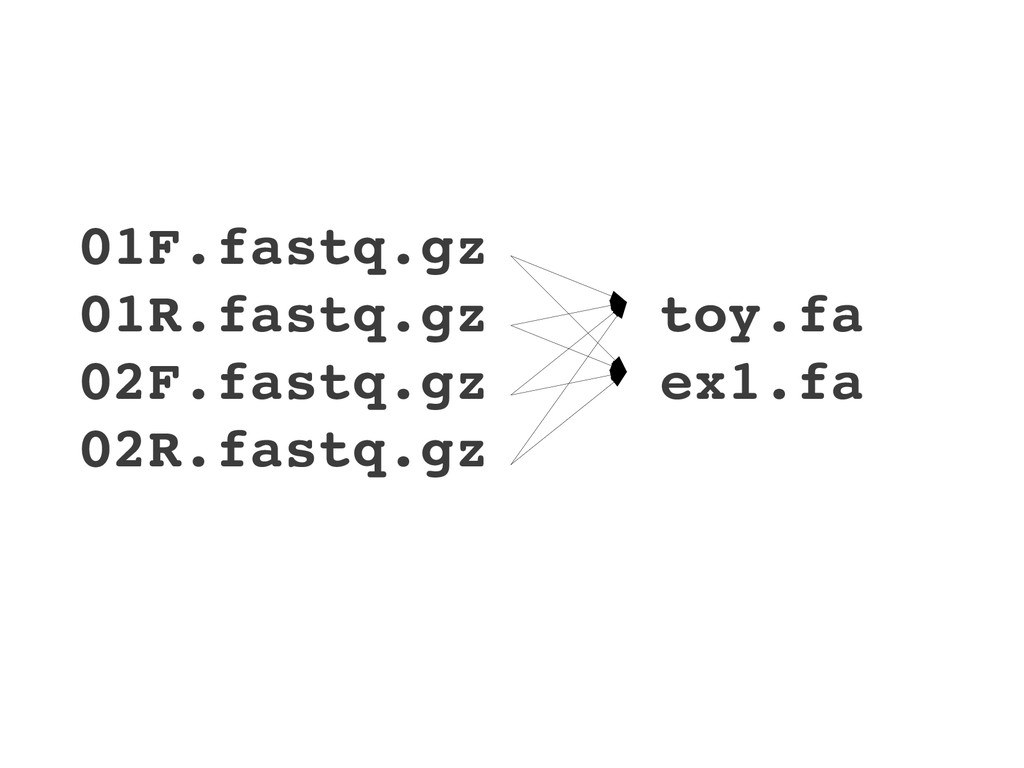
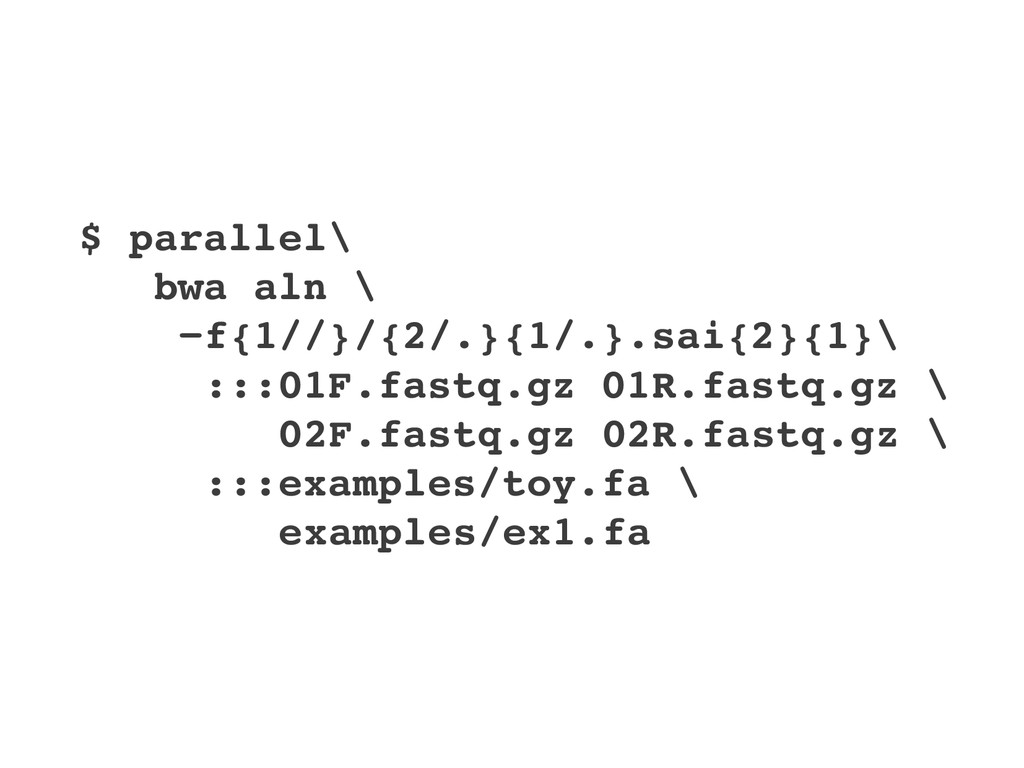
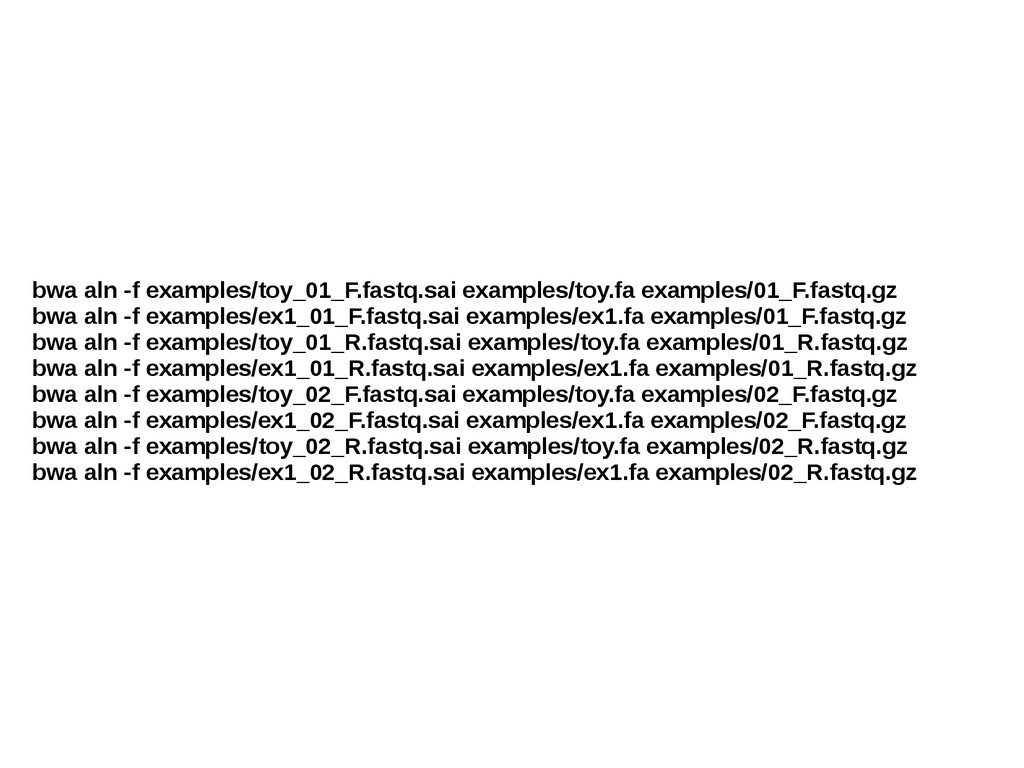
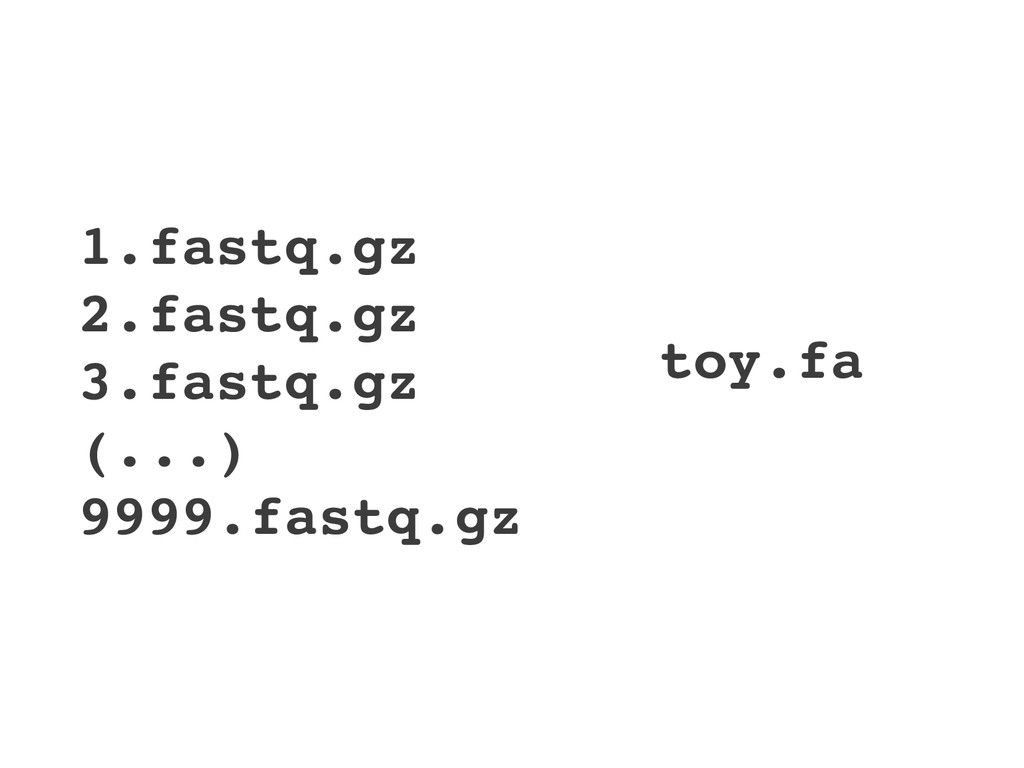
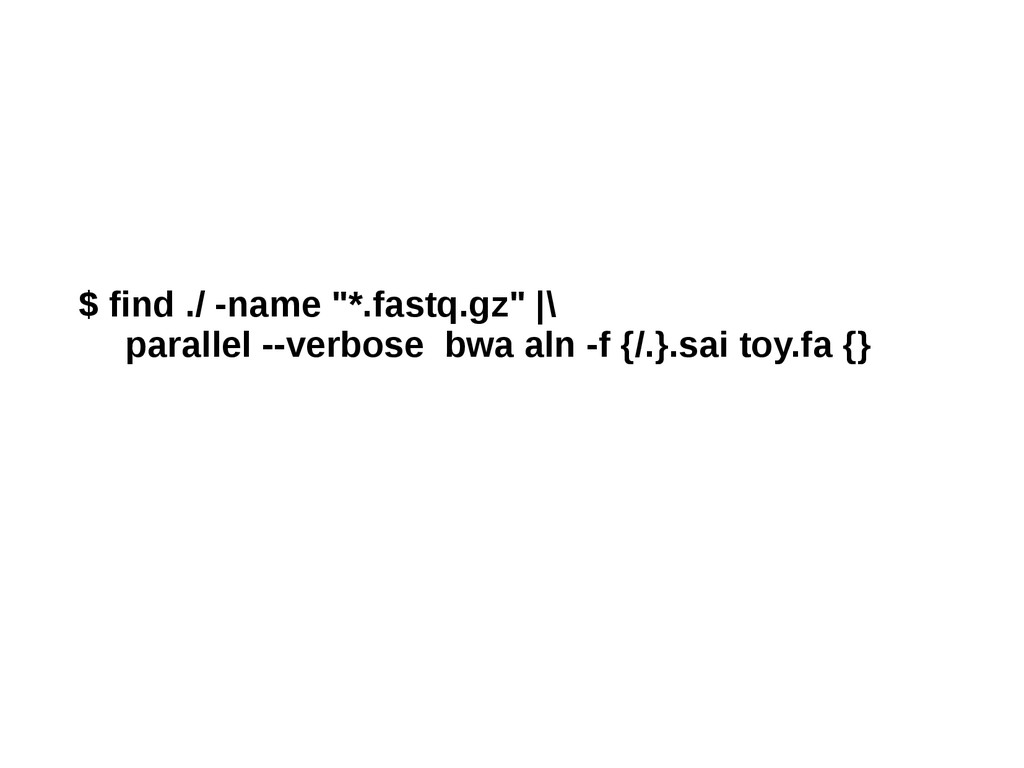
  3. bwa aln -f examples/toy_01_F.fastq.sai examples/toy.fa examples/01_F.fastq.gz bwa aln -f examples/ex1_01_F.fastq.sai

    examples/ex1.fa examples/01_F.fastq.gz bwa aln -f examples/toy_01_R.fastq.sai examples/toy.fa examples/01_R.fastq.gz bwa aln -f examples/ex1_01_R.fastq.sai examples/ex1.fa examples/01_R.fastq.gz bwa aln -f examples/toy_02_F.fastq.sai examples/toy.fa examples/02_F.fastq.gz bwa aln -f examples/ex1_02_F.fastq.sai examples/ex1.fa examples/02_F.fastq.gz bwa aln -f examples/toy_02_R.fastq.sai examples/toy.fa examples/02_R.fastq.gz bwa aln -f examples/ex1_02_R.fastq.sai examples/ex1.fa examples/02_R.fastq.gz
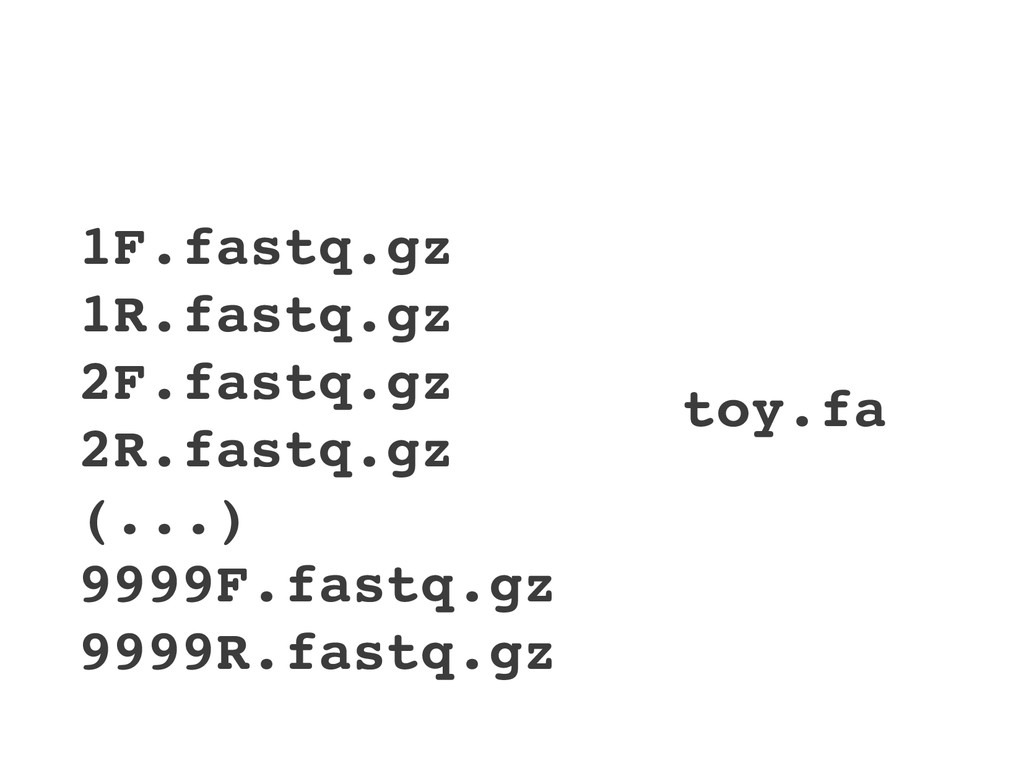
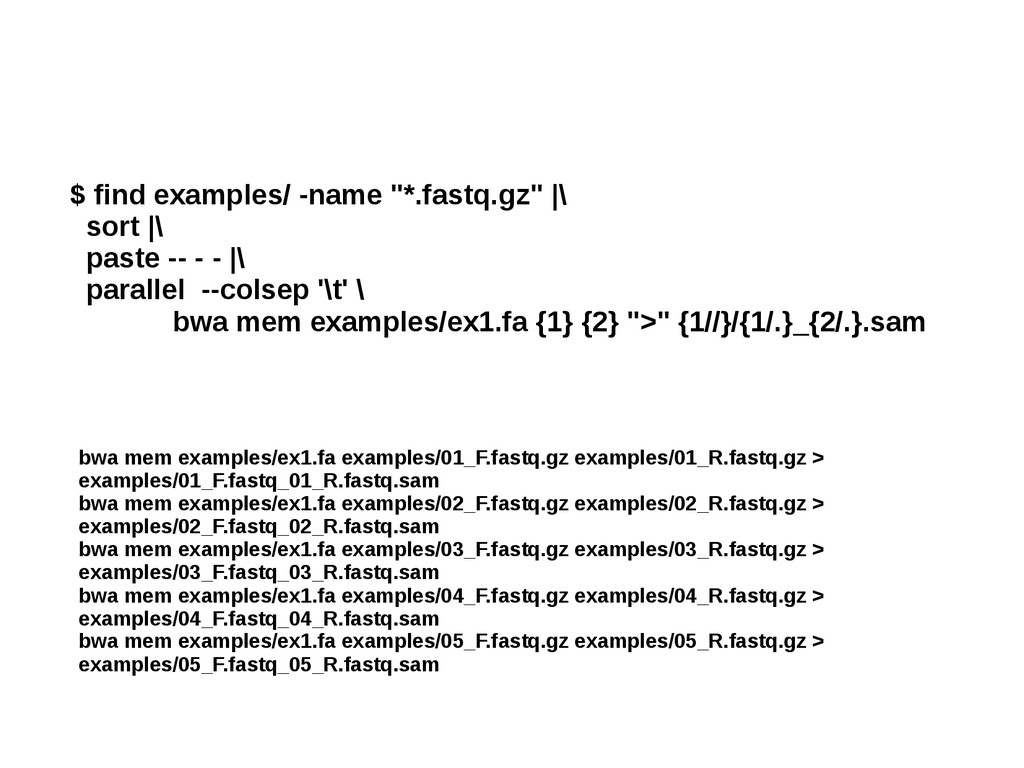
  4. $ find examples/ -name "*.fastq.gz" |\ sort |\ paste --

    - - |\ parallel --colsep '\t' \ bwa mem examples/ex1.fa {1} {2} ">" {1//}/{1/.}_{2/.}.sam bwa mem examples/ex1.fa examples/01_F.fastq.gz examples/01_R.fastq.gz > examples/01_F.fastq_01_R.fastq.sam bwa mem examples/ex1.fa examples/02_F.fastq.gz examples/02_R.fastq.gz > examples/02_F.fastq_02_R.fastq.sam bwa mem examples/ex1.fa examples/03_F.fastq.gz examples/03_R.fastq.gz > examples/03_F.fastq_03_R.fastq.sam bwa mem examples/ex1.fa examples/04_F.fastq.gz examples/04_R.fastq.gz > examples/04_F.fastq_04_R.fastq.sam bwa mem examples/ex1.fa examples/05_F.fastq.gz examples/05_R.fastq.gz > examples/05_F.fastq_05_R.fastq.sam
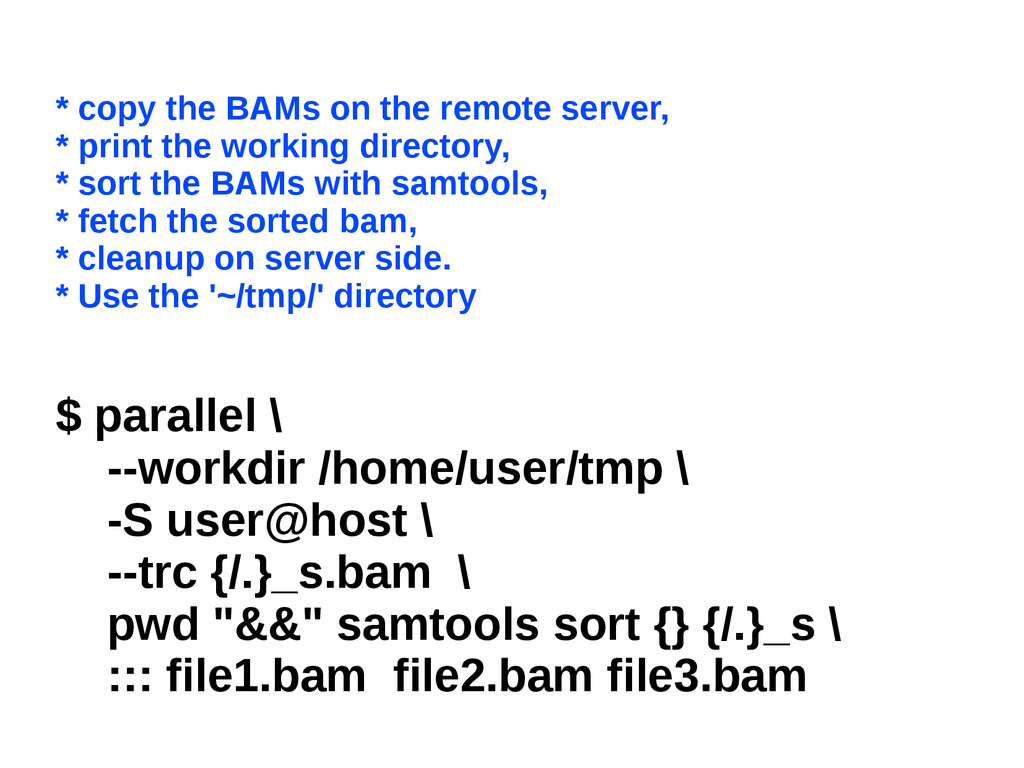
  5. * copy the BAMs on the remote server, * print

    the working directory, * sort the BAMs with samtools, * fetch the sorted bam, * cleanup on server side. * Use the '~/tmp/' directory $ parallel \ --workdir /home/user/tmp \ -S user@host \ --trc {/.}_s.bam \ pwd "&&" samtools sort {} {/.}_s \ ::: file1.bam file2.bam file3.bam
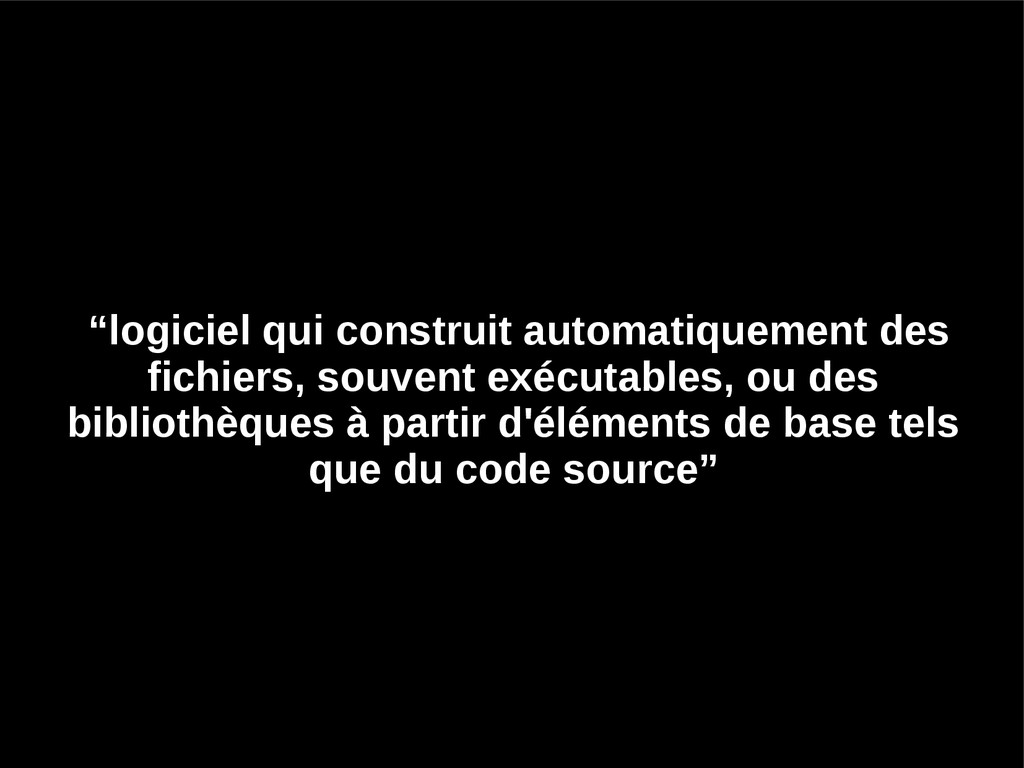
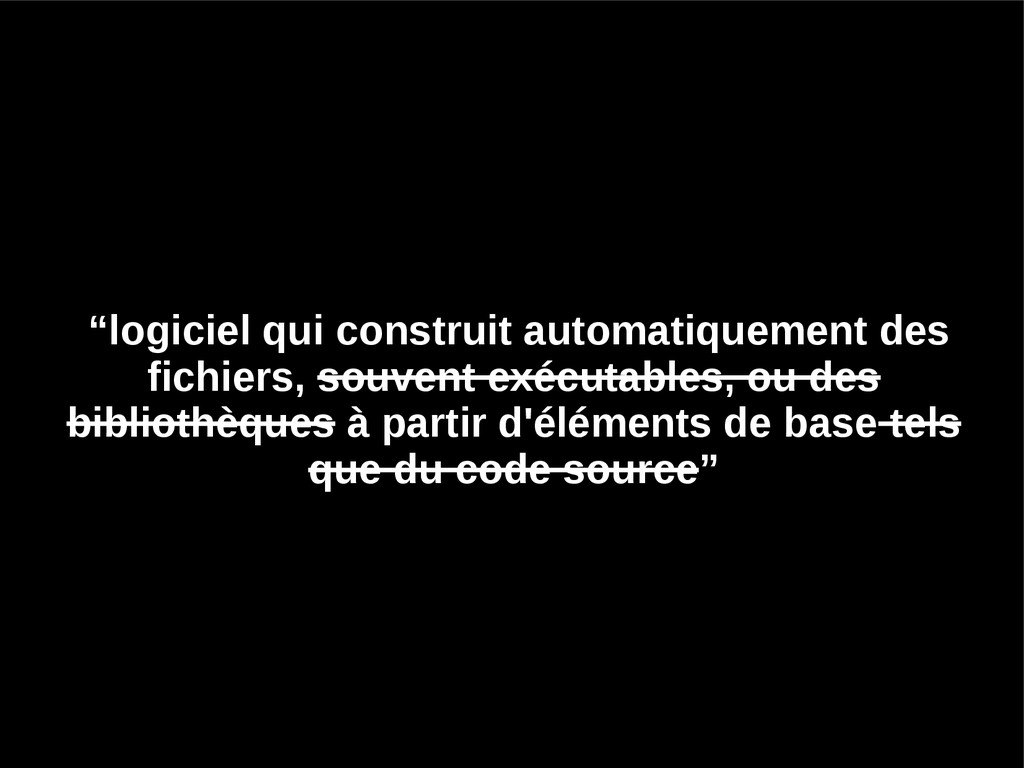
  6. “logiciel qui construit automatiquement des fichiers, souvent exécutables, ou des

    bibliothèques à partir d'éléments de base tels que du code source”
  7. “logiciel qui construit automatiquement des fichiers, souvent exécutables, ou des

    bibliothèques à partir d'éléments de base tels que du code source”
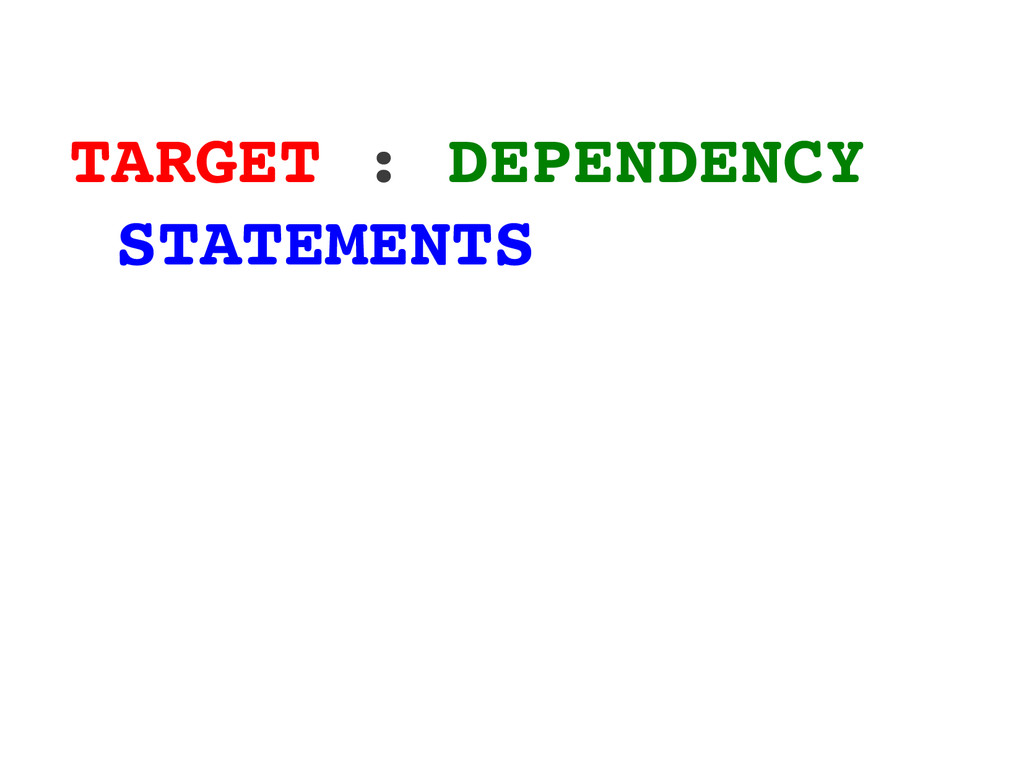
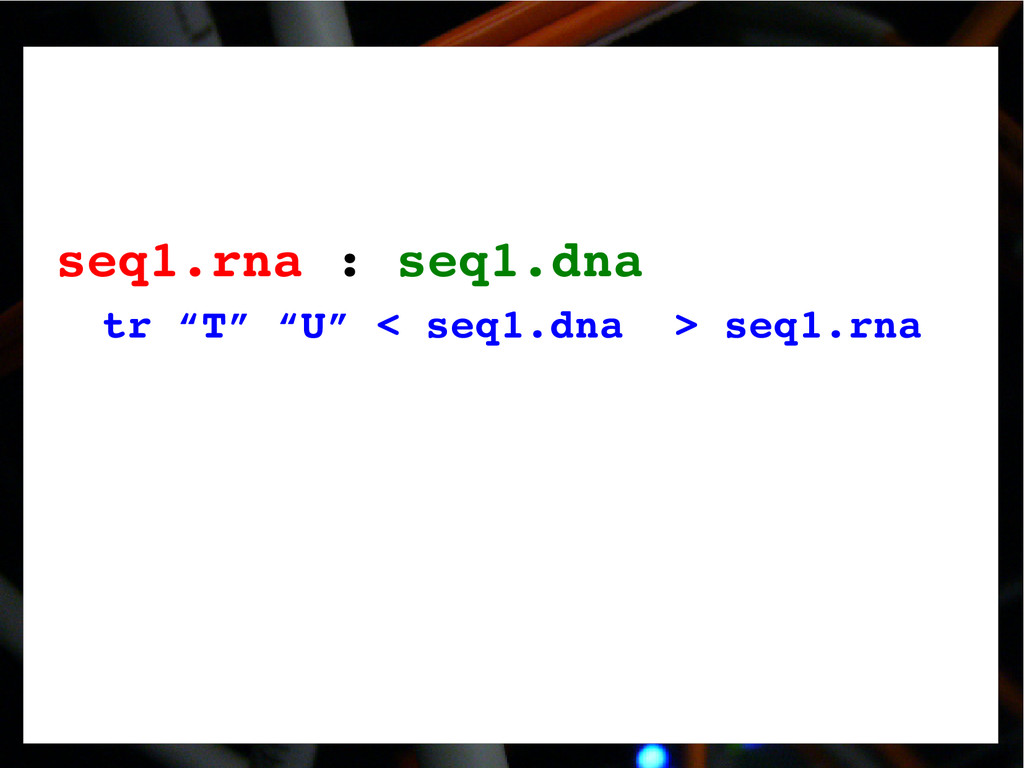
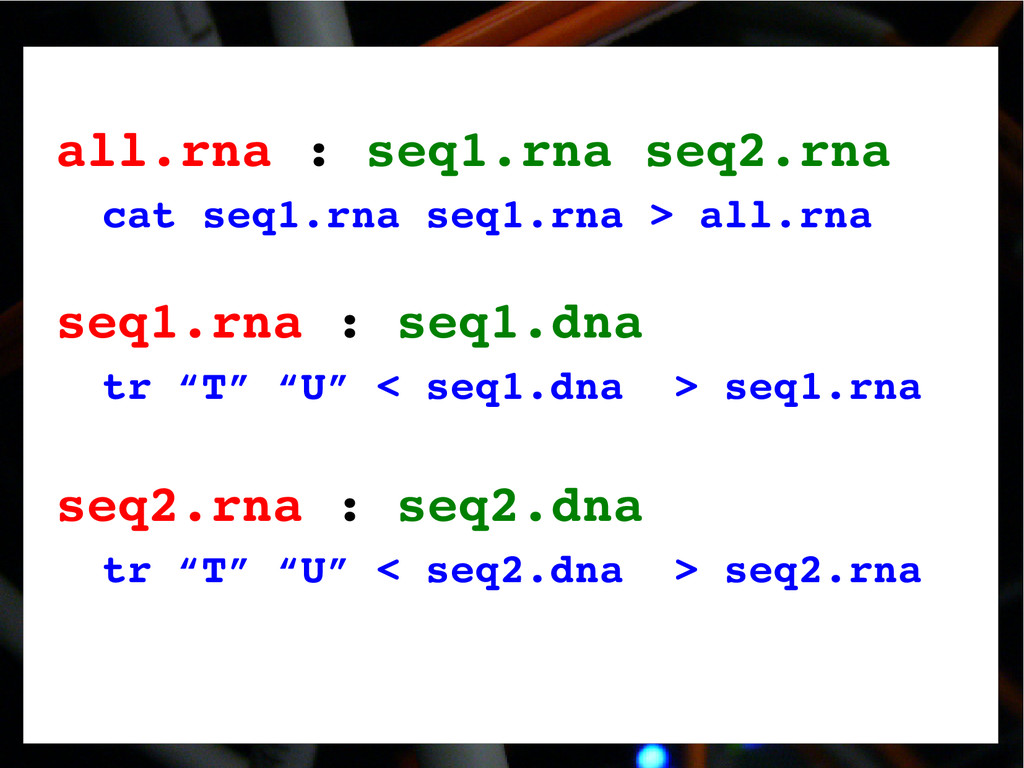
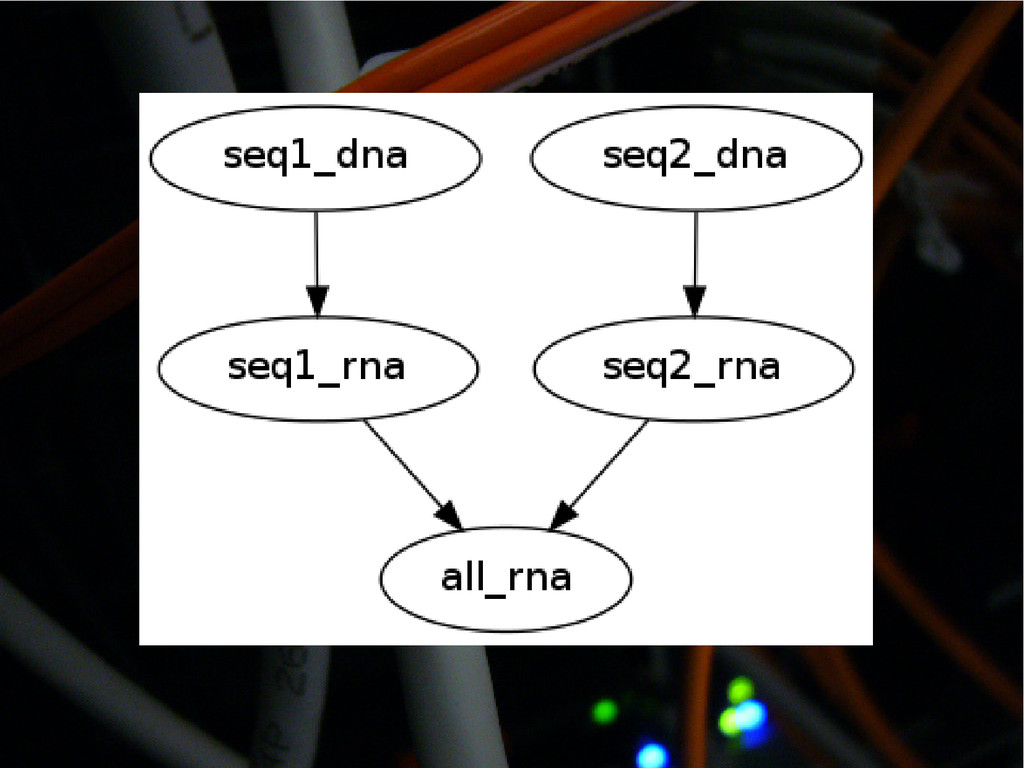
  8. all.rna : seq1.rna seq2.rna cat seq1.rna seq1.rna > all.rna seq1.rna

    : seq1.dna tr “T” “U” < seq1.dna > seq1.rna seq2.rna : seq2.dna tr “T” “U” < seq2.dna > seq2.rna
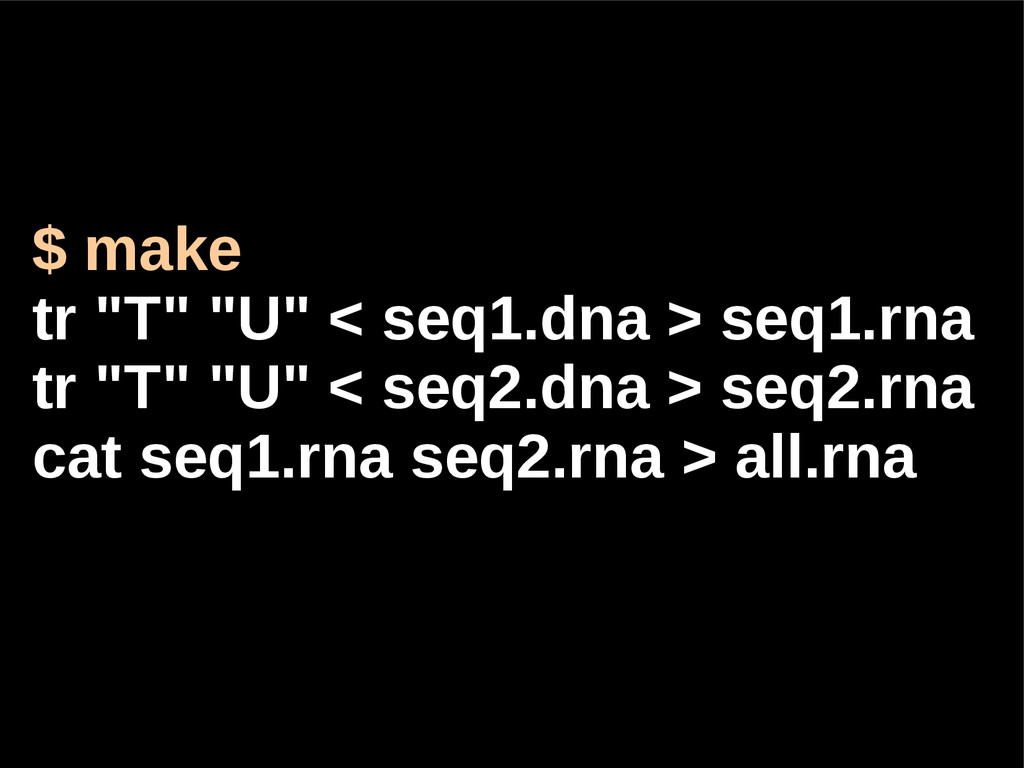
  9. $ make tr "T" "U" < seq1.dna > seq1.rna tr

    "T" "U" < seq2.dna > seq2.rna cat seq1.rna seq2.rna > all.rna
  10. v

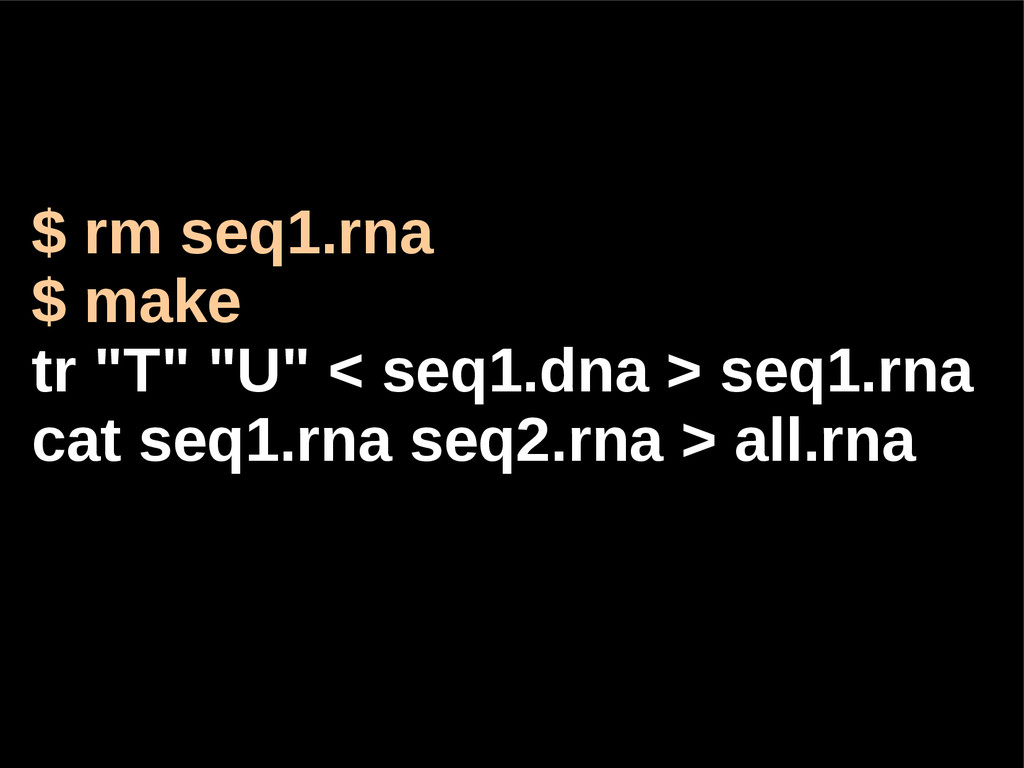
  11. $ rm seq1.rna $ make tr "T" "U" < seq1.dna

    > seq1.rna cat seq1.rna seq2.rna > all.rna
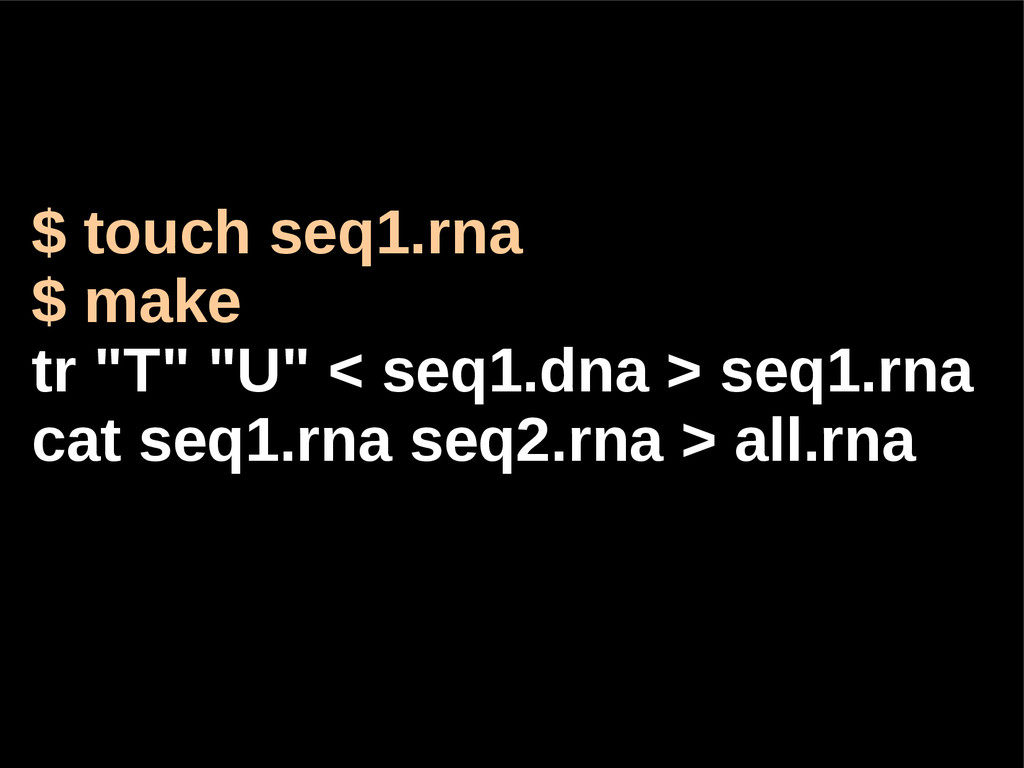
  12. $ touch seq1.rna $ make tr "T" "U" < seq1.dna

    > seq1.rna cat seq1.rna seq2.rna > all.rna
  13. NAME qmake - distributed parallel make, scheduling by Sun Grid

    Engine. SYNTAX qmake [ options ] -- [ gmake options ] DESCRIPTION Qmake is a parallel, distributed make utility. Scheduling of the parallel make tasks is done by Sun Grid Engine. It is based on gmake (GNU make), version 3.78.1. Both Sun Grid Engine and gmake command line options can be specified. They are separated by "--".
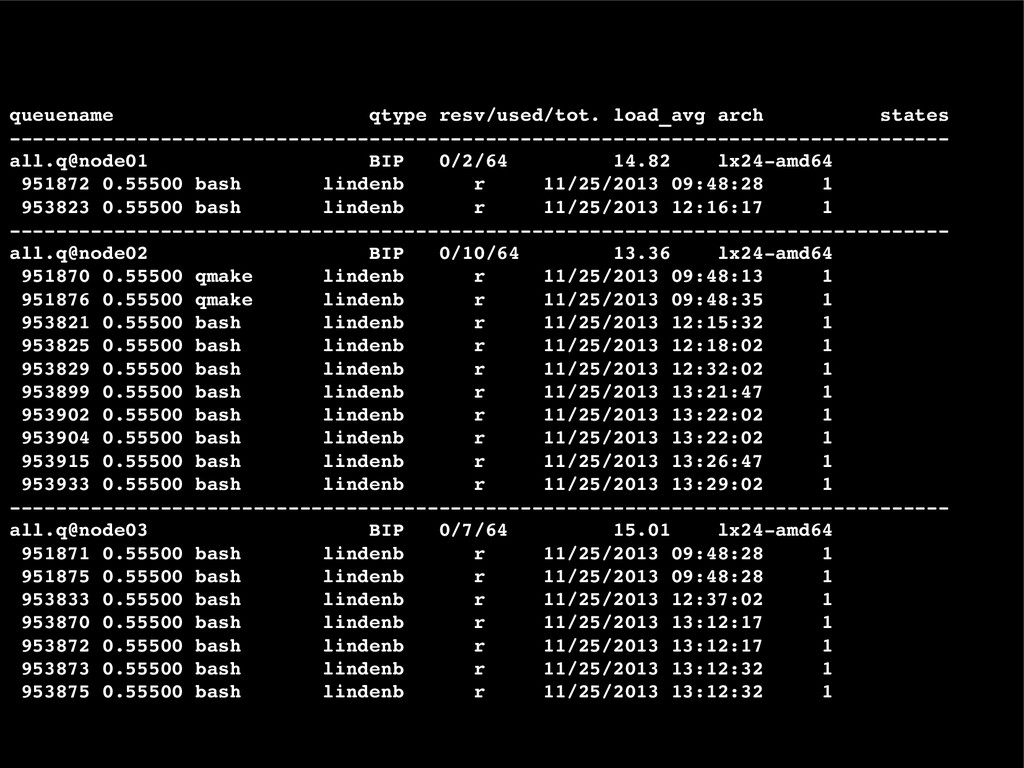
  14. queuename qtype resv/used/tot. load_avg arch states ­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­ all.q@node01 BIP 0/2/64

    14.82 lx24­amd64 951872 0.55500 bash lindenb r 11/25/2013 09:48:28 1 953823 0.55500 bash lindenb r 11/25/2013 12:16:17 1 ­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­ all.q@node02 BIP 0/10/64 13.36 lx24­amd64 951870 0.55500 qmake lindenb r 11/25/2013 09:48:13 1 951876 0.55500 qmake lindenb r 11/25/2013 09:48:35 1 953821 0.55500 bash lindenb r 11/25/2013 12:15:32 1 953825 0.55500 bash lindenb r 11/25/2013 12:18:02 1 953829 0.55500 bash lindenb r 11/25/2013 12:32:02 1 953899 0.55500 bash lindenb r 11/25/2013 13:21:47 1 953902 0.55500 bash lindenb r 11/25/2013 13:22:02 1 953904 0.55500 bash lindenb r 11/25/2013 13:22:02 1 953915 0.55500 bash lindenb r 11/25/2013 13:26:47 1 953933 0.55500 bash lindenb r 11/25/2013 13:29:02 1 ­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­­ all.q@node03 BIP 0/7/64 15.01 lx24­amd64 951871 0.55500 bash lindenb r 11/25/2013 09:48:28 1 951875 0.55500 bash lindenb r 11/25/2013 09:48:28 1 953833 0.55500 bash lindenb r 11/25/2013 12:37:02 1 953870 0.55500 bash lindenb r 11/25/2013 13:12:17 1 953872 0.55500 bash lindenb r 11/25/2013 13:12:17 1 953873 0.55500 bash lindenb r 11/25/2013 13:12:32 1 953875 0.55500 bash lindenb r 11/25/2013 13:12:32 1
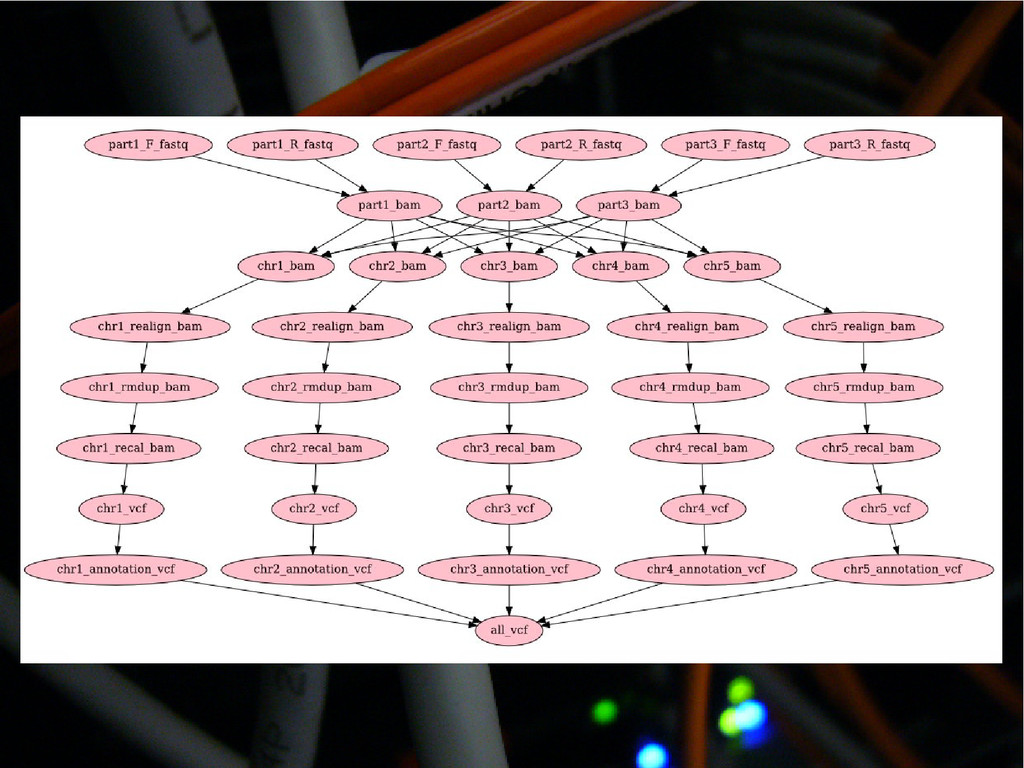
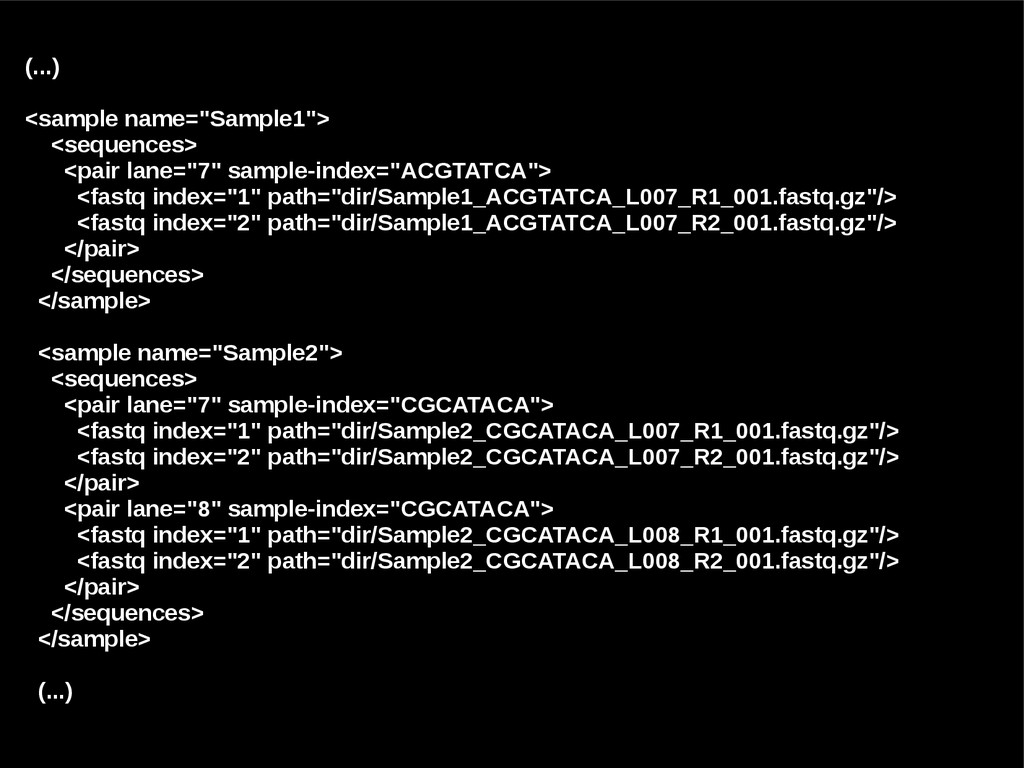
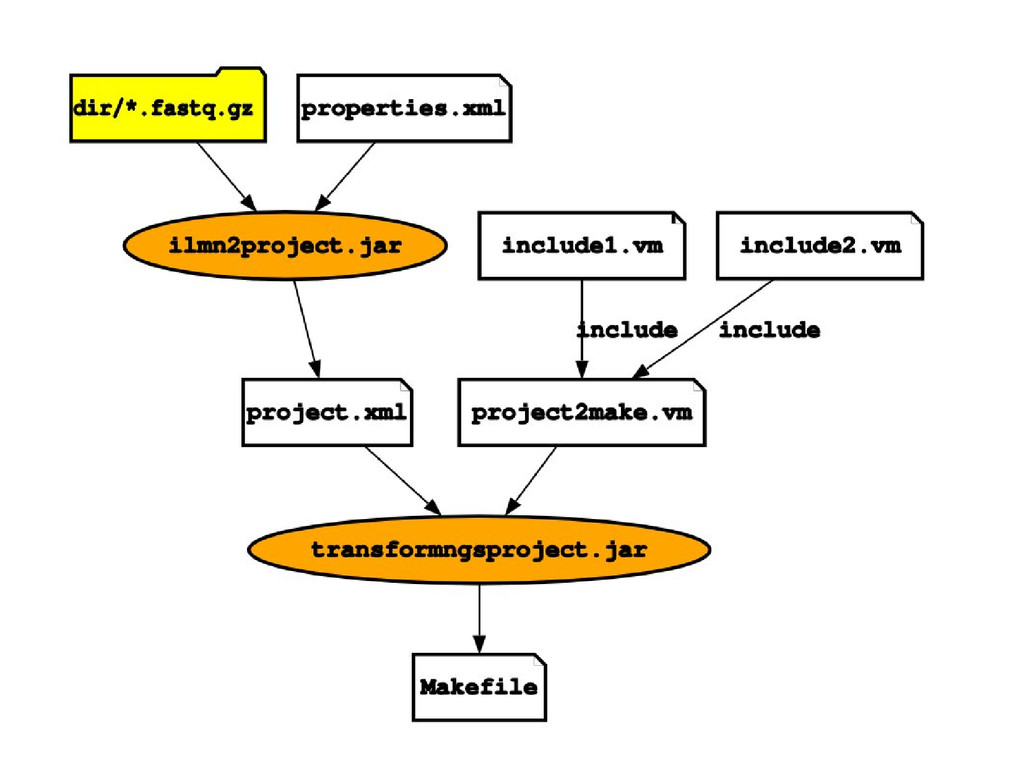
  15. (...) <sample name="Sample1"> <sequences> <pair lane="7" sample-index="ACGTATCA"> <fastq index="1" path="dir/Sample1_ACGTATCA_L007_R1_001.fastq.gz"/>

    <fastq index="2" path="dir/Sample1_ACGTATCA_L007_R2_001.fastq.gz"/> </pair> </sequences> </sample> <sample name="Sample2"> <sequences> <pair lane="7" sample-index="CGCATACA"> <fastq index="1" path="dir/Sample2_CGCATACA_L007_R1_001.fastq.gz"/> <fastq index="2" path="dir/Sample2_CGCATACA_L007_R2_001.fastq.gz"/> </pair> <pair lane="8" sample-index="CGCATACA"> <fastq index="1" path="dir/Sample2_CGCATACA_L008_R1_001.fastq.gz"/> <fastq index="2" path="dir/Sample2_CGCATACA_L008_R2_001.fastq.gz"/> </pair> </sequences> </sample> (...)
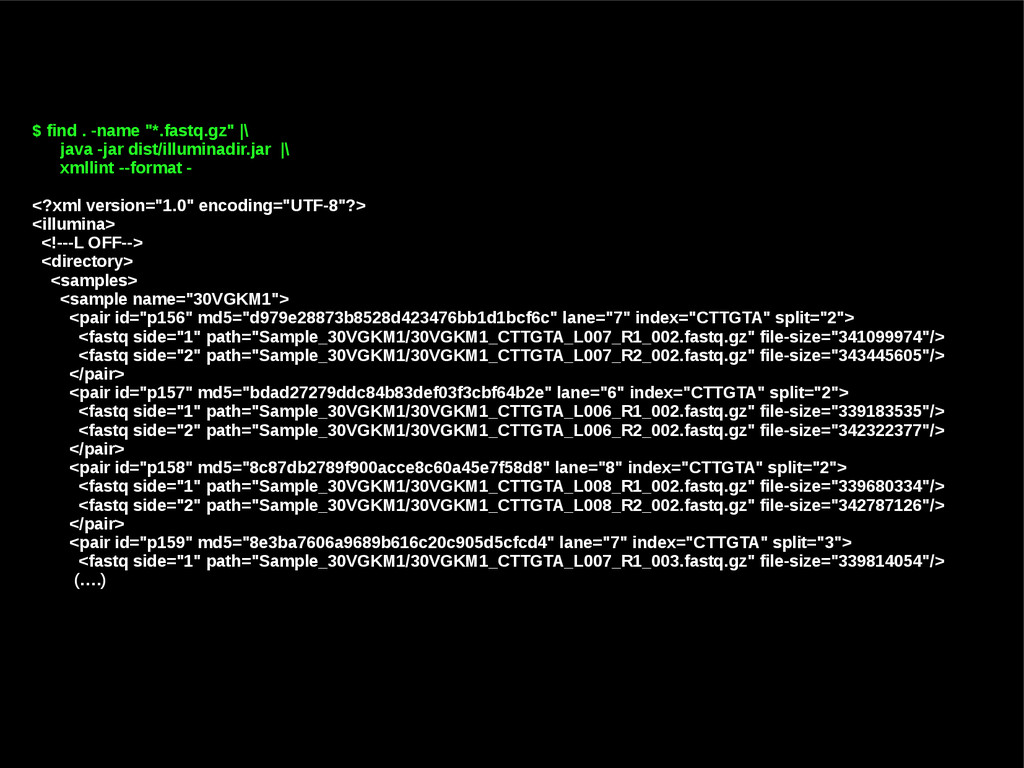
  16. $ find . -name "*.fastq.gz" |\ java -jar dist/illuminadir.jar |\

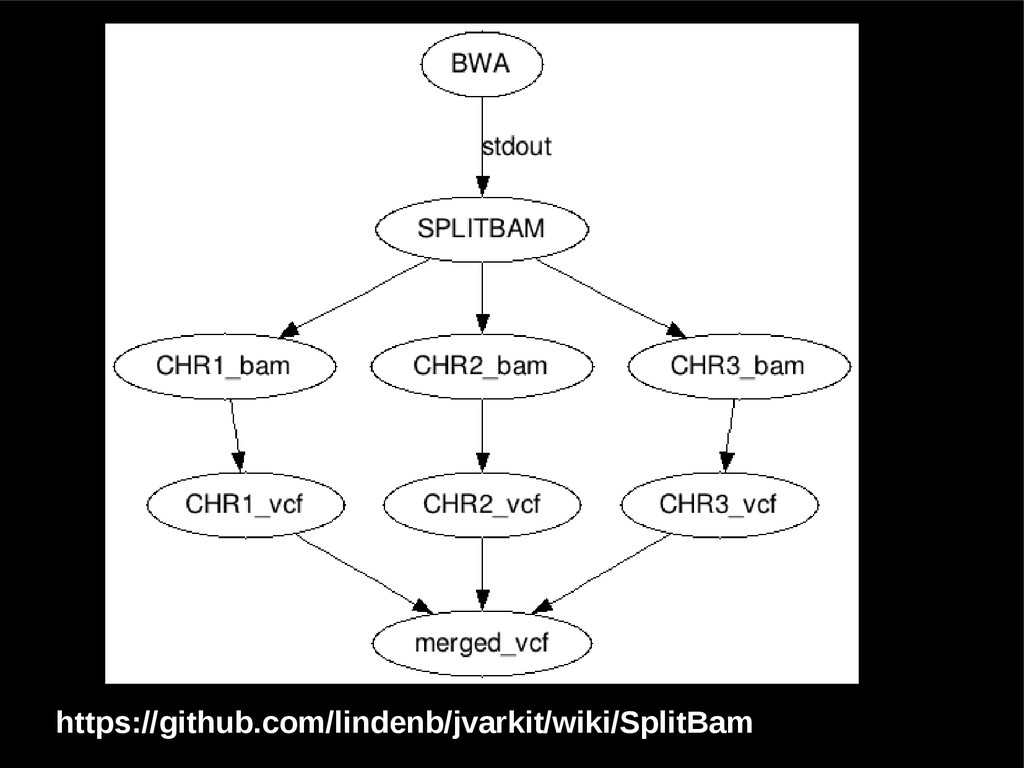
    xmllint --format - <?xml version="1.0" encoding="UTF-8"?> <illumina> <!---L OFF--> <directory> <samples> <sample name="30VGKM1"> <pair id="p156" md5="d979e28873b8528d423476bb1d1bcf6c" lane="7" index="CTTGTA" split="2"> <fastq side="1" path="Sample_30VGKM1/30VGKM1_CTTGTA_L007_R1_002.fastq.gz" file-size="341099974"/> <fastq side="2" path="Sample_30VGKM1/30VGKM1_CTTGTA_L007_R2_002.fastq.gz" file-size="343445605"/> </pair> <pair id="p157" md5="bdad27279ddc84b83def03f3cbf64b2e" lane="6" index="CTTGTA" split="2"> <fastq side="1" path="Sample_30VGKM1/30VGKM1_CTTGTA_L006_R1_002.fastq.gz" file-size="339183535"/> <fastq side="2" path="Sample_30VGKM1/30VGKM1_CTTGTA_L006_R2_002.fastq.gz" file-size="342322377"/> </pair> <pair id="p158" md5="8c87db2789f900acce8c60a45e7f58d8" lane="8" index="CTTGTA" split="2"> <fastq side="1" path="Sample_30VGKM1/30VGKM1_CTTGTA_L008_R1_002.fastq.gz" file-size="339680334"/> <fastq side="2" path="Sample_30VGKM1/30VGKM1_CTTGTA_L008_R2_002.fastq.gz" file-size="342787126"/> </pair> <pair id="p159" md5="8e3ba7606a9689b616c20c905d5cfcd4" lane="7" index="CTTGTA" split="3"> <fastq side="1" path="Sample_30VGKM1/30VGKM1_CTTGTA_L007_R1_003.fastq.gz" file-size="339814054"/> (….)
  17. [{"directory":"RUN62_XFC2DM8ACXX/data","samples":[{"sample":"SAMPLE1","files":[{ "md5pair":"cd4b436ce7aff4cf669d282c6d9a7899","lane":8,"index":"ATCACG","split":2 ,"forward":{"md5filename":"3369c3457d6603f06379b654cb78e696","path":"20131001_SN L149_0062_XFC2DM8ACXX/data/OUT/Sample_SAMPLE1/SAMPLE1_ATCACG_L008_R1_002.fastq.g z","side":1,"file-size":359046311},"reverse":{"md5filename":"832039fa00b5f401088 48e48eb437e0b","path":"20131001_SNL149_0062_XFC2DM8ACXX/data/OUT/Sample_SAMPLE1/ SAMPLE1_ATCACG_L008_R2_002.fastq.gz","side":2,"file-size":359659451}},{"md5pair" :"b3050fa3307e63ab9790b0e263c5d240","lane":8,"index":"ATCACG","split":3,"forward ":{"md5filename":"091727bb6b300e463c3d708e157436ab","path":"20131001_SNL149_0062 _XFC2DM8ACXX/data/OUT/Sample_SAMPLE1/SAMPLE1_ATCACG_L008_R1_003.fastq.gz","side"

    :1,"file-size":206660736},"reverse":{"md5filename":"20235ef4ec8845515beb4e13da34 b5d3","path":"20131001_SNL149_0062_XFC2DM8ACXX/data/OUT/Sample_SAMPLE1/SAMPLE1_A TCACG_L008_R2_003.fastq.gz","side":2,"file-size":206715143}},{"md5pair":"9f7ee49 e87d01610372c43ab928939f6","lane":8,"index":"ATCACG","split":1,"forward":{"md5fi lename":"54cb2fd33edd5c2e787287ccf1595952","path":"20131001_SNL149_0062_XFC2DM8A CXX/data/OUT/Sample_SAMPLE1/SAMPLE1_ATCACG_L008_R1_001.fastq.gz","side":1,"file- size":354530831},"reverse":{"md5filename":"e937cbdf32020074e50d3332c67cf6b3","pa th":"20131001_SNL149_0062_XFC2DM8ACXX/data/OUT/Sample_SAMPLE1/SAMPLE1_ATCACG_L00 8_R2_001.fastq.gz","side":2,"file-size":356908963}},{"md5pair":"0697846a504158ee f523c0f4ede85288","lane":7,"index":"ATCACG","split":2,"forward":{"md5filename":" https://github.com/lindenb/jvarkit/wiki/Illuminadir
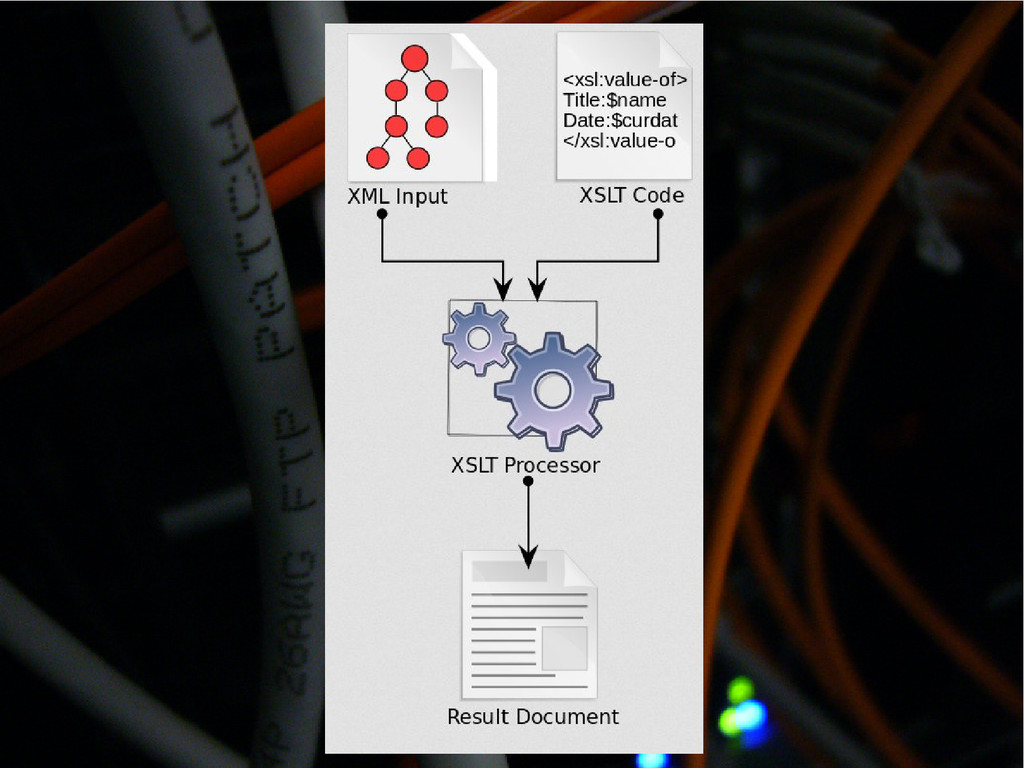
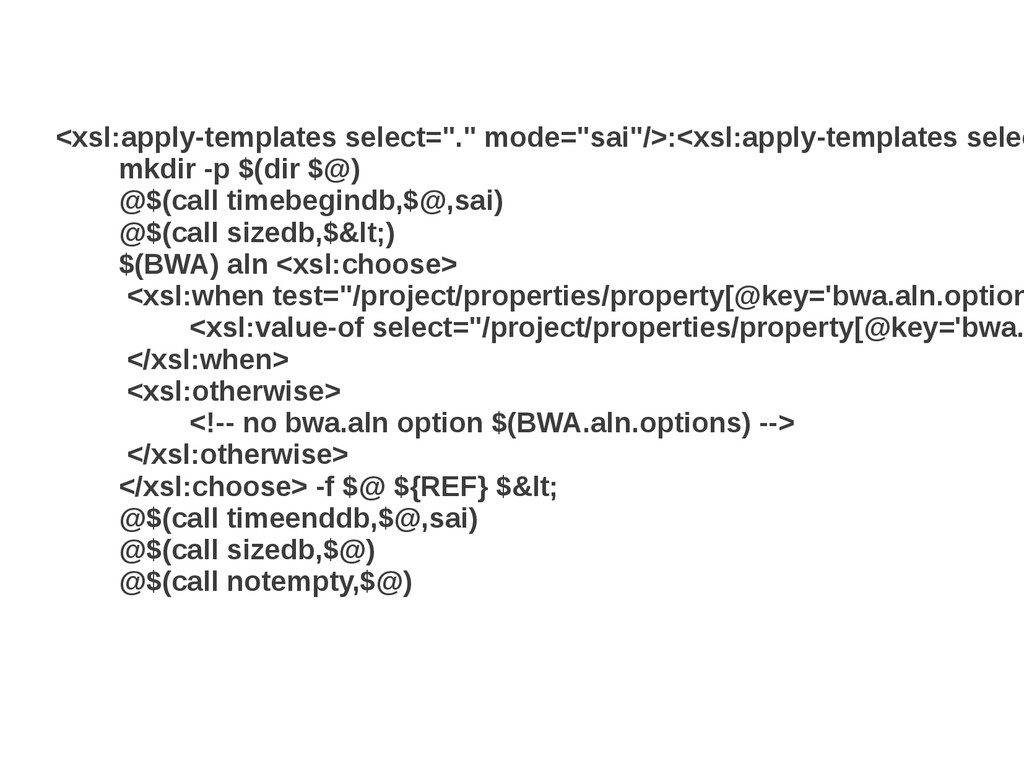
  18. <xsl:apply-templates select="." mode="sai"/>:<xsl:apply-templates selec mkdir -p $(dir $@) @$(call timebegindb,$@,sai)

    @$(call sizedb,$&lt;) $(BWA) aln <xsl:choose> <xsl:when test="/project/properties/property[@key='bwa.aln.option <xsl:value-of select="/project/properties/property[@key='bwa. </xsl:when> <xsl:otherwise> <!-- no bwa.aln option $(BWA.aln.options) --> </xsl:otherwise> </xsl:choose> -f $@ ${REF} $&lt; @$(call timeenddb,$@,sai) @$(call sizedb,$@) @$(call notempty,$@)
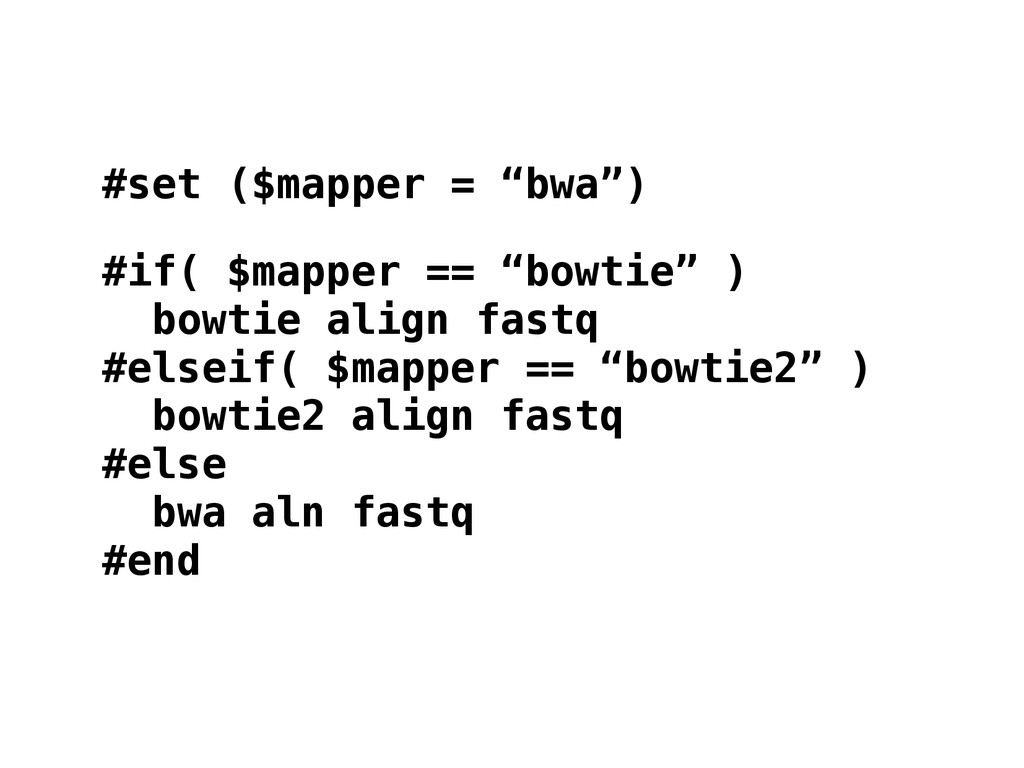
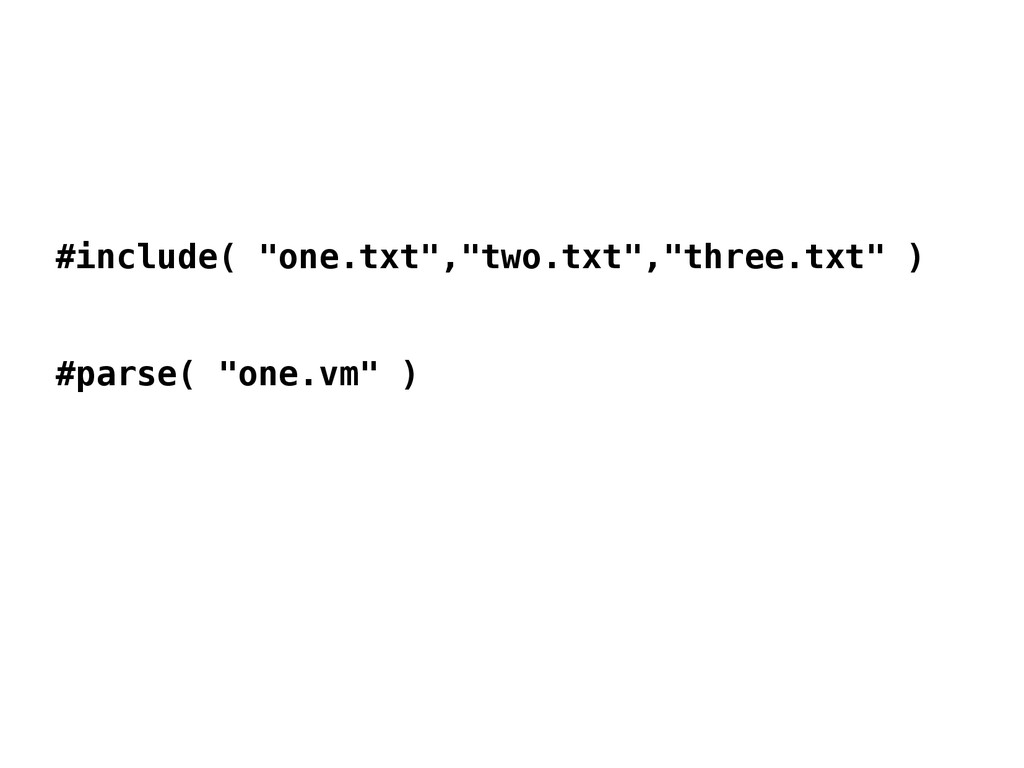
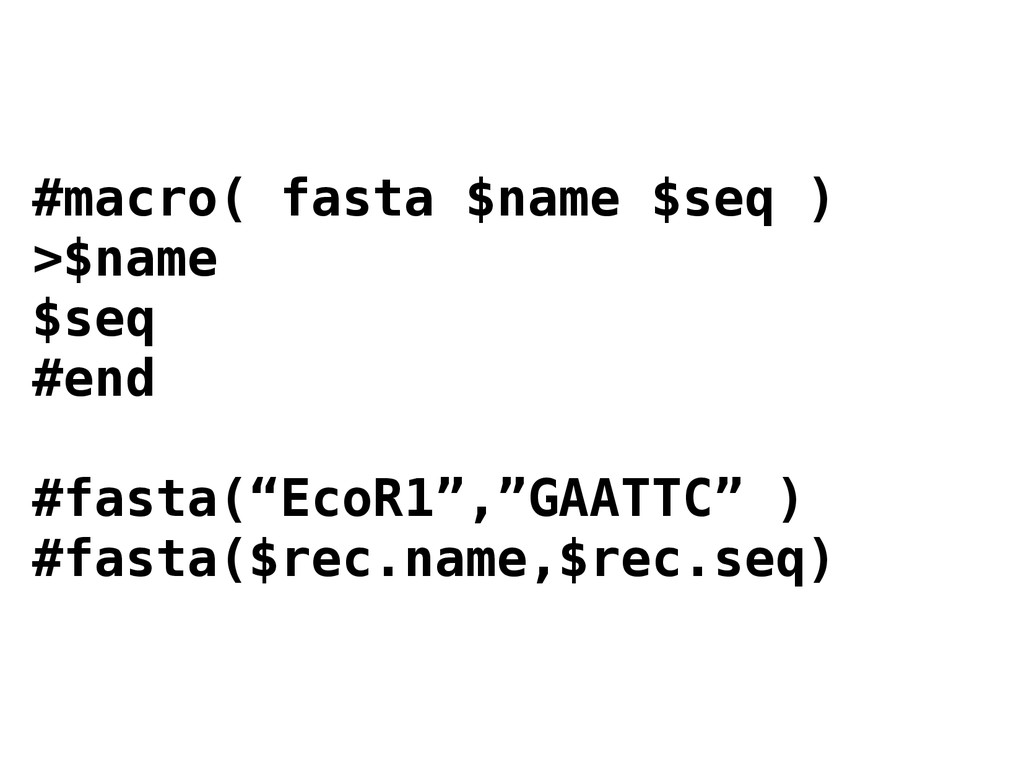
  19. #set ($mapper = “bwa”) #if( $mapper == “bowtie” ) bowtie

    align fastq #elseif( $mapper == “bowtie2” ) bowtie2 align fastq #else bwa aln fastq #end
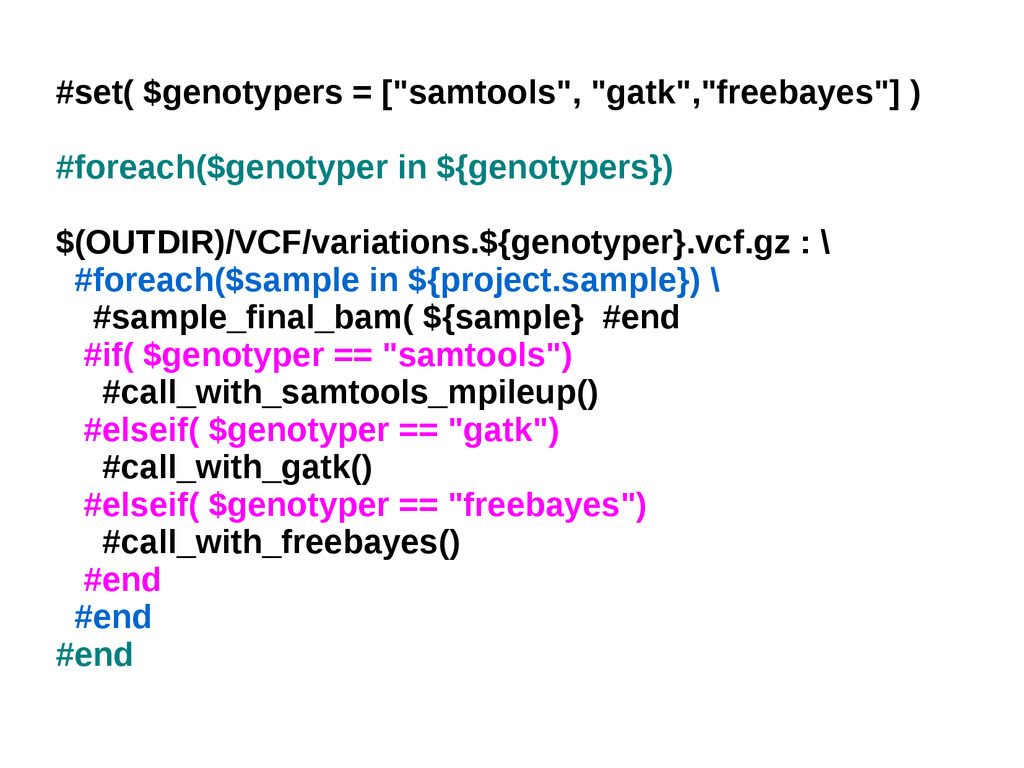
  20. #set( $genotypers = ["samtools", "gatk","freebayes"] ) #foreach($genotyper in ${genotypers}) $(OUTDIR)/VCF/variations.${genotyper}.vcf.gz

    : \ #foreach($sample in ${project.sample}) \ #sample_final_bam( ${sample} #end #if( $genotyper == "samtools") #call_with_samtools_mpileup() #elseif( $genotyper == "gatk") #call_with_gatk() #elseif( $genotyper == "freebayes") #call_with_freebayes() #end #end #end
  21. Can you test with a minimal DEPTH=20 ? And with

    minimal number of affected samples per SNP = 2 ? And with minimal number of affected samples per gene = 4 ? Can you remove all the known SNPs ? Did we try fix the maximum number of SNPs per gene ? Did we ever find this gene ? Did you include the results from Polyphen ?
  22. Merci Laetitia Duboscq-Bidot Mathieu Le-Neue Audrey Bihouée Eric Charpentier Edouard
    Hirchaud Flogiane Simoget Solena le Scouarnec Raluca Teusan Richard Redon The Biostars.org community