Share
Embed
Mining genetic variation in any species with GEMINI
Plant and Animal Genomics
January 9, 2016
Aaron Quinlan
January 10, 2016
Transcript
41 slides
-
Aaron Quinlan University of Utah quinlanlab.org @aaronquinlan Plant and Animal
Genome Conference January 9, 2016 Mining genetic variation in any species with GEMINI -
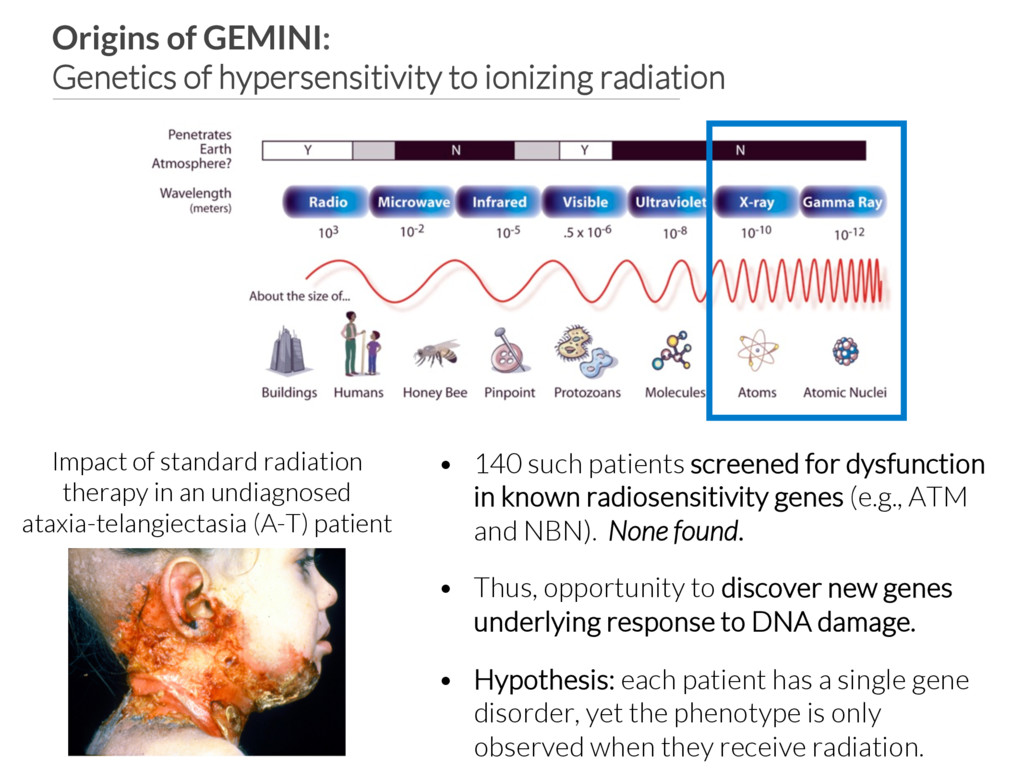
Origins of GEMINI: Genetics of hypersensitivity to ionizing radiation Impact
of standard radiation therapy in an undiagnosed ataxia-telangiectasia (A-T) patient • 140 such patients screened for dysfunction in known radiosensitivity genes (e.g., ATM and NBN). None found. • Thus, opportunity to discover new genes underlying response to DNA damage. • Hypothesis: each patient has a single gene disorder, yet the phenotype is only observed when they receive radiation. -
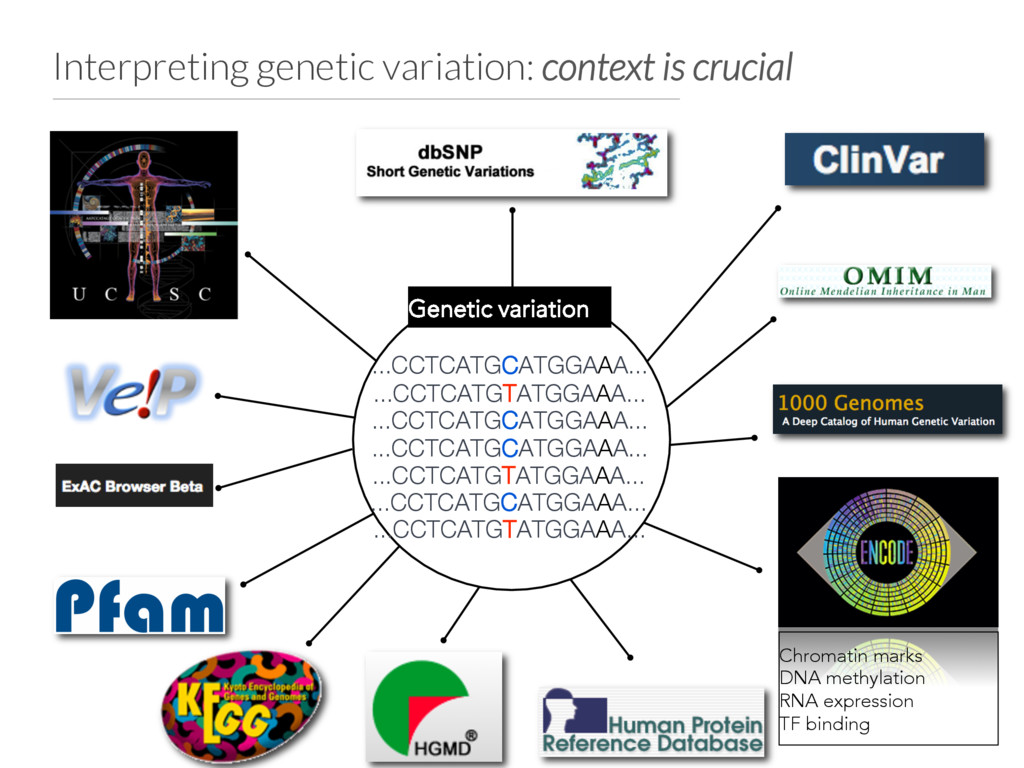
Interpreting genetic variation: context is crucial ...CCTCATGCATGGAAA... Genetic variation ...CCTCATGTATGGAAA...
...CCTCATGCATGGAAA... ...CCTCATGCATGGAAA... ...CCTCATGTATGGAAA... ...CCTCATGCATGGAAA... ...CCTCATGTATGGAAA... Chromatin marks DNA methylation RNA expression TF binding -
GEMINI: a flexible framework for exploring genome variation Uma Paila
Brad Chapman github.com/arq5x/gemini gemini.rtfd.org Brent Pedersen -
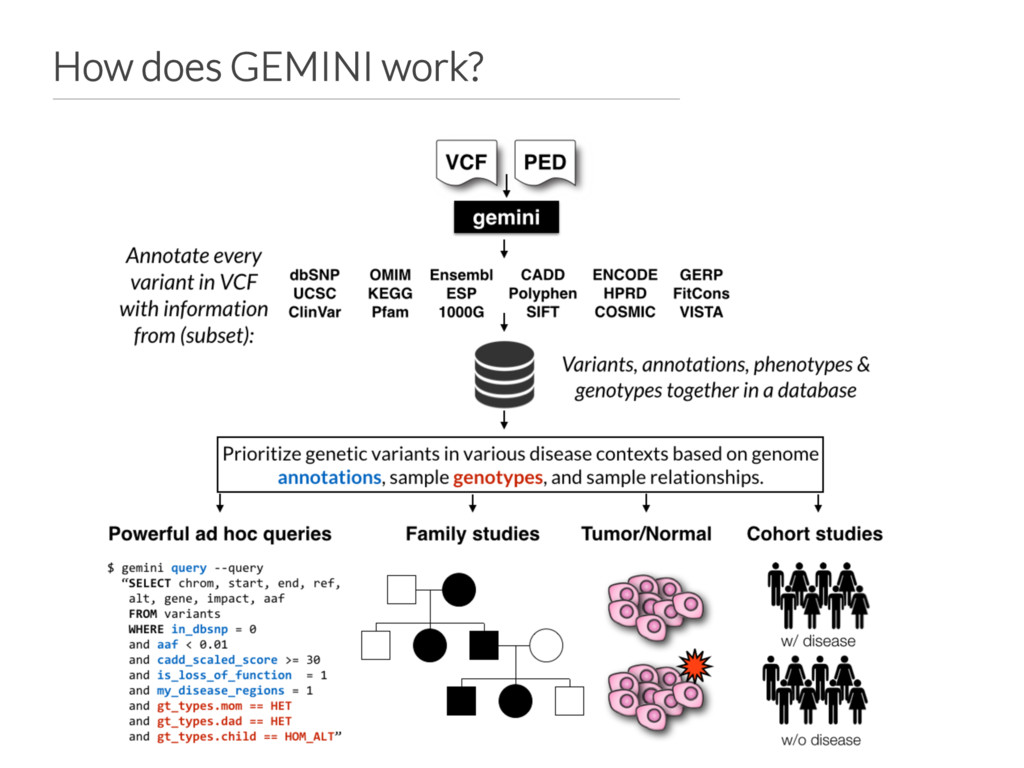
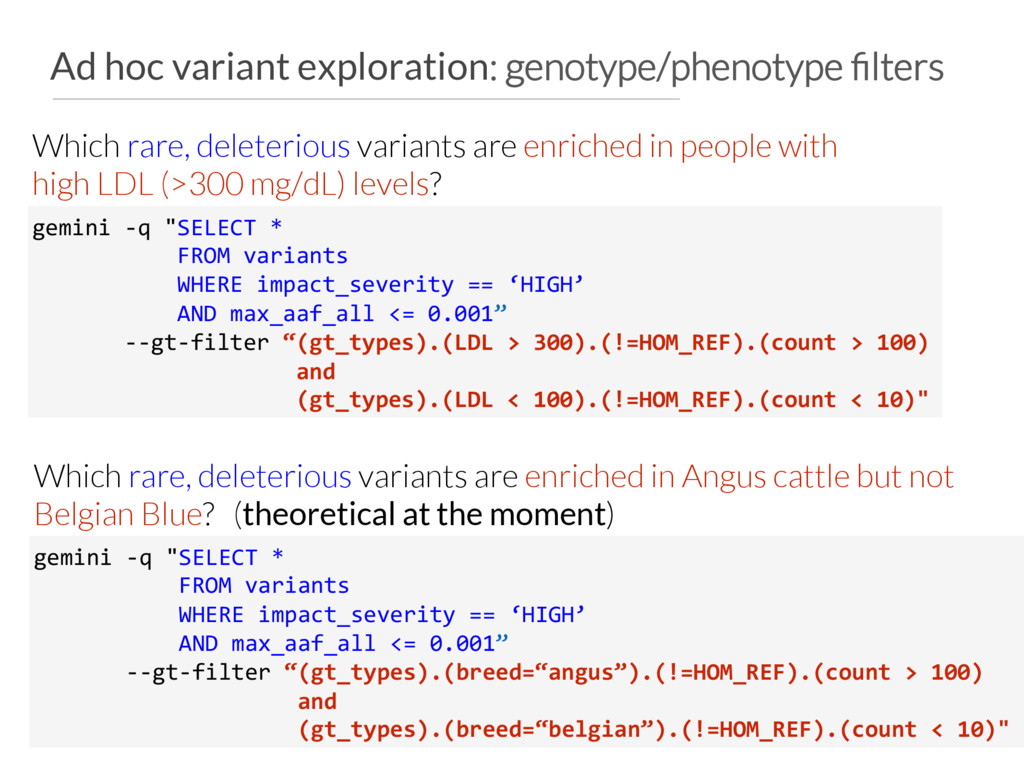
Ad hoc variant exploration: genotype/phenotype filters gemini -q "SELECT *
FROM variants WHERE impact_severity == ‘HIGH’ AND max_aaf_all <= 0.001” --gt-filter “(gt_types).(LDL > 300).(!=HOM_REF).(count > 100) and (gt_types).(LDL < 100).(!=HOM_REF).(count < 10)" Which rare, deleterious variants are enriched in people with high LDL (>300 mg/dL) levels? gemini -q "SELECT * FROM variants WHERE impact_severity == ‘HIGH’ AND max_aaf_all <= 0.001” --gt-filter “(gt_types).(breed=“angus”).(!=HOM_REF).(count > 100) and (gt_types).(breed=“belgian”).(!=HOM_REF).(count < 10)" Which rare, deleterious variants are enriched in Angus cattle but not Belgian Blue? (theoretical at the moment) -
Automated tools for disease inheritance models A/A A/G A/G Dominant
A/G G/G A/G Recessive (consang.) C/C A/G A/A A/G C/T C/T Recessive (compound heterozygous) A/A A/G A/A De novo -
GEMINI is popular for rare disease research. UW Center for
Mendelian Genomics -
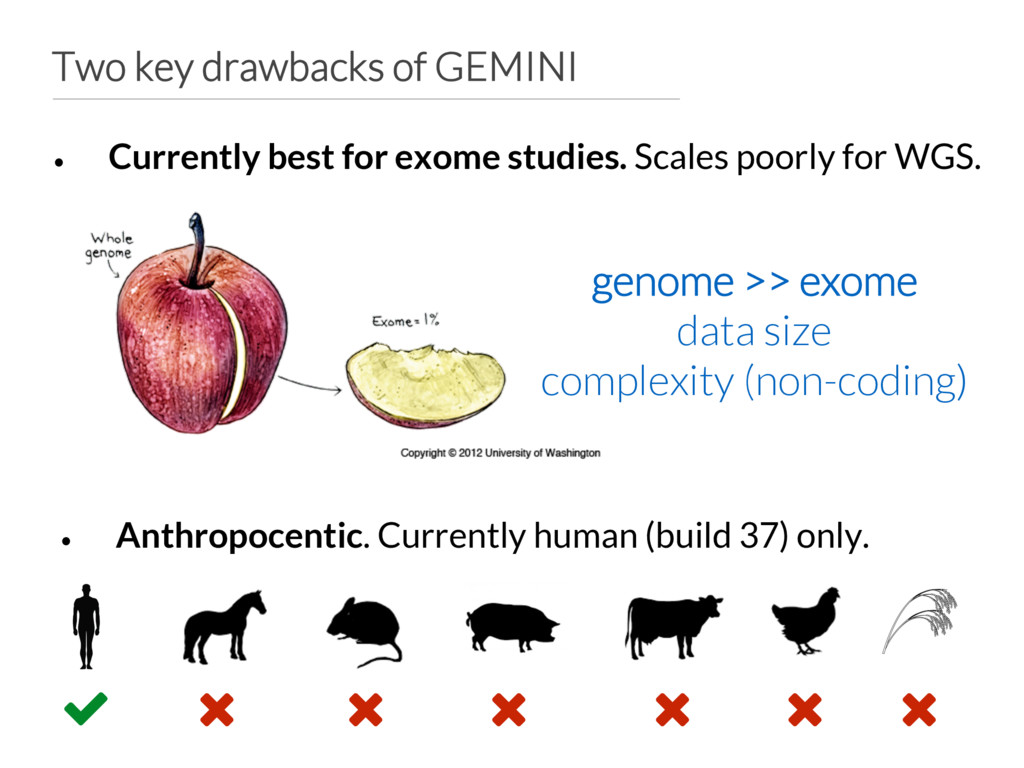
Two key drawbacks of GEMINI • Currently best for exome
studies. Scales poorly for WGS. genome >> exome data size complexity (non-coding) • Anthropocentic. Currently human (build 37) only. ! " " " " " " -
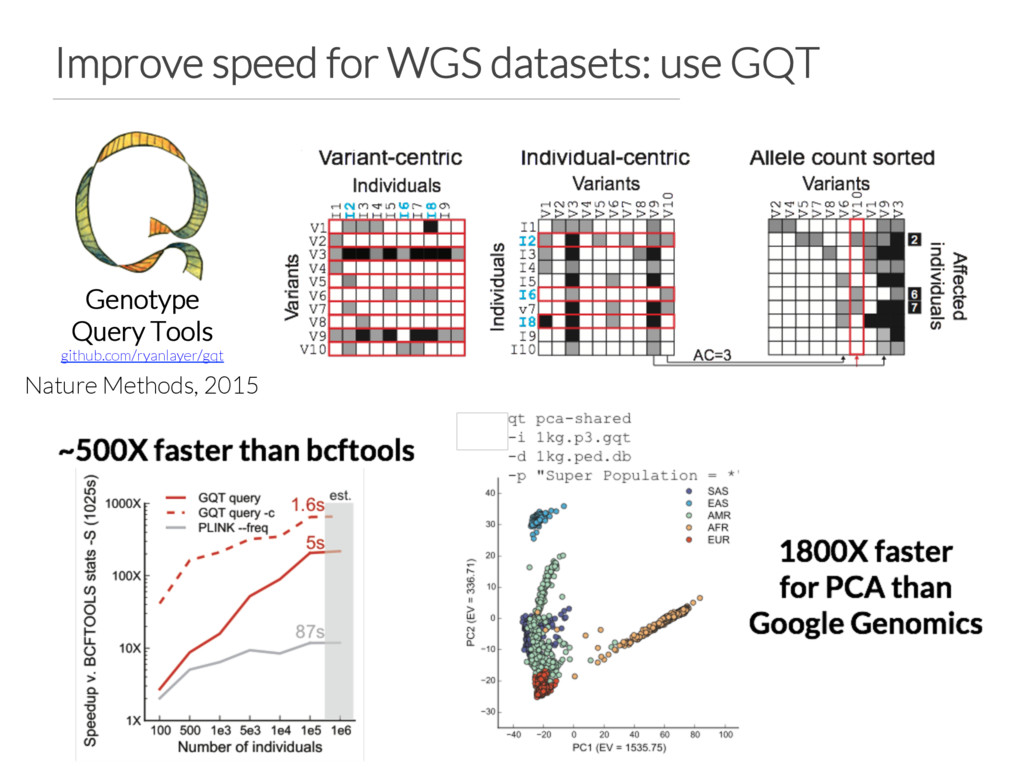
Improve speed for WGS datasets: use GQT Genotype Query Tools
github.com/ryanlayer/gqt Nature Methods, 2015 -
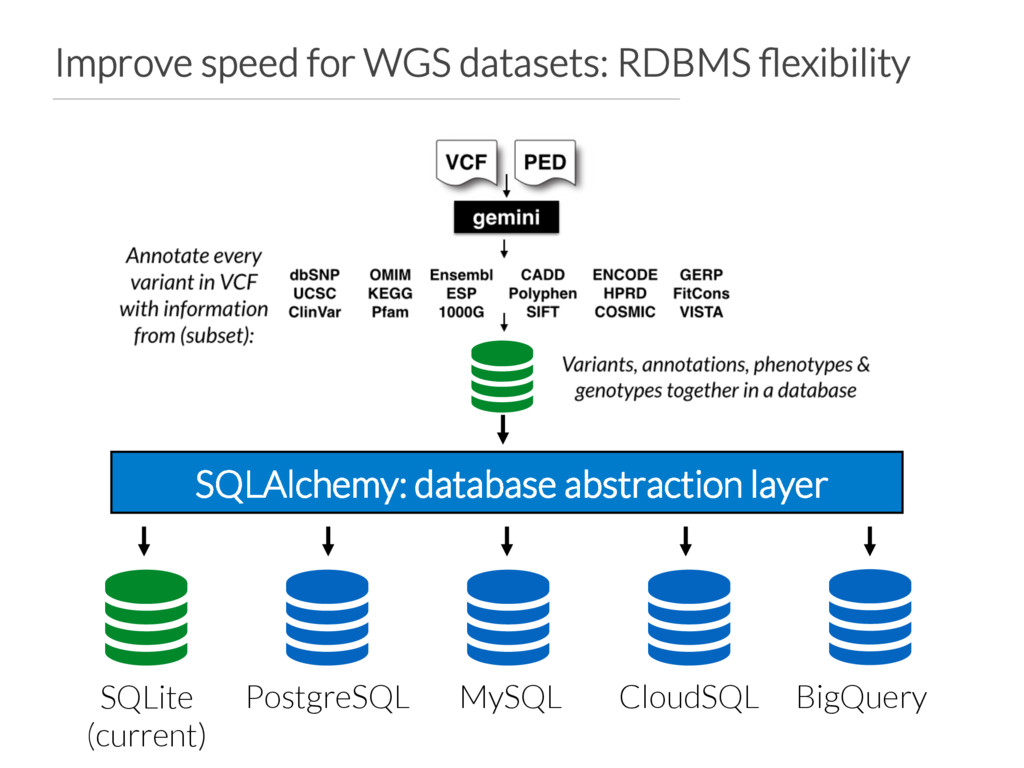
Improve speed for WGS datasets: RDBMS flexibility
SQLite (current) PostgreSQL MySQL CloudSQL BigQuery SQLAlchemy: database abstraction layer -
Improve variant annotation speed and flexibility #CHROM POS ID REF
ALT QUAL FILTER 2 41647 . A G 4495.41 PASS 2 45895 . A G 463.75 PASS 2 224970 . C T 4241.64 PASS 2 229934 . A G 5037.95 PASS 2 234130 . T G 3958 PASS 2 242732 . T TAAC 3193.19 PASS 2 242800 . T C 3929.77 PASS 2 243504 . C T 6628.06 PASS 2 243567 . T TA 3398.03 HRunFilter 2 262553 . T C 3503.49 PASS 2 264895 . G C 3774.13 PASS 2 269352 . G A 9802.28 PASS 2 276942 . A G 5878.58 PASS 2 277250 . G A 7051.35 PASS 2 279705 . C T 7139.54 PASS 2 283231 . A AT 6976.81 HRunFilter 2 675831 . G T 865.05 PASS 2 676177 . C G 4961.19 PASS 2 905368 . C G 101.98 ABFilter; 2 905369 . C G 28.97 ABFilter 2 905393 . C G 930.81 QDFilter 2 905427 . C G 140.17 QDFilter 2 905442 . A T 131.51 ABFilter 2 905492 . T G 550.3 QDFilter 2 905494 . C G 48.5 ABFilter 2 905533 . C T 320.33 ABFilter 2 905576 . T G 72.09 QDFilter 2 905581 . C T 1276.63 QDFilter 2 905595 . G C 390.15 ABFilter 2 905634 . A C 393.91 QDFilter 2 905687 . C G 3233.06 ABFilter 2 905736 . A T 1324.63 QDFilter 2 905763 . G C 15.12 ABFilter Tabix’ed . . . -
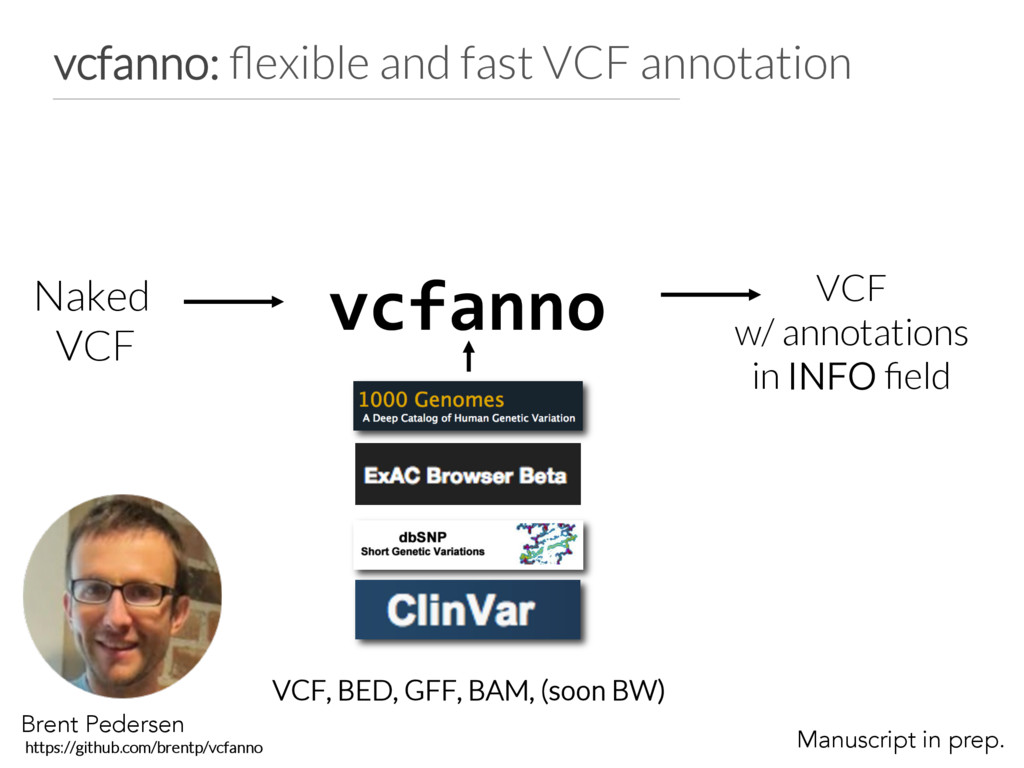
vcfanno: flexible and fast VCF annotation Naked VCF vcfanno VCF
w/ annotations in INFO field Brent Pedersen https://github.com/brentp/vcfanno VCF, BED, GFF, BAM, (soon BW) Manuscript in prep. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”] ops=[“first”, “first”] [[annotation]] file="dbsnp.b141.vcf.gz"
fields=["ID"] names=["rs_ids"] ops=[“concat"] [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "lua:variance(vals)"] vcfanno configuration file. Allows multiple annotations from each file Can rename the annotations in the resulting VCF Multiple operations to summarize the results of multiple hits in annot. file: mean, max, min concat, count, uniq first, flag Match on POS+REF+ALT for VCF annotations. Lua for custom computations. variance() defined in custom.js -
before and after vcfanno AC=11;AF=0.017 AC=11;AF=0.017; exac_aaf=0.0012; exac_num_het=8; rs_ids=1234; gerp_mean=7.25e-07
gerp_var=1.39e-08 Naked VCF Dressed VCF [[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”] ops=[“first”, “first”] [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file -
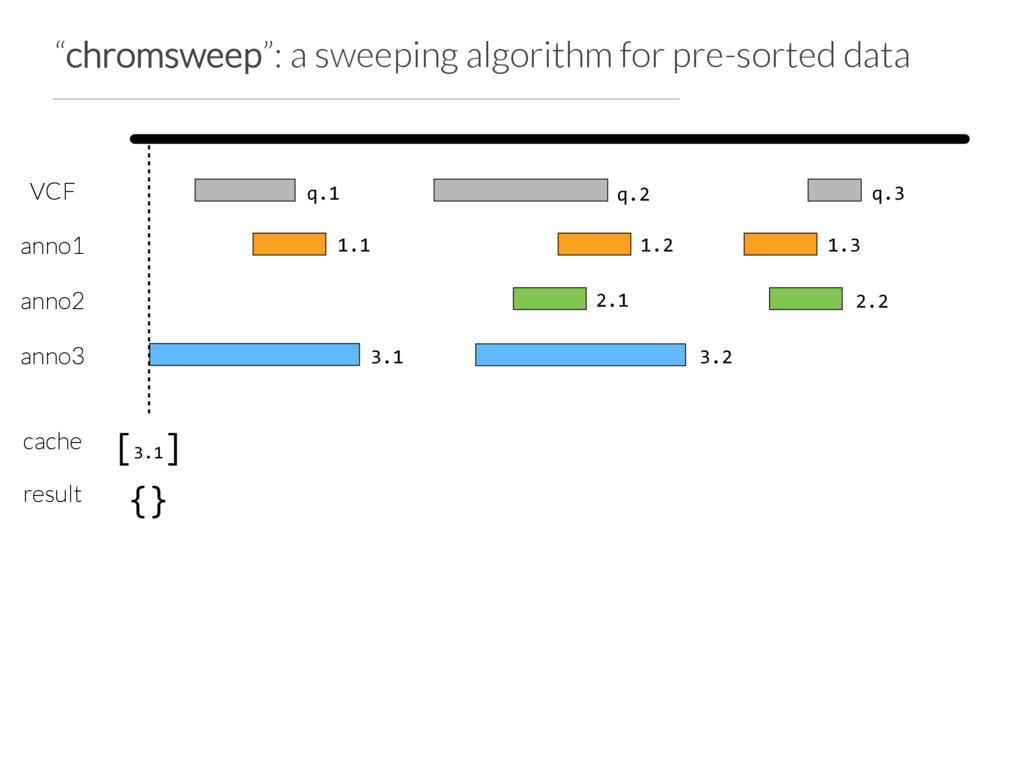
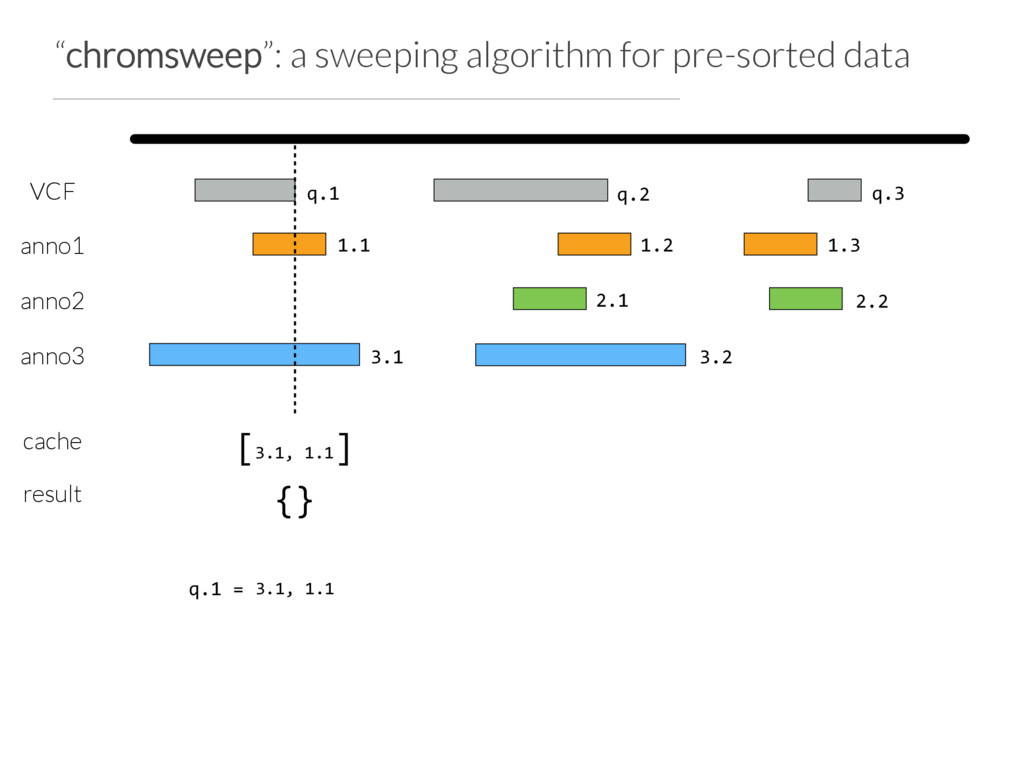
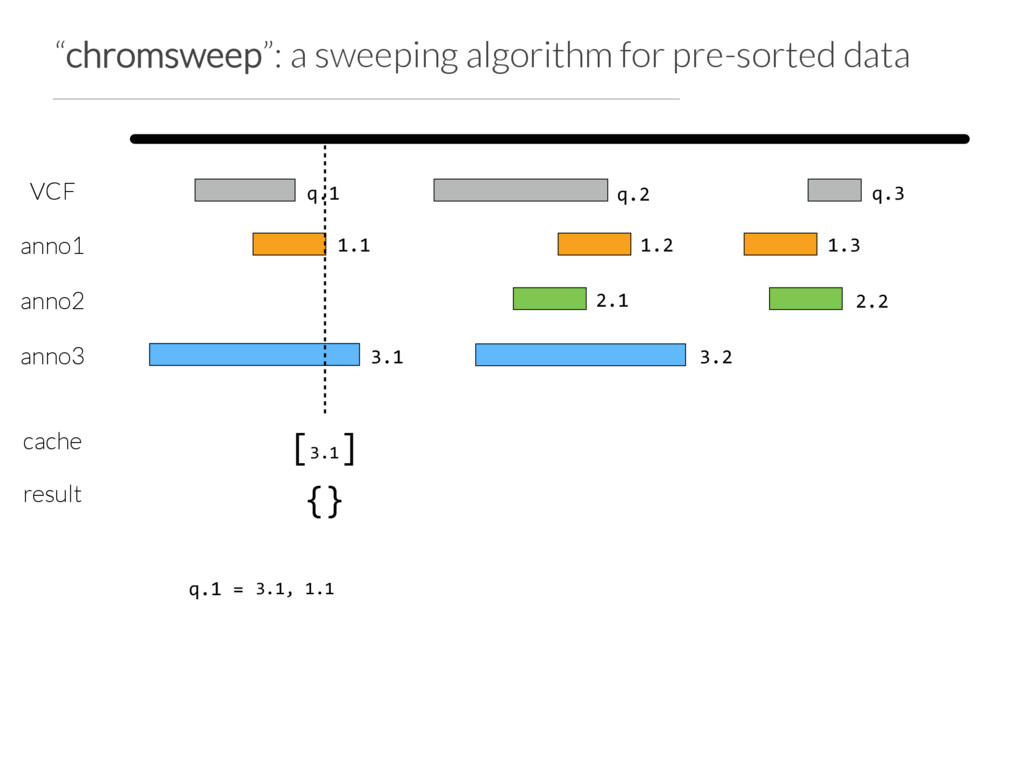
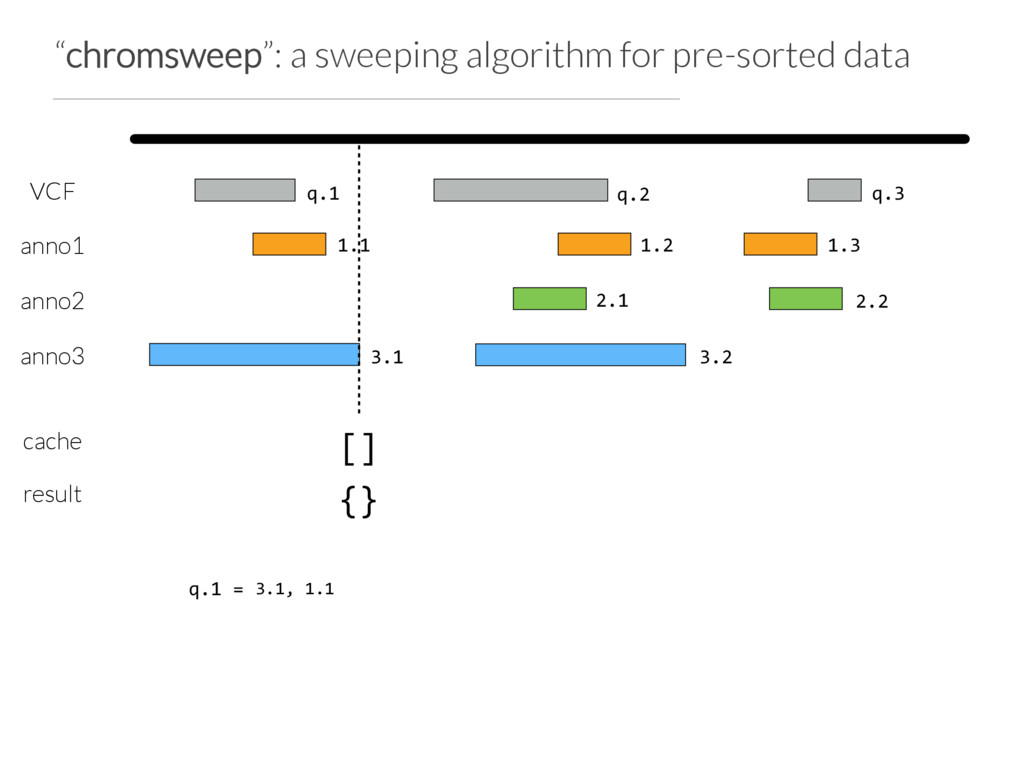
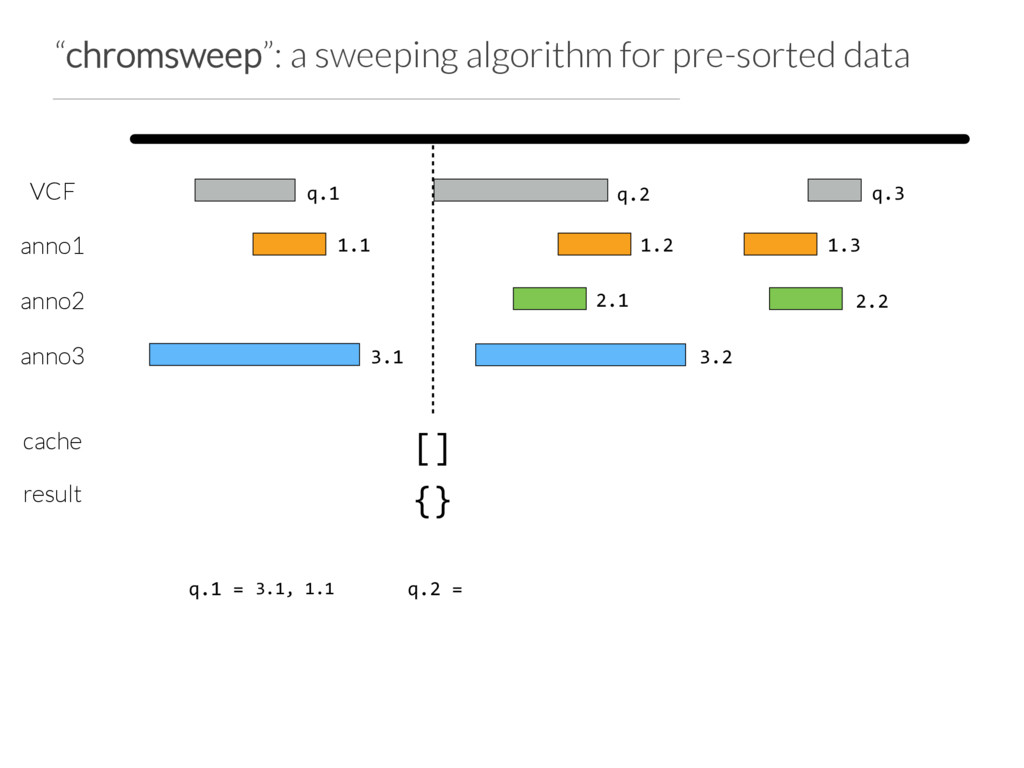
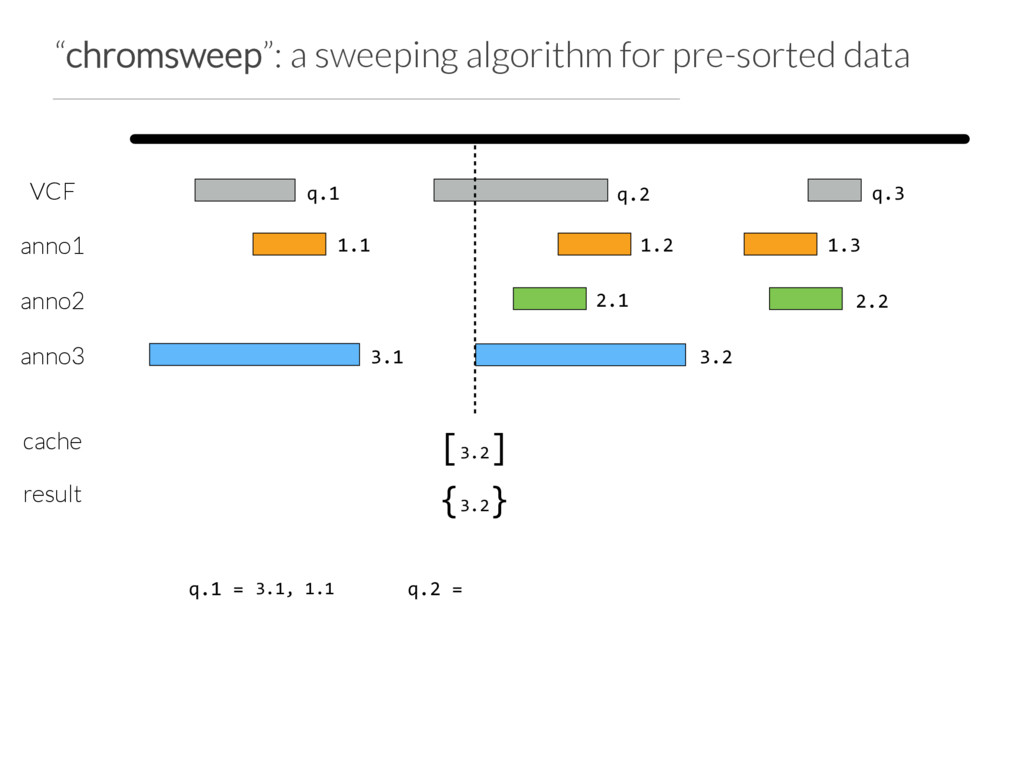
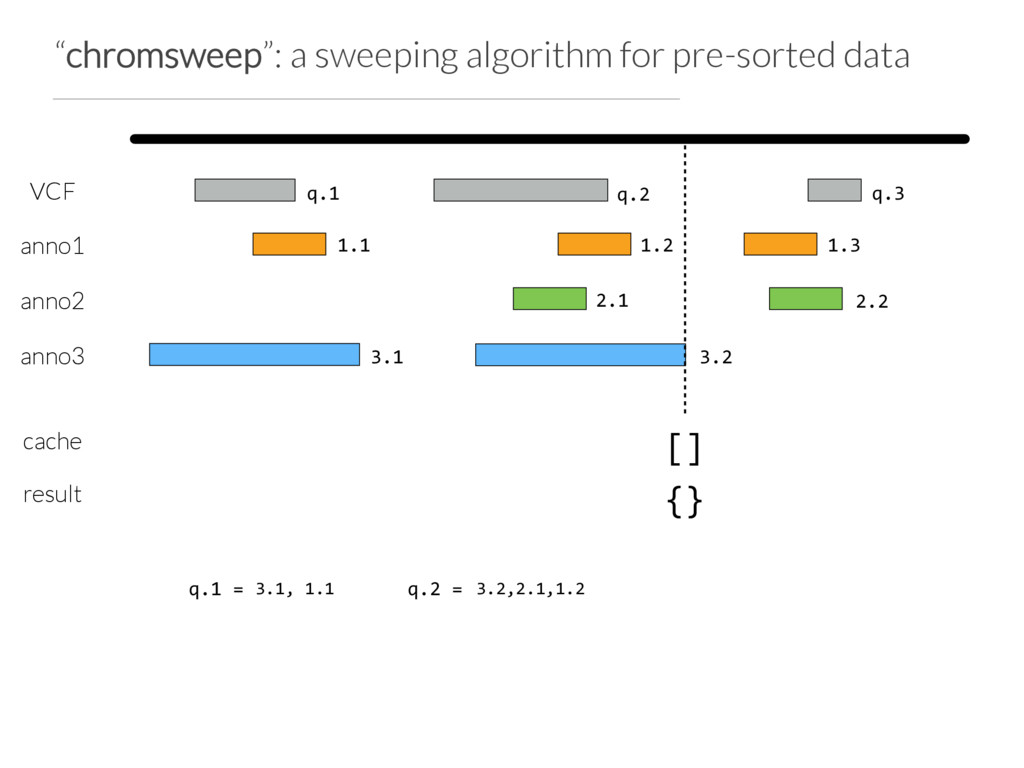
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [] cache result q.1 q.2 q.3 chromsweep is the fundamental algorithm underlying our bedtools software -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [3.1 ] cache result q.1 q.2 q.3 -
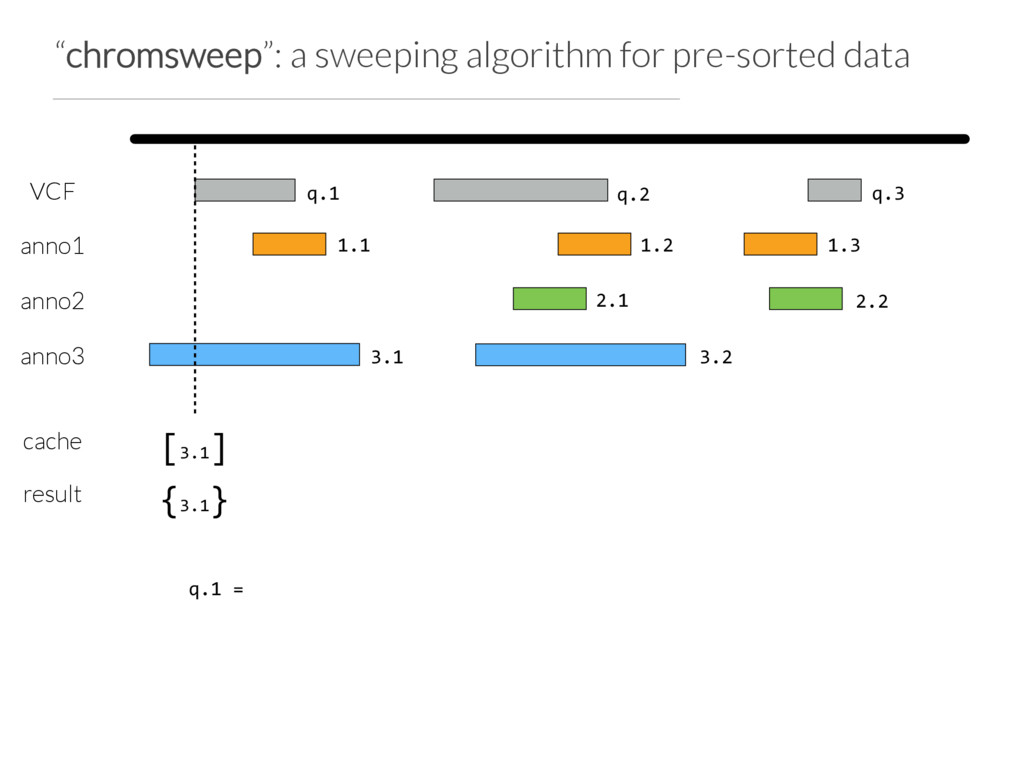
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.1 } [3.1 ] cache result q.1 q.2 q.3 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.1, 1.1 } [3.1, 1.1 ] cache result q.1 q.2 q.3 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [3.1, 1.1 ] cache result q.1 q.2 q.3 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [3.1 ] cache result q.1 q.2 q.3 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [] cache result q.1 q.2 q.3 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [] cache result q.1 q.2 q.3 q.2 = 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.2 } [3.2 ] cache result q.1 q.2 q.3 q.2 = 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
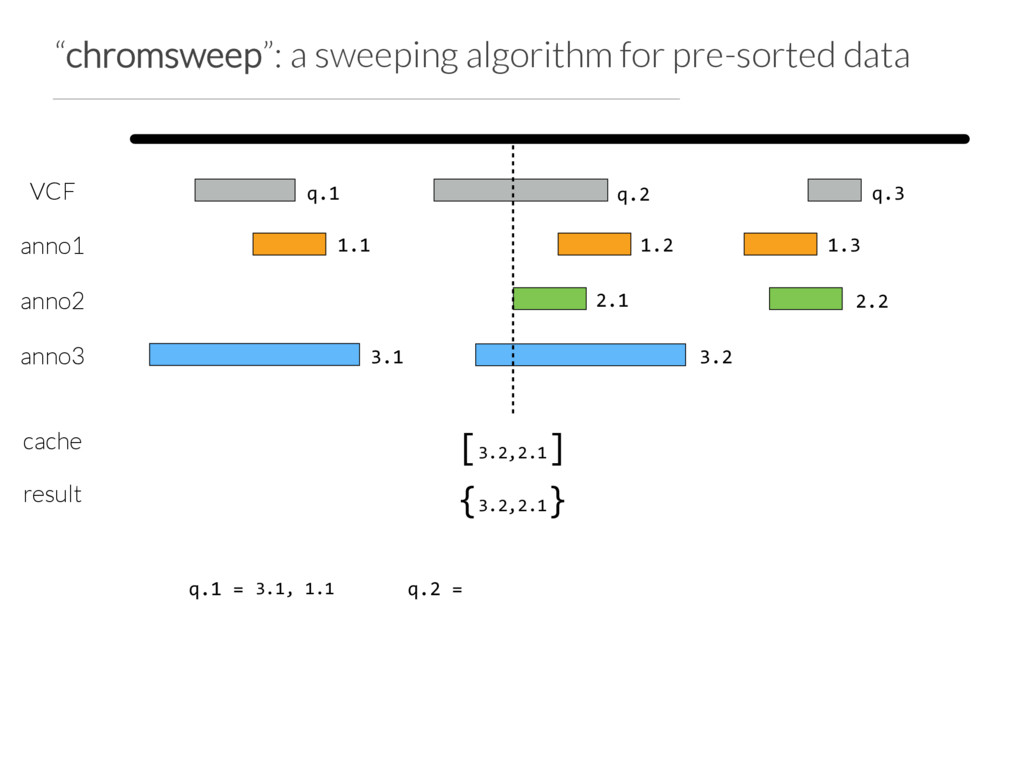
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.2,2.1 } [3.2,2.1 ] cache result q.1 q.2 q.3 q.2 = 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.2,2.1,1.2 } [3.2,2.1,1.2 ] cache result q.1 q.2 q.3 q.2 = 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {3.2,2.1,1.2 } [3.2,2.1,1.2 ] cache result q.1 q.2 q.3 q.2 = *2.1 stays in the cache 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
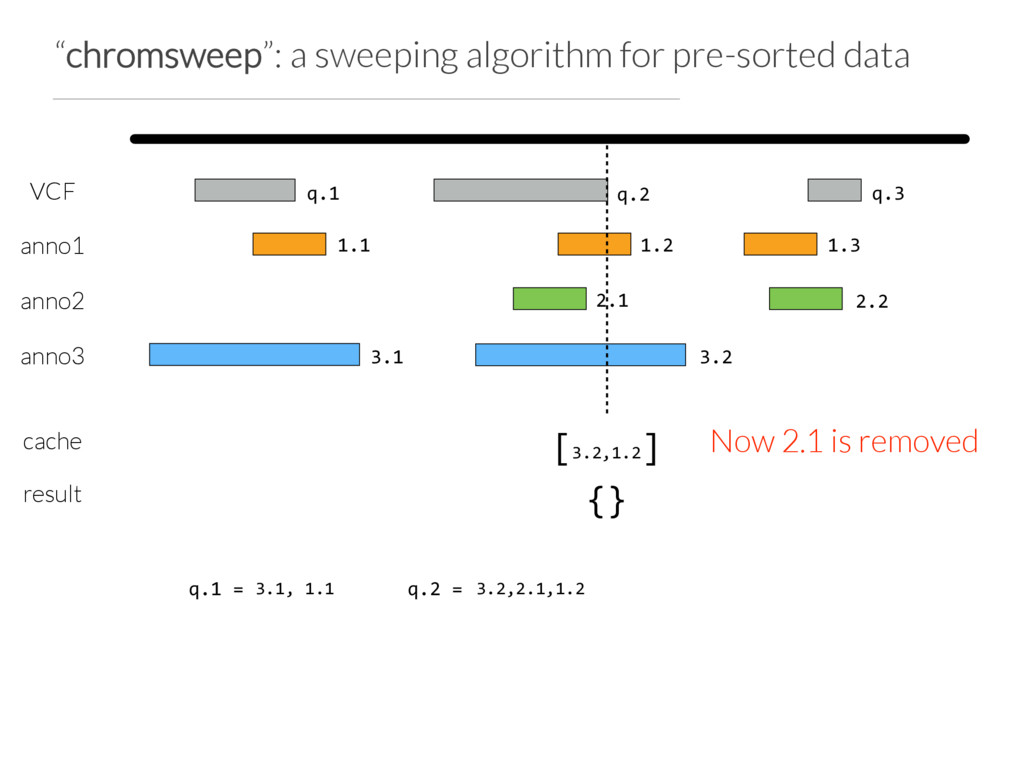
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [3.2,1.2 ] cache result q.1 q.2 q.3 q.2 = Now 2.1 is removed 3.2,2.1,1.2 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
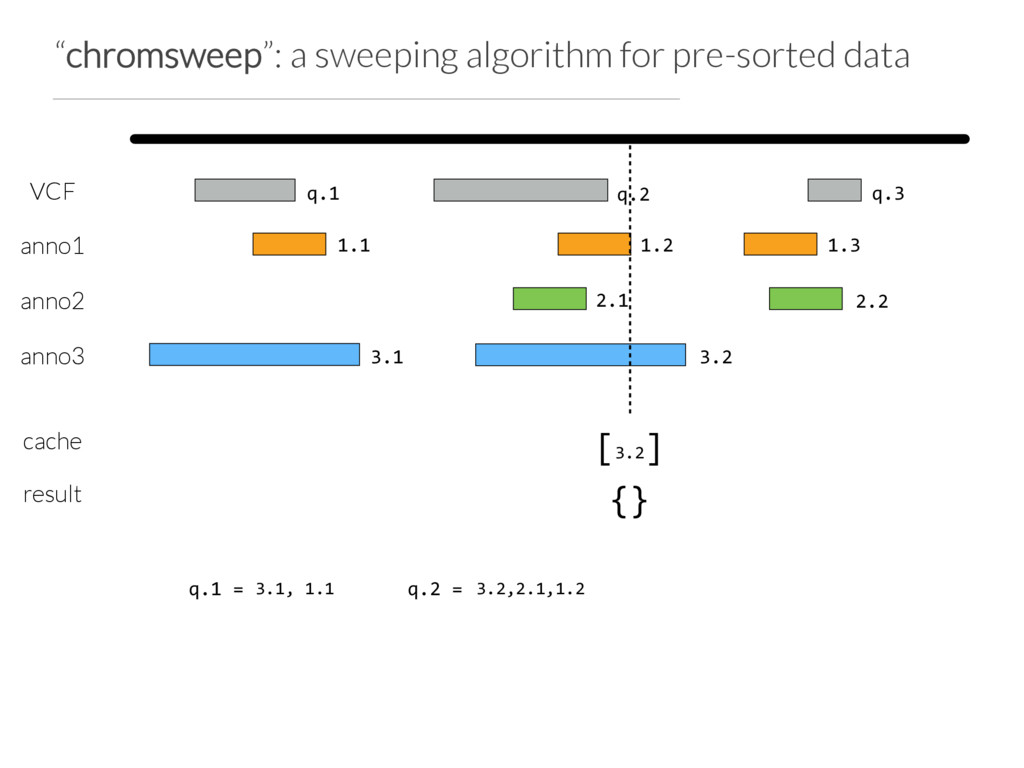
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [3.2 ] cache result q.1 q.2 q.3 q.2 = 3.2,2.1,1.2 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
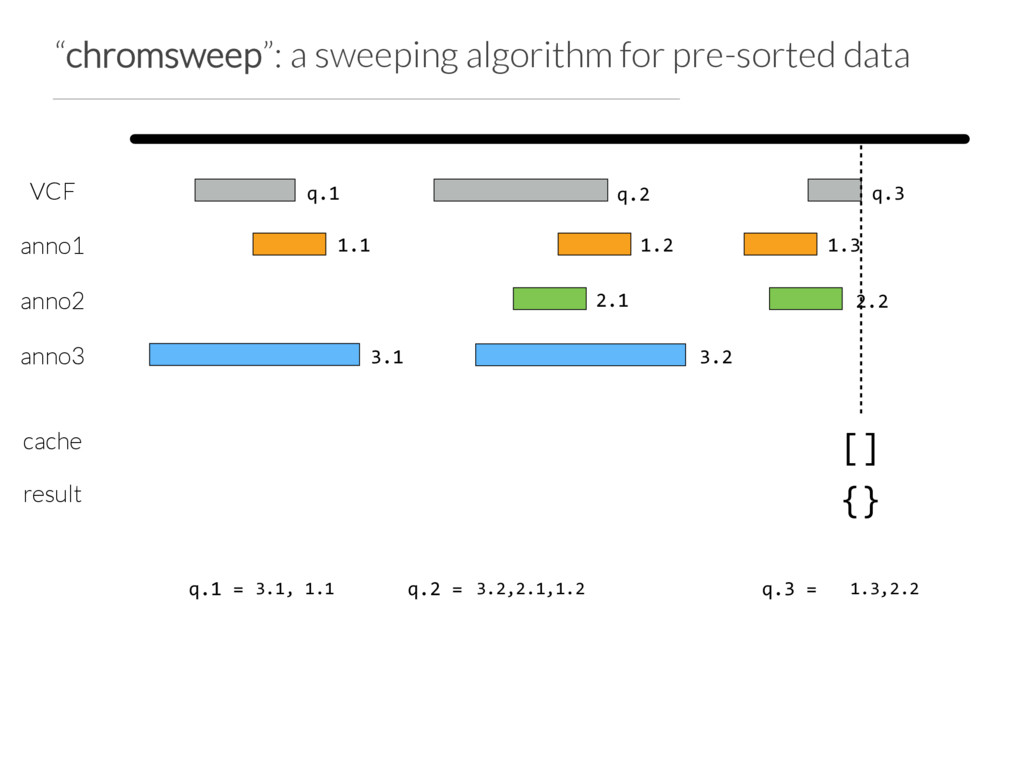
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [] cache result q.1 q.2 q.3 q.2 = 3.2,2.1,1.2 3.1, 1.1 q.1 = -
“chromsweep”: a sweeping algorithm for pre-sorted data VCF anno1 anno2
anno3 1.1 1.2 1.3 2.1 2.2 3.1 3.2 {} [] cache result q.1 q.2 q.3 q.3 = 1.3,2.2 q.2 = 3.2,2.1,1.2 3.1, 1.1 q.1 = -
vcfanno implements the first parallel chromsweep VCF anno1 anno2 anno3
Step 1: partition the query set at “breaks” in the data or when N (e.g. 10) intervals are found Step 2: Use Tabix to extract the records germane to a chunk from each annotation file Step 3: Chromsweep each chunk independently. -
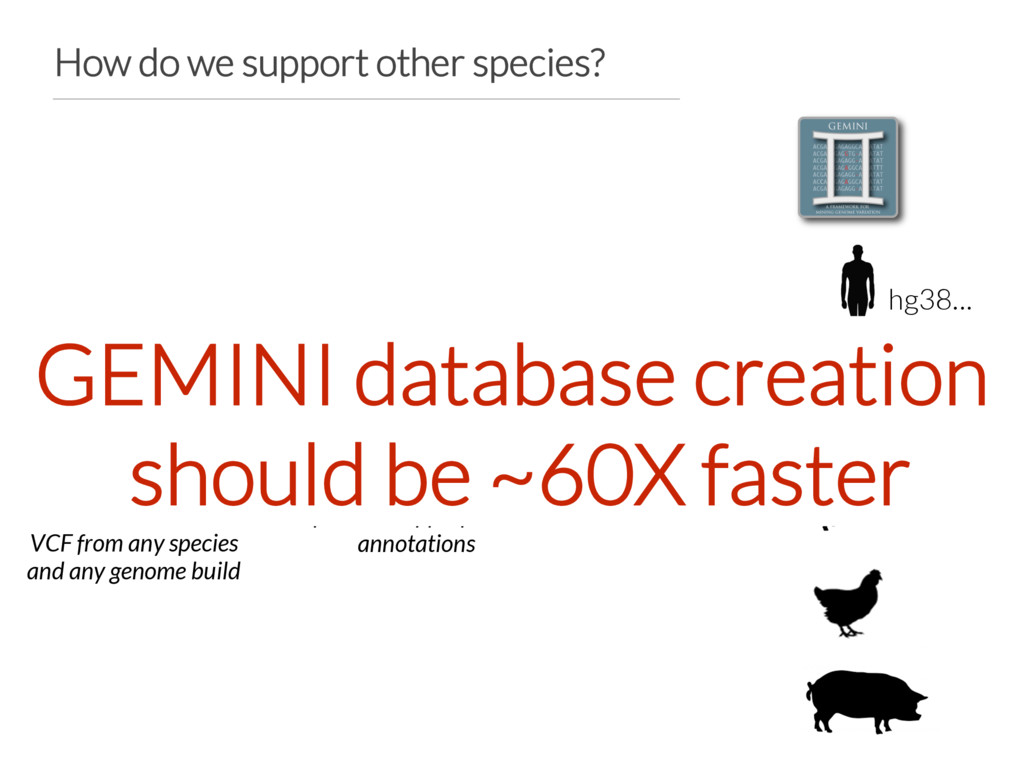
vcfanno VCF hg38… VCF from any species and any genome
build Vcfanno configuration file points to appropriate annotations GEMINI database is created based on vcfanno configuration file GEMINI database creation should be ~60X faster How do we support other species? -
[[annotation]] file=“cpg.hg38.bed.gz" fields=[4] names=[“cpg_density"] ops=[“mean"] [[annotation]] file=“rmsk.hg38.bed.gz" fields=[4] names=[“rmsk"] ops=[“concat”]
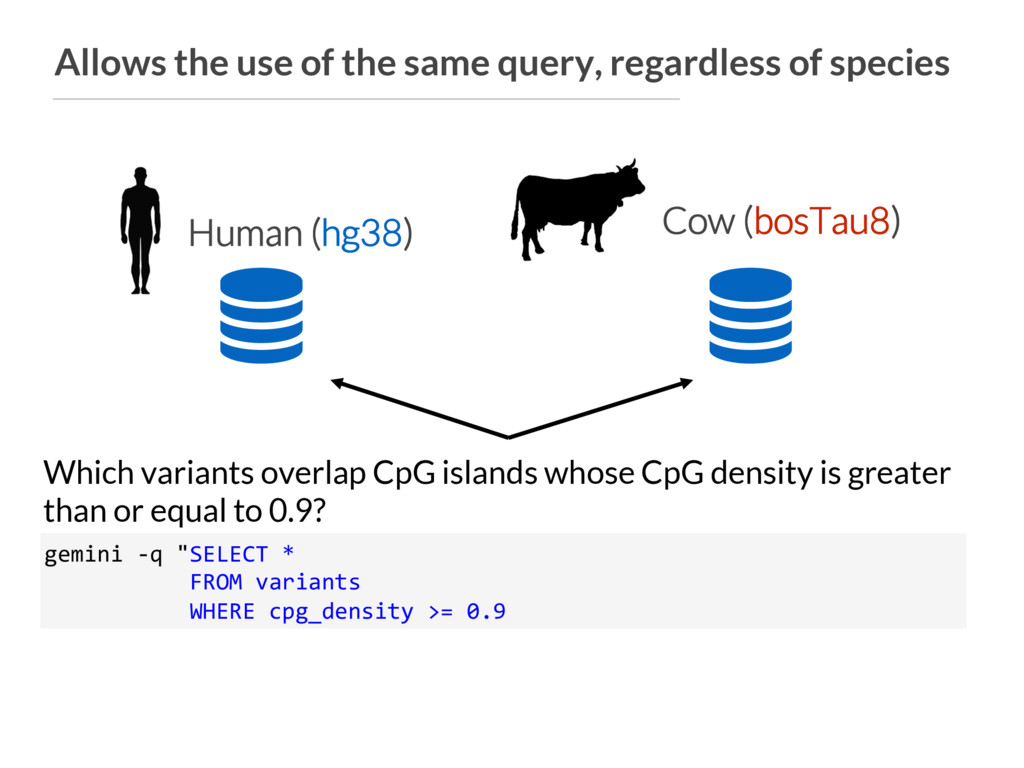
[[annotation]] file="cytoband.hg38.bed.gz" fields=[4] names=[“cytoband”] ops=[“distinct"] How? Simply point vcfanno to the relevant annotations Human (hg38) [[annotation]] file=“cpg.bosTau8.bed.gz" fields=[4] names=[“cpg_density"] ops=[“mean"] [[annotation]] file=“rmsk.bosTau8.bed.gz" fields=[4] names=[“rmsk"] ops=[“concat”] [[annotation]] file="cytoband.bosTau8.bed.gz" fields=[4] names=[“cytoband”] ops=[“distinct"] Cow (bosTau8) -
Allows the use of the same query, regardless of species
gemini -q "SELECT * FROM variants WHERE cpg_density >= 0.9 Which variants overlap CpG islands whose CpG density is greater than or equal to 0.9? Human (hg38) Cow (bosTau8) -
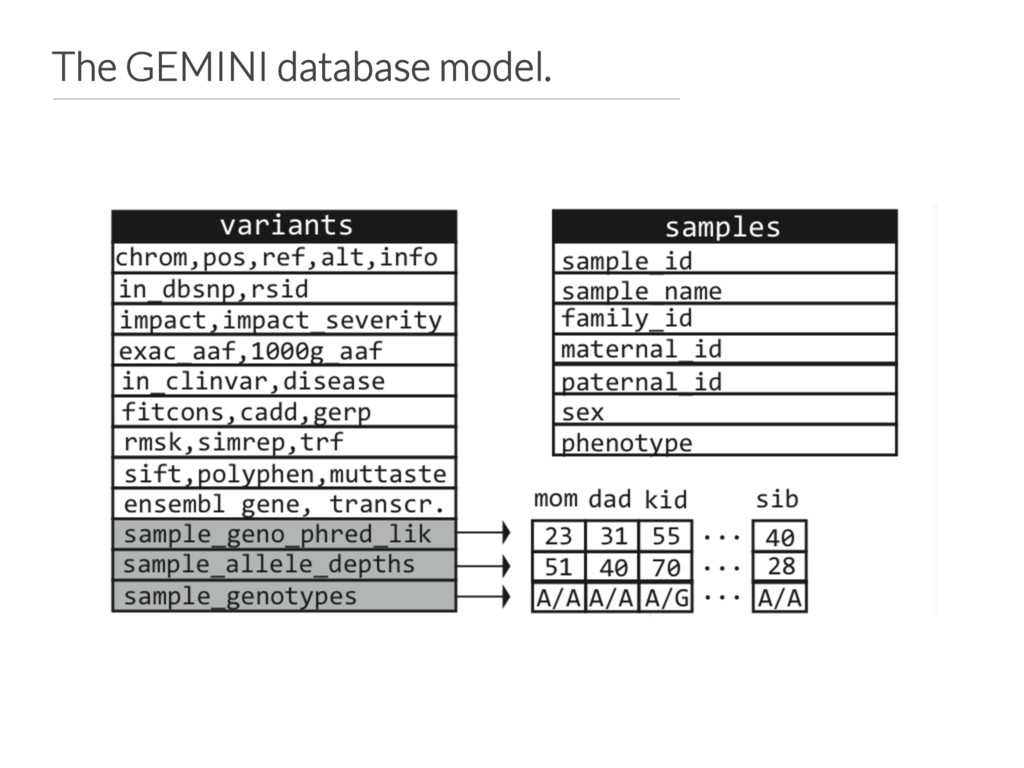
Summary • GEMINI is a flexible framework for exploring genetic
variation from WES and WGS studies. • Integrates variants, genotypes, phenotypes and annotations into a simple database. • Current focus: • Improving scalability for WGS • Support for any (diploid) species • Expected release: April 2016 github.com/arq5x/gemini gemini.rtfd.org -
Challenges • Multi-allelic variants are a bugger. • Even harder
with polyploidy. The VCF format is ill-suited to this. • Versioning & distributing annos. See GGD: https://github.com/arq5x/ggd -
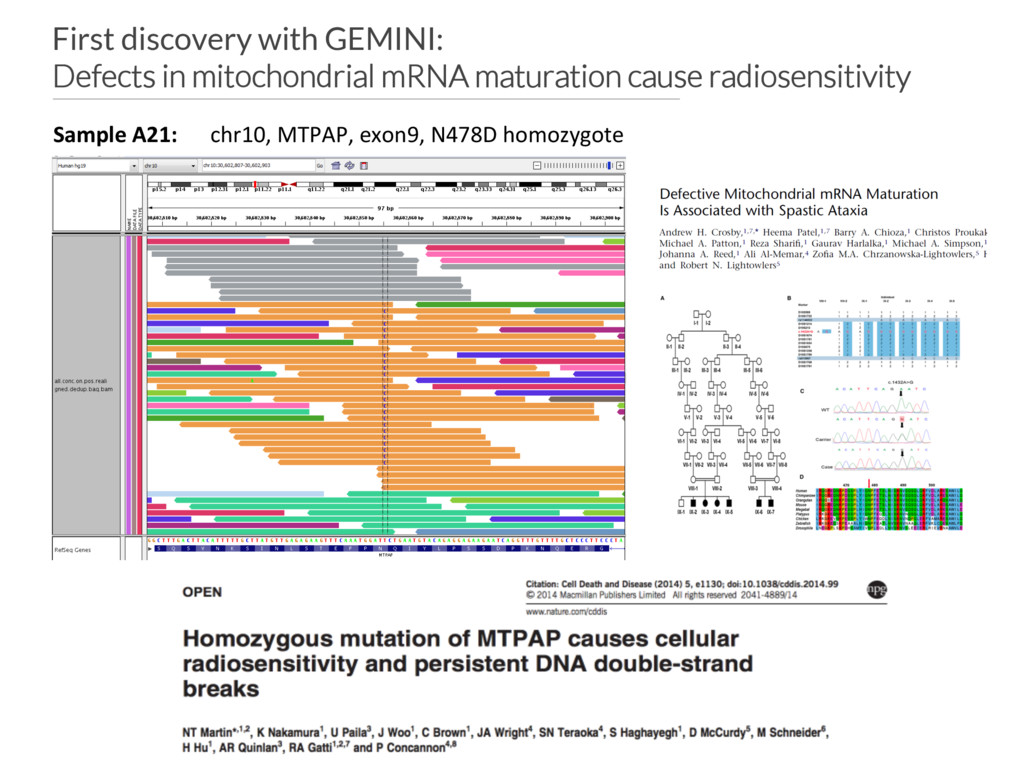
First discovery with GEMINI: Defects in mitochondrial mRNA maturation cause
radiosensitivity Sample A21: chr10, MTPAP, exon9, N478D homozygote