Share
Embed
Transcript
32 slides
-
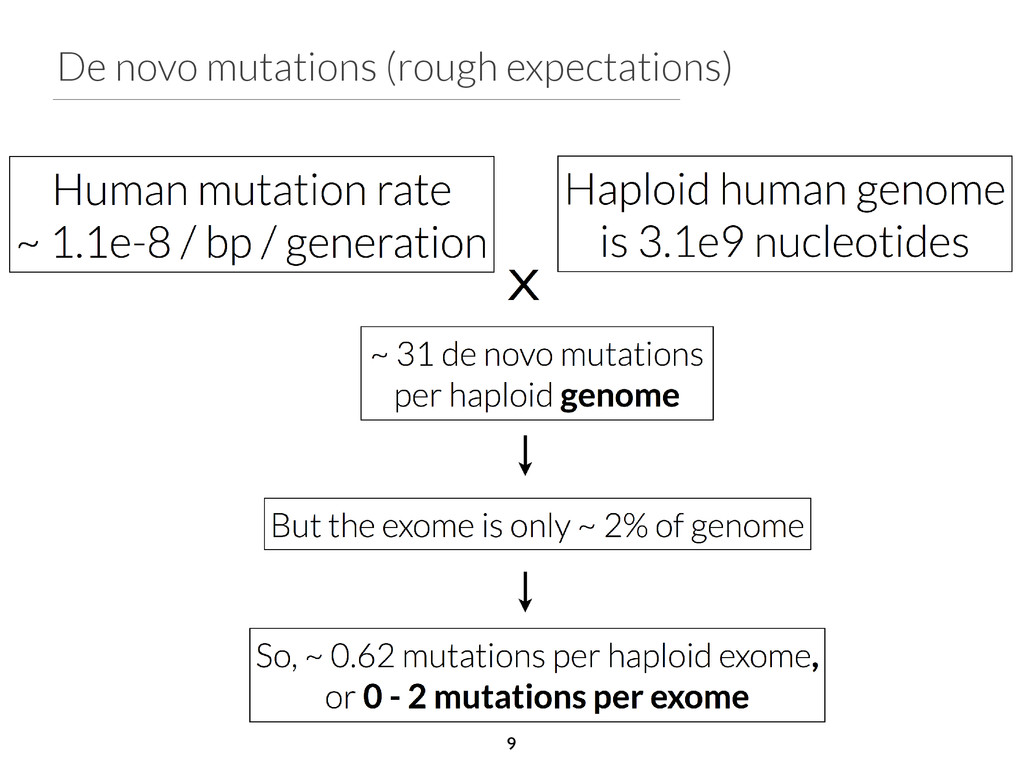
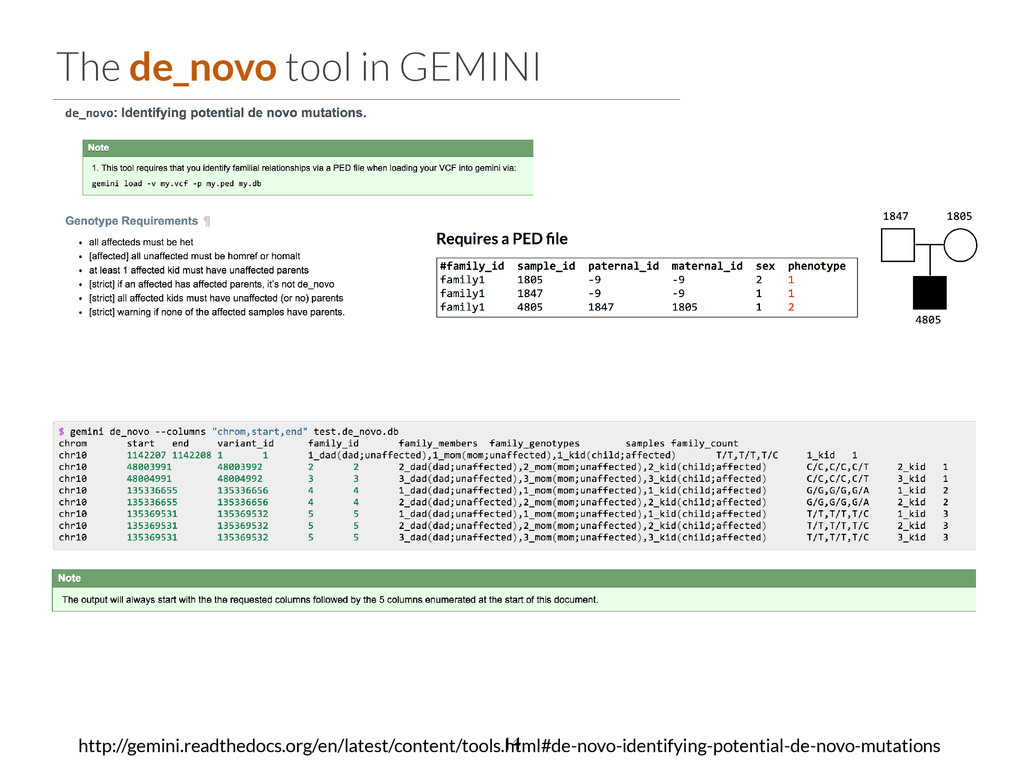
Identifying de novo mutations with GEMINI Please refer to the
following Github Gist to find each command for this session. Commands should be copy/pasted from this Gist Aaron Quinlan University of Utah ! ! ! ! ! quinlanlab.org 1 https://gist.github.com/arq5x/9e1928638397ba45da2e#file-denovo-sh -
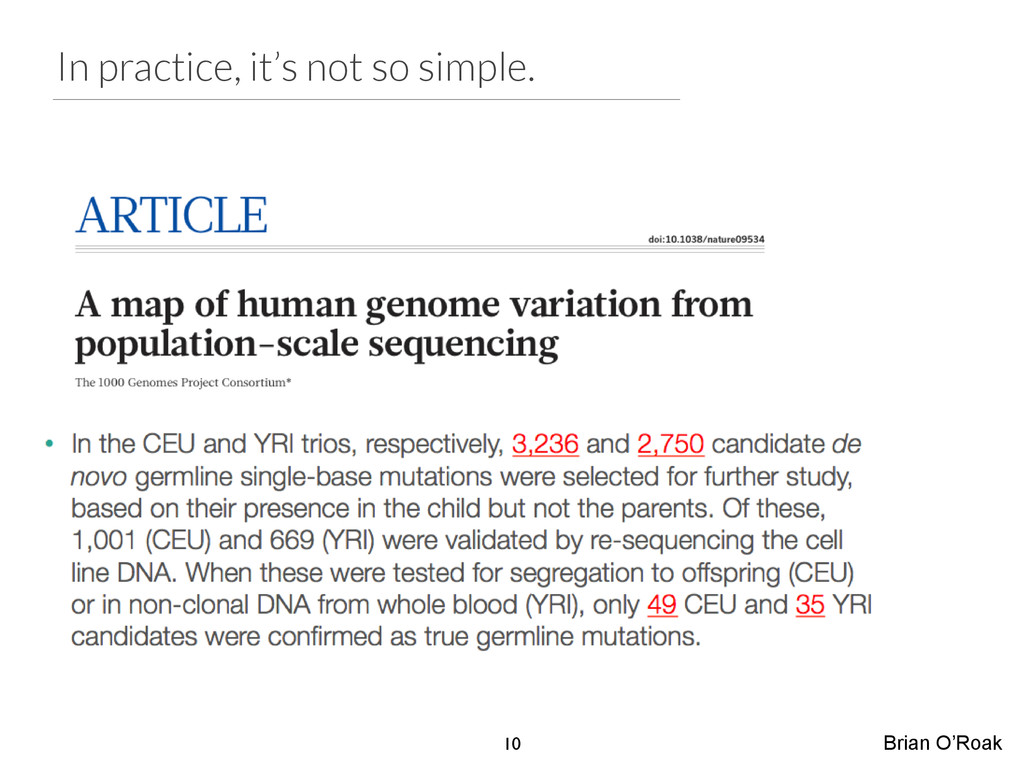
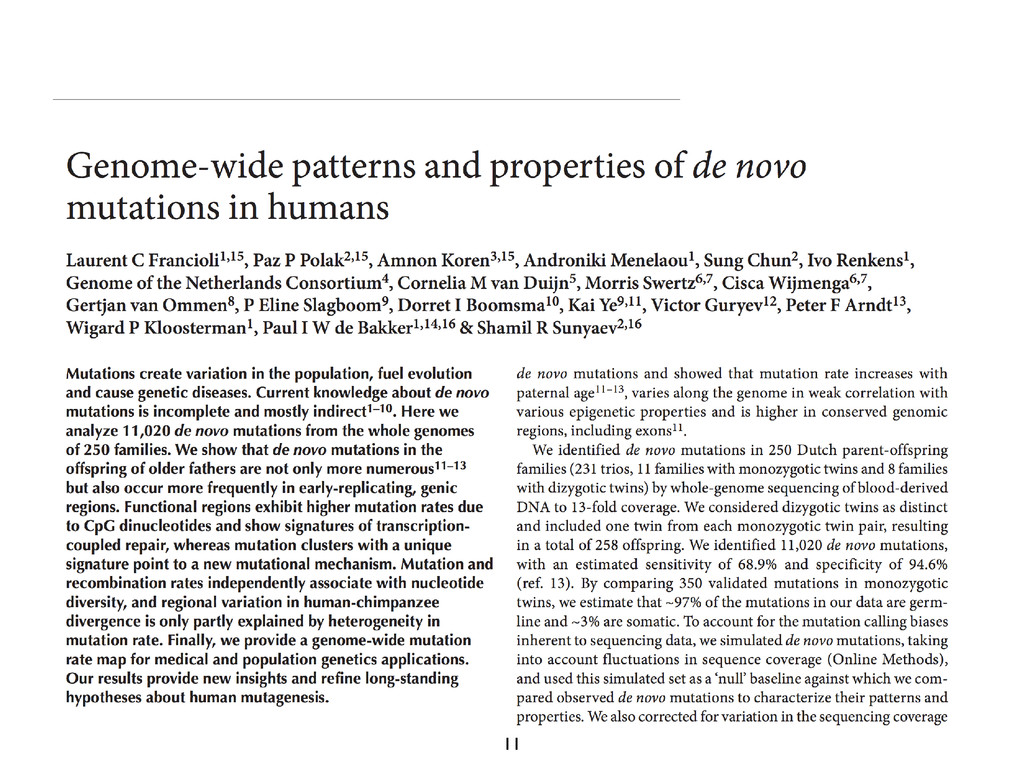
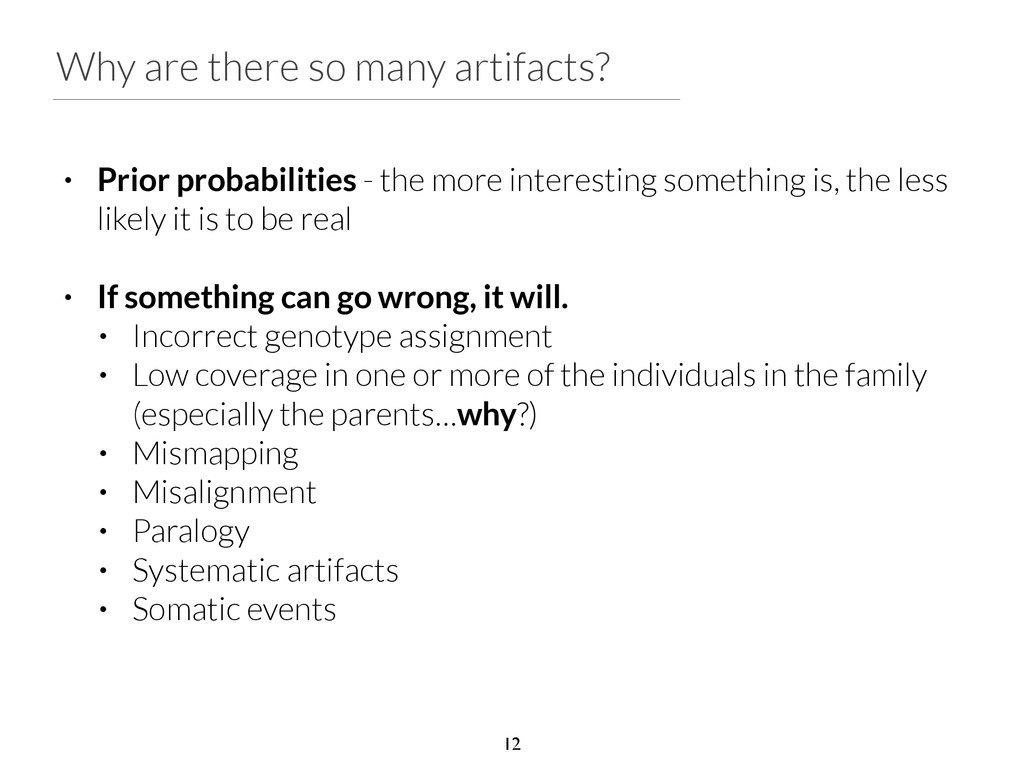
Why are there so many artifacts? • Prior probabilities -
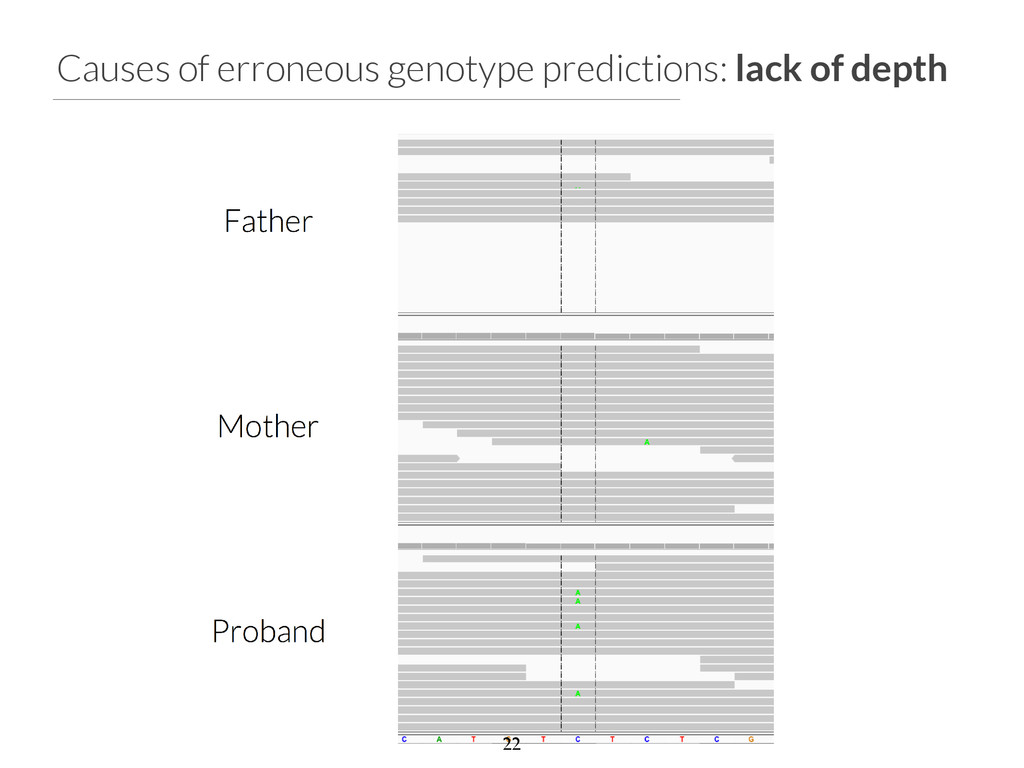
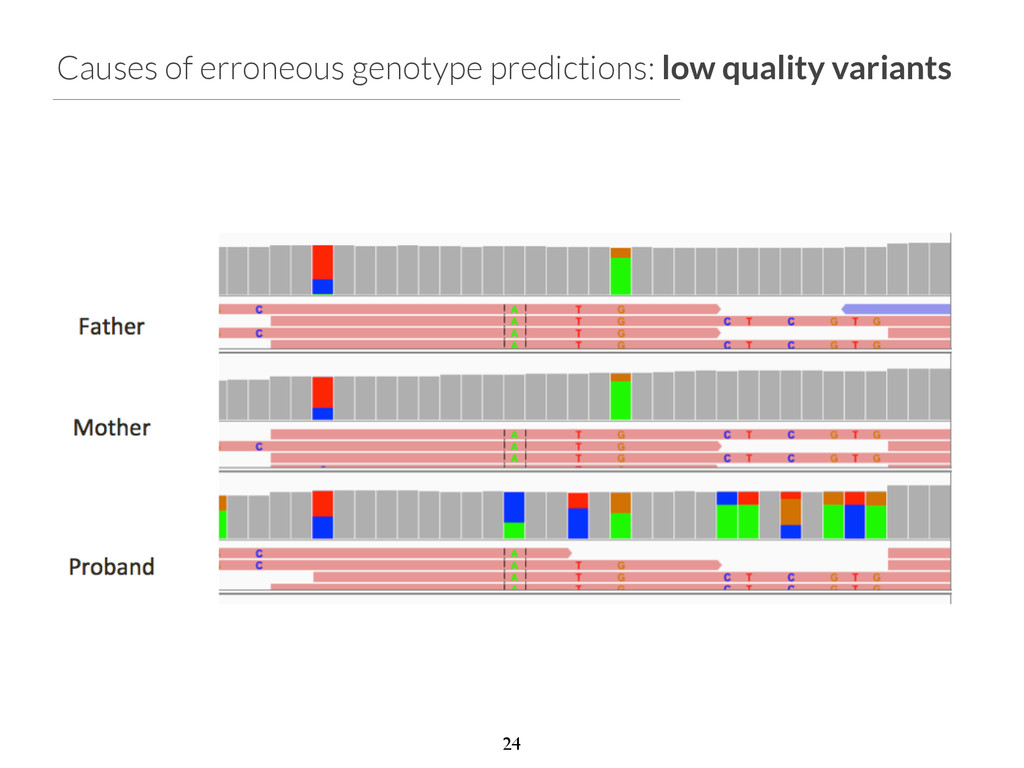
the more interesting something is, the less likely it is to be real ! • If something can go wrong, it will. • Incorrect genotype assignment • Low coverage in one or more of the individuals in the family (especially the parents…why?) • Mismapping • Misalignment • Paralogy • Systematic artifacts • Somatic events 12 -
Create a GEMINI database from a VCF Notes: 1. The
VCF has been normalized and decomposed with VT 2. The VCF has been annotated with VEP. $ curl https://s3.amazonaws.com/gemini-‐tutorials/trio.trim.vep.vcf.gz > trio.trim.vep.vcf.gz $ curl https://s3.amazonaws.com/gemini-‐tutorials/denovo.ped > denovo.ped $ gemini load -‐-‐cores 4 \ -‐v trio.trim.vep.vcf.gz \ -‐t VEP \ -‐-‐skip-‐gene-‐tables -‐-‐skip-‐cadd -‐-‐skip-‐gerp-‐bp \ -‐p de_novo.ped \ ! trio.trim.vep.denovo.db Note: copy and paste the full command from the Github Gist to avoid errors ~8 minutes http://gemini.readthedocs.org/en/latest/content/preprocessing.html#step-1-split-left-align-and-trim-variants 15 -
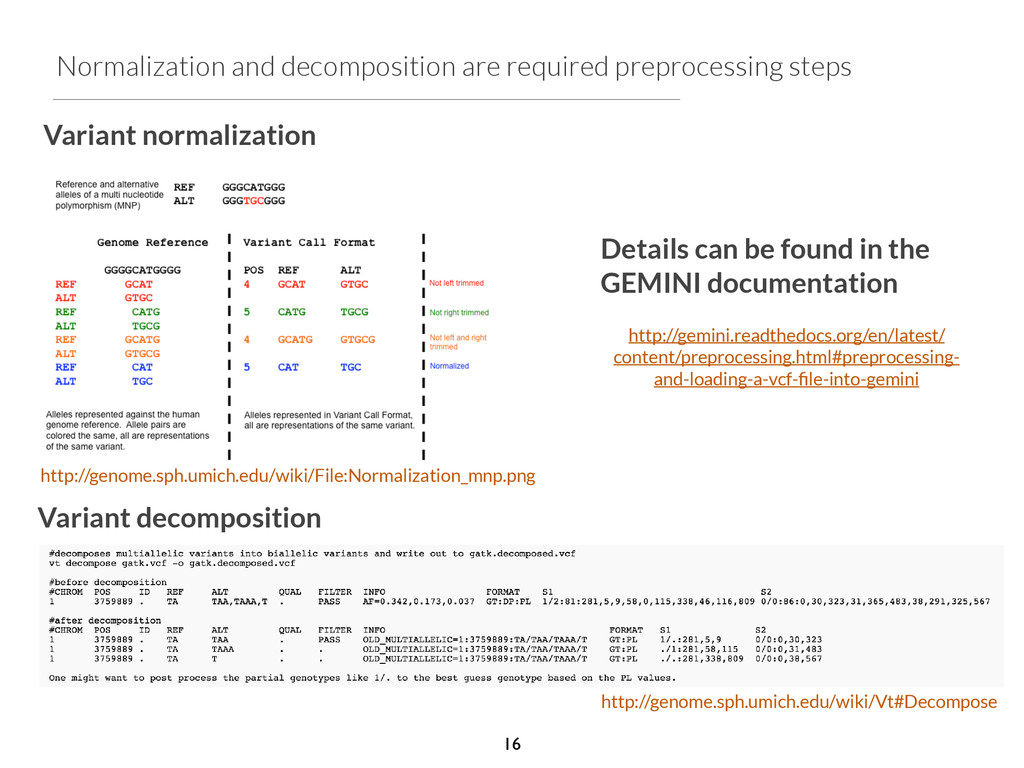
Normalization and decomposition are required preprocessing steps Variant decomposition http://genome.sph.umich.edu/wiki/Vt#Decompose
Variant normalization http://genome.sph.umich.edu/wiki/File:Normalization_mnp.png http://gemini.readthedocs.org/en/latest/ content/preprocessing.html#preprocessing- and-loading-a-vcf-file-into-gemini Details can be found in the GEMINI documentation 16 -
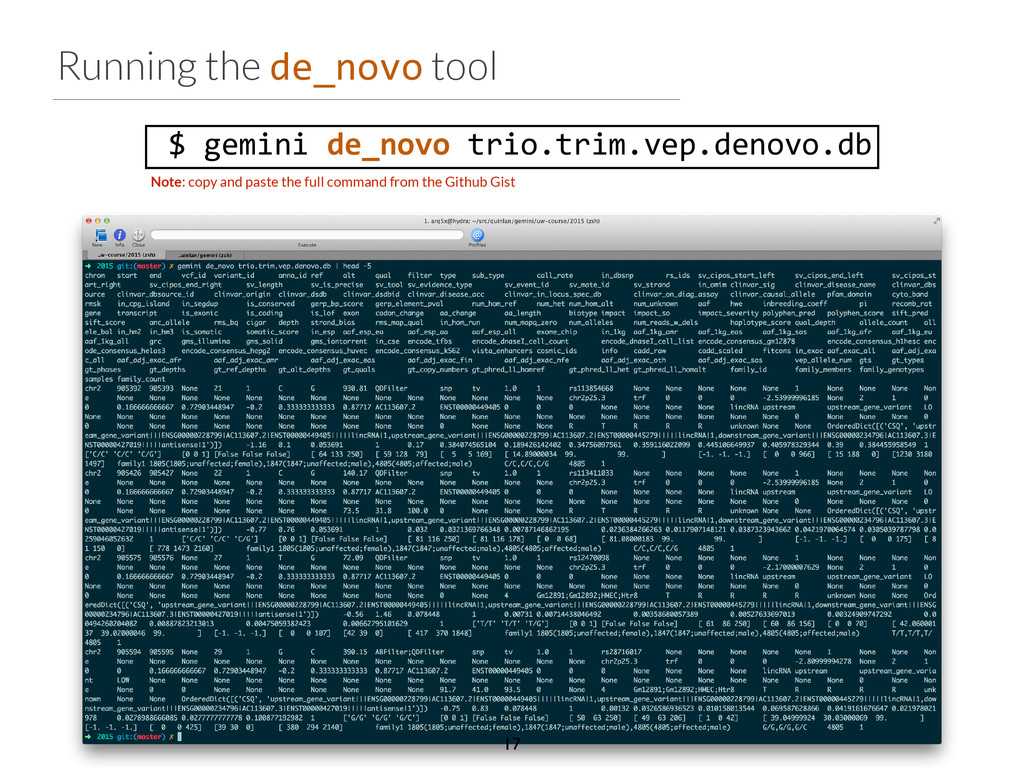
Running the de_novo tool $ gemini de_novo trio.trim.vep.denovo.db Note: copy
and paste the full command from the Github Gist 17 -
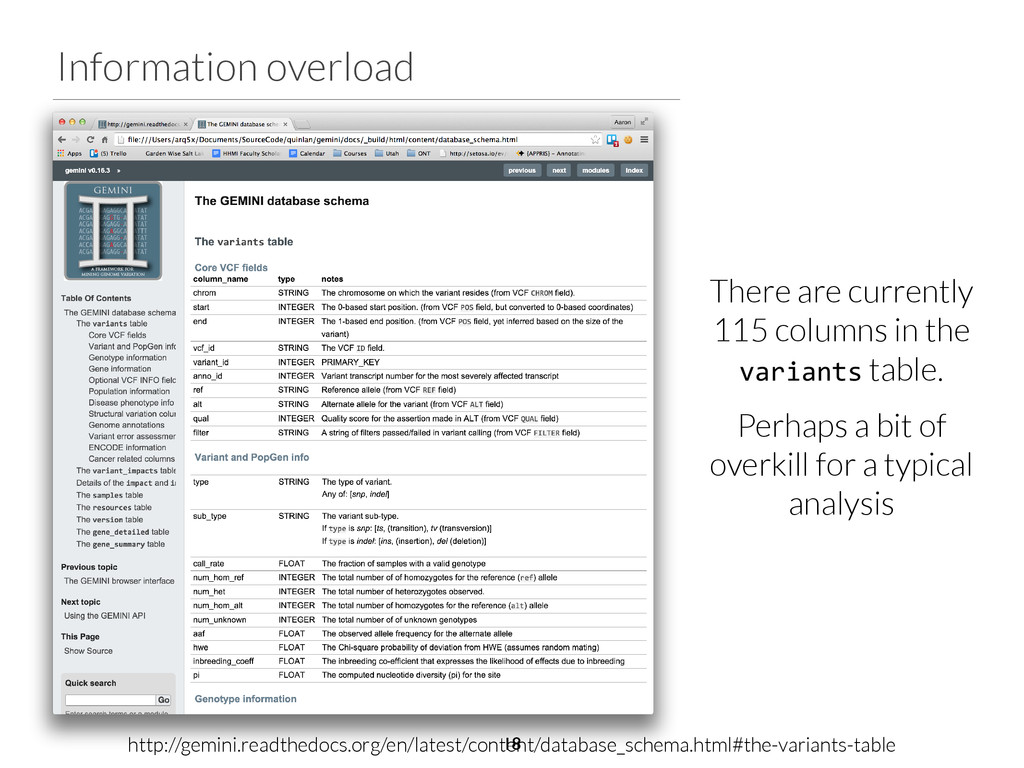
Information overload There are currently 115 columns in the variants
table. Perhaps a bit of overkill for a typical analysis http://gemini.readthedocs.org/en/latest/content/database_schema.html#the-variants-table 18 -
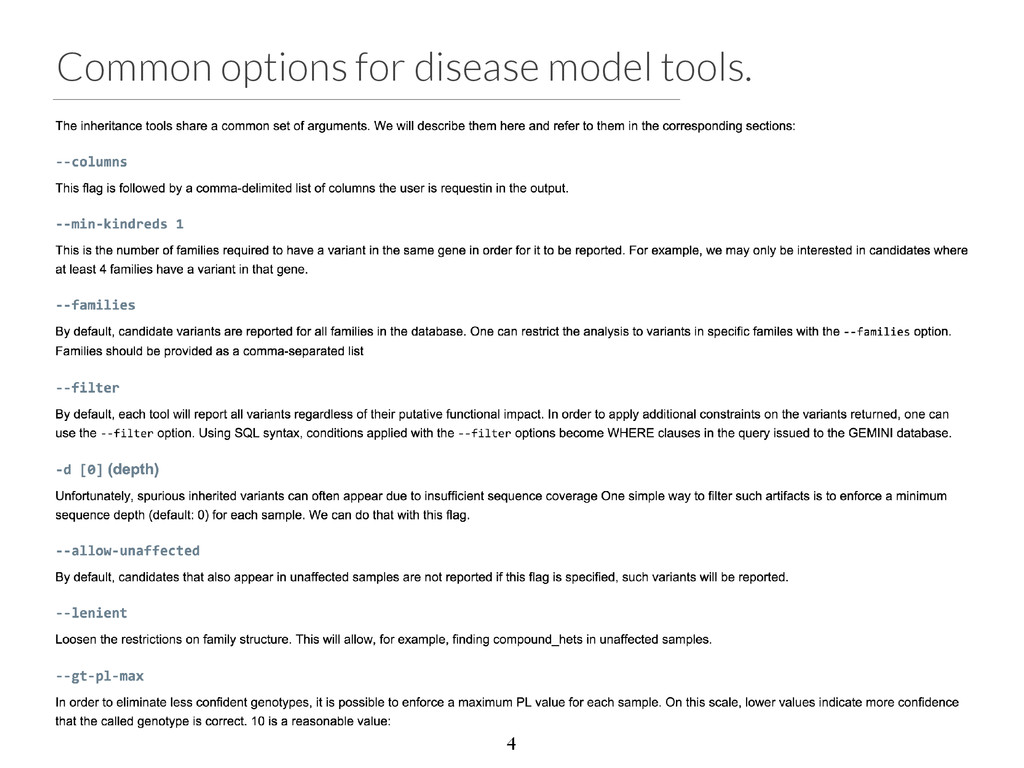
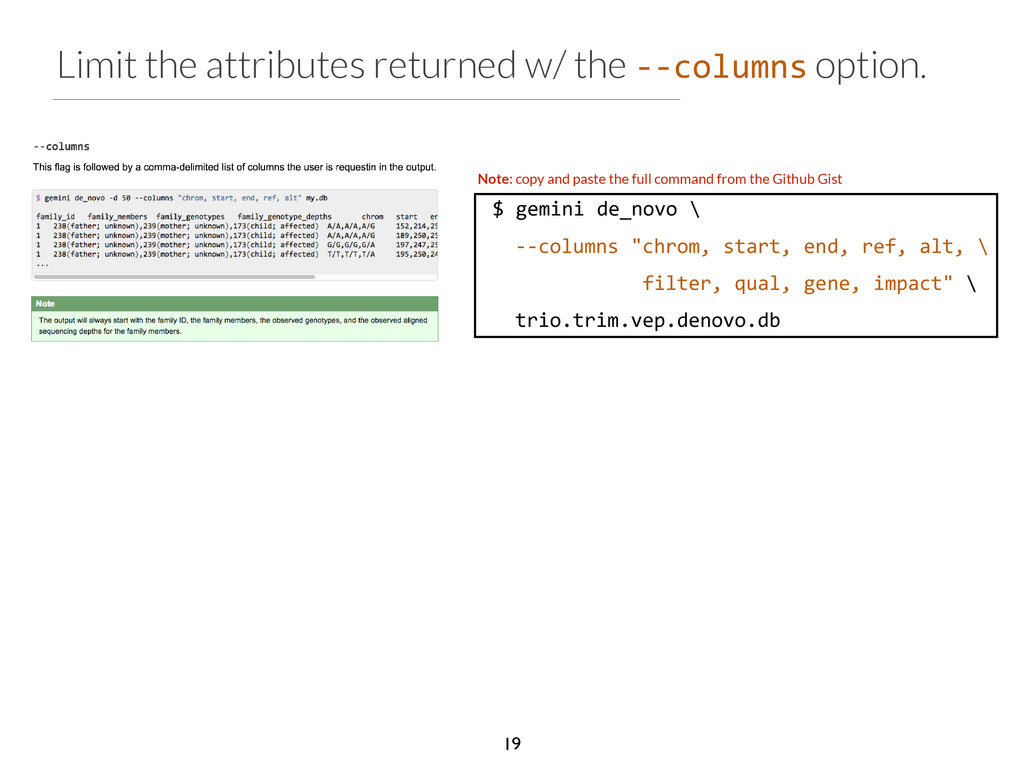
Limit the attributes returned w/ the -‐-‐columns option. $ gemini
de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ trio.trim.vep.denovo.db Note: copy and paste the full command from the Github Gist 19 -
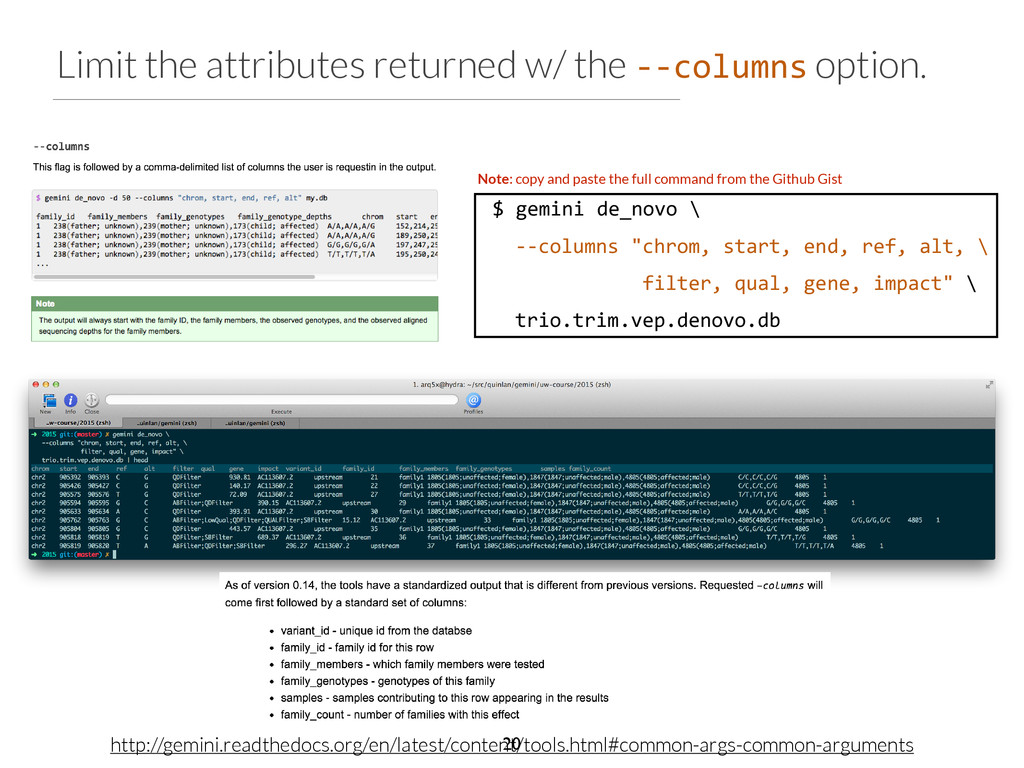
Limit the attributes returned w/ the -‐-‐columns option. http://gemini.readthedocs.org/en/latest/content/tools.html#common-args-common-arguments $
gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ trio.trim.vep.denovo.db Note: copy and paste the full command from the Github Gist 20 -
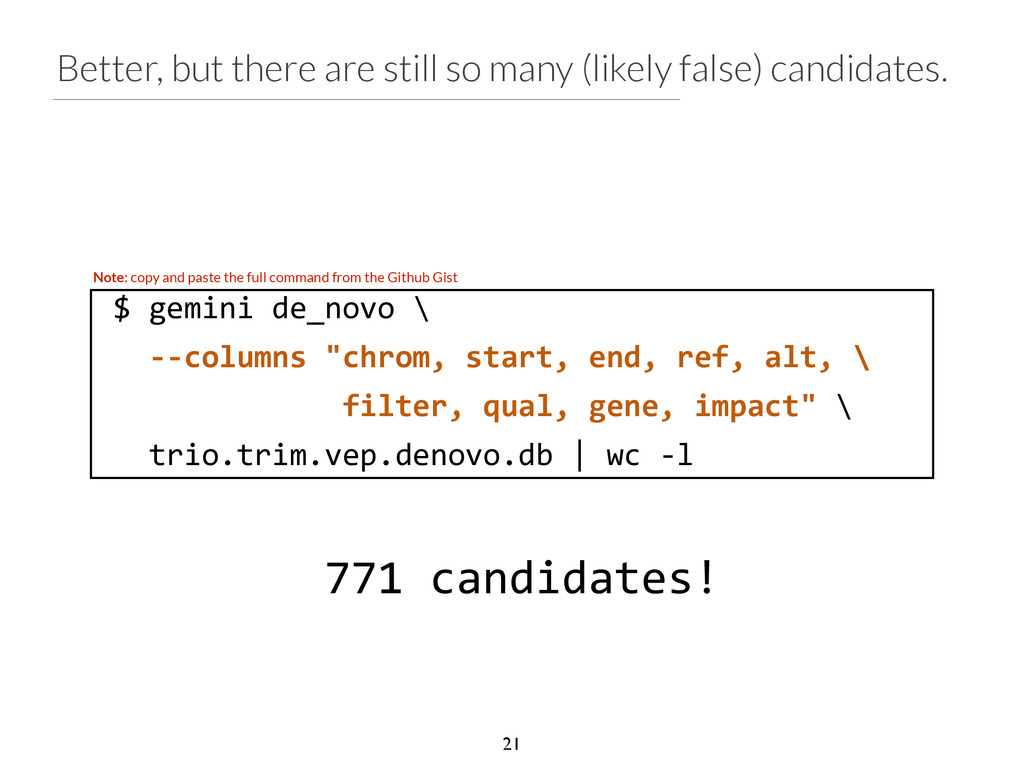
Better, but there are still so many (likely false) candidates.
$ gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ trio.trim.vep.denovo.db | wc -‐l Note: copy and paste the full command from the Github Gist 771 candidates! 21 -
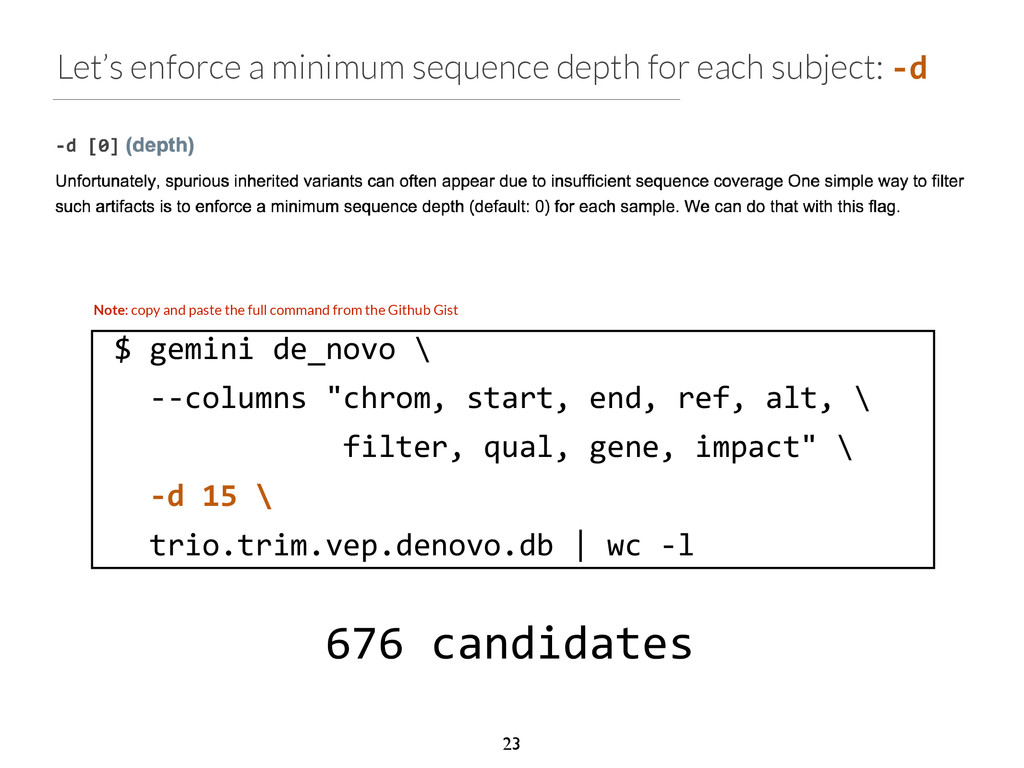
Let’s enforce a minimum sequence depth for each subject: -‐d
$ gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ -‐d 15 \ trio.trim.vep.denovo.db | wc -‐l Note: copy and paste the full command from the Github Gist 676 candidates 23 -
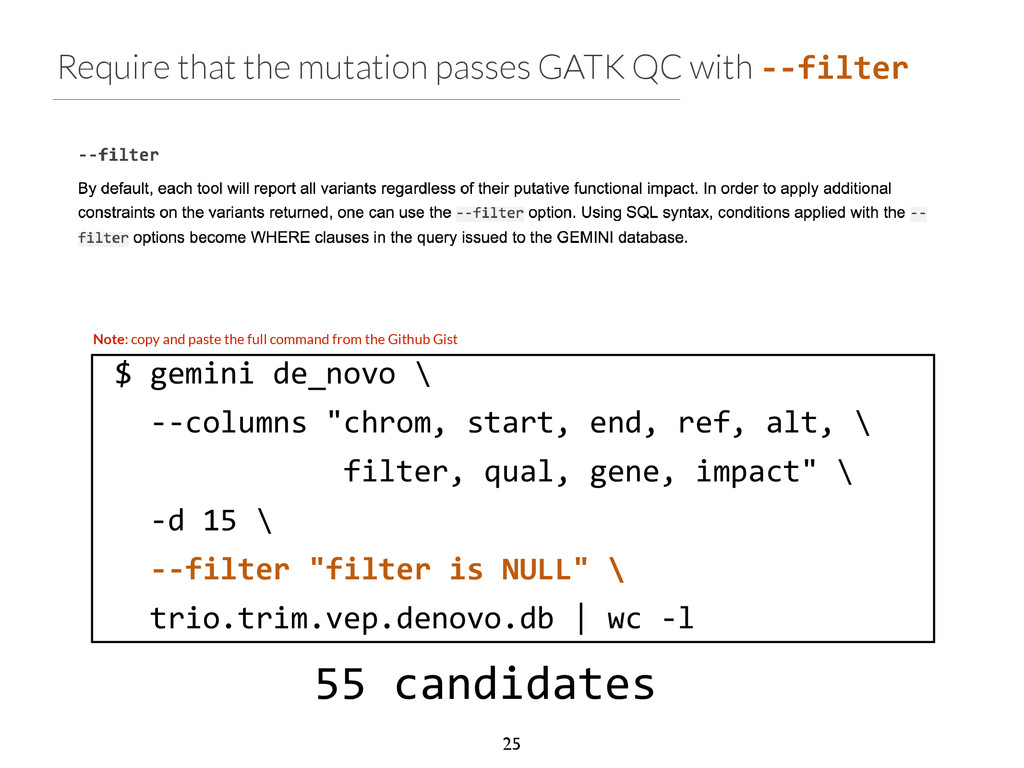
Require that the mutation passes GATK QC with -‐-‐filter $
gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ -‐d 15 \ -‐-‐filter "filter is NULL" \ trio.trim.vep.denovo.db | wc -‐l Note: copy and paste the full command from the Github Gist 55 candidates 25 -
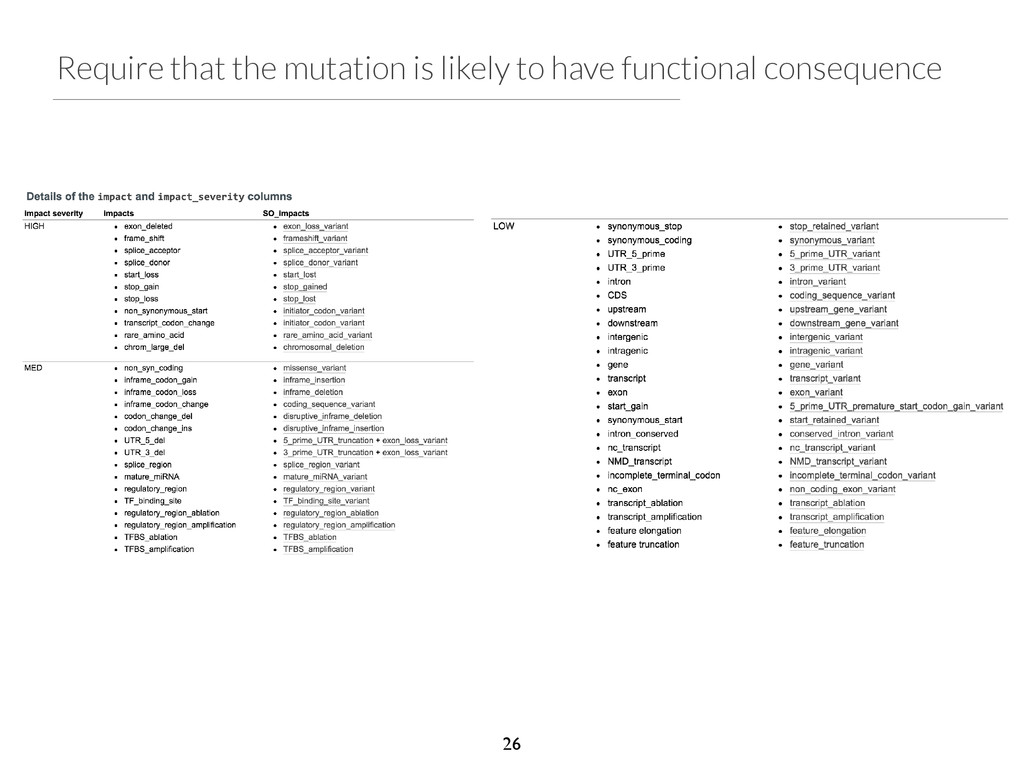
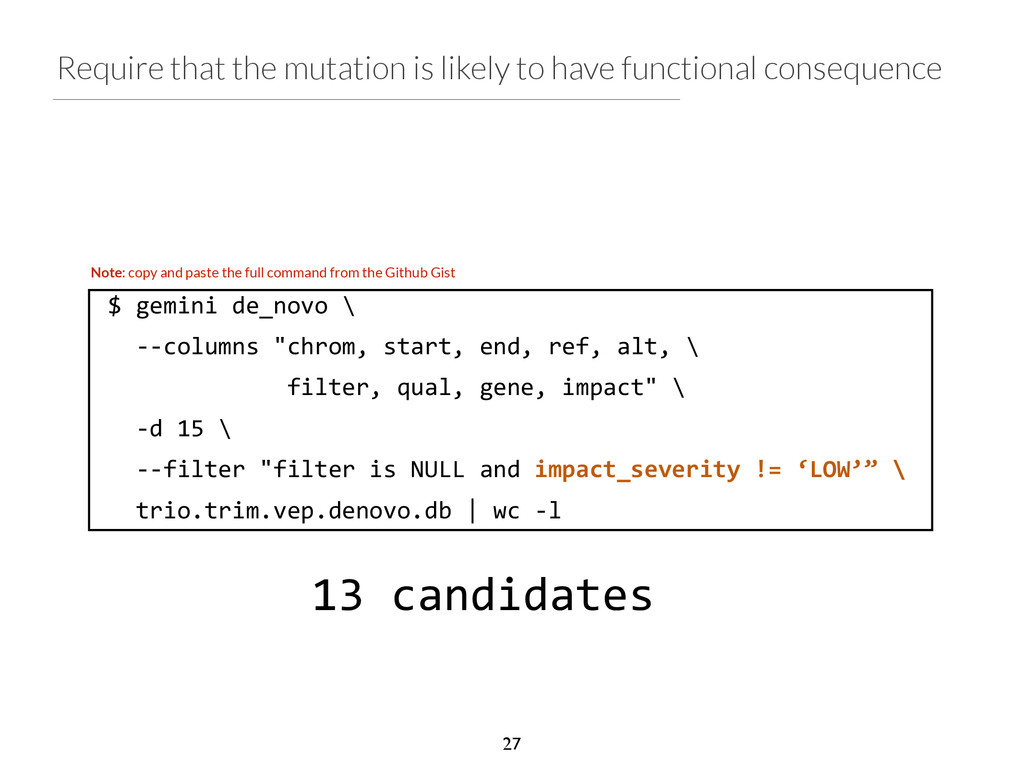
Require that the mutation is likely to have functional consequence
$ gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ -‐d 15 \ -‐-‐filter "filter is NULL and impact_severity != ‘LOW’” \ trio.trim.vep.denovo.db | wc -‐l Note: copy and paste the full command from the Github Gist 13 candidates 27 -
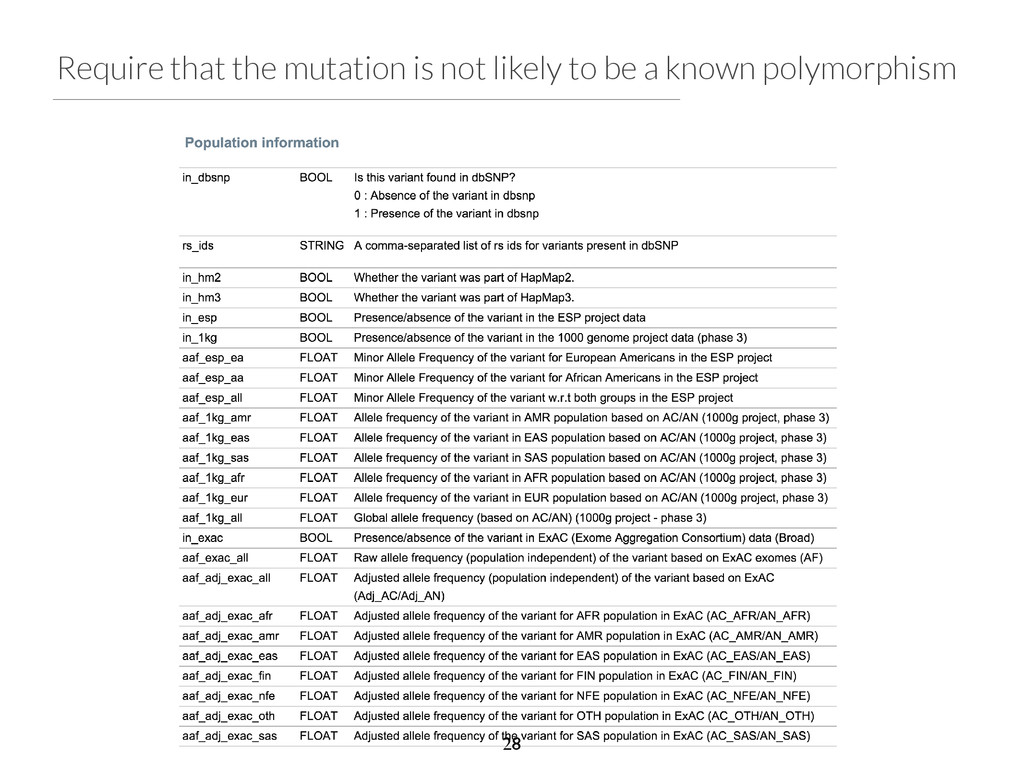
Require that the mutation is not likely to be a
known polymorphism 28 -
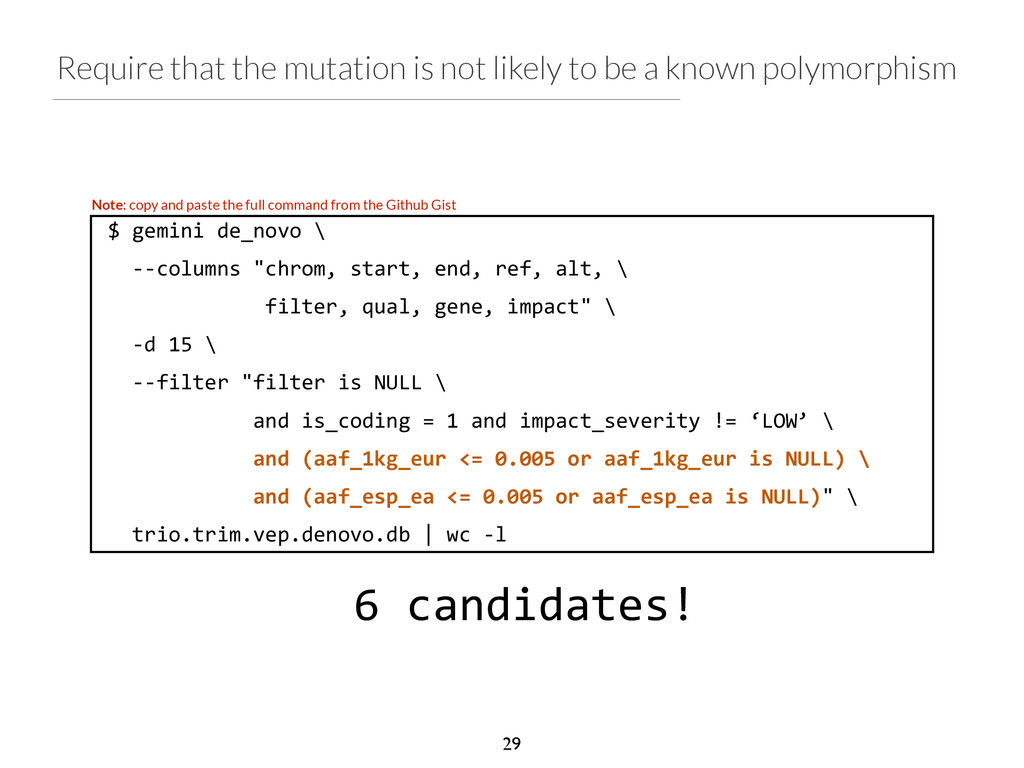
Require that the mutation is not likely to be a
known polymorphism Note: copy and paste the full command from the Github Gist $ gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ -‐d 15 \ -‐-‐filter "filter is NULL \ and is_coding = 1 and impact_severity != ‘LOW’ \ and (aaf_1kg_eur <= 0.005 or aaf_1kg_eur is NULL) \ and (aaf_esp_ea <= 0.005 or aaf_esp_ea is NULL)" \ trio.trim.vep.denovo.db | wc -‐l 6 candidates! 29 -
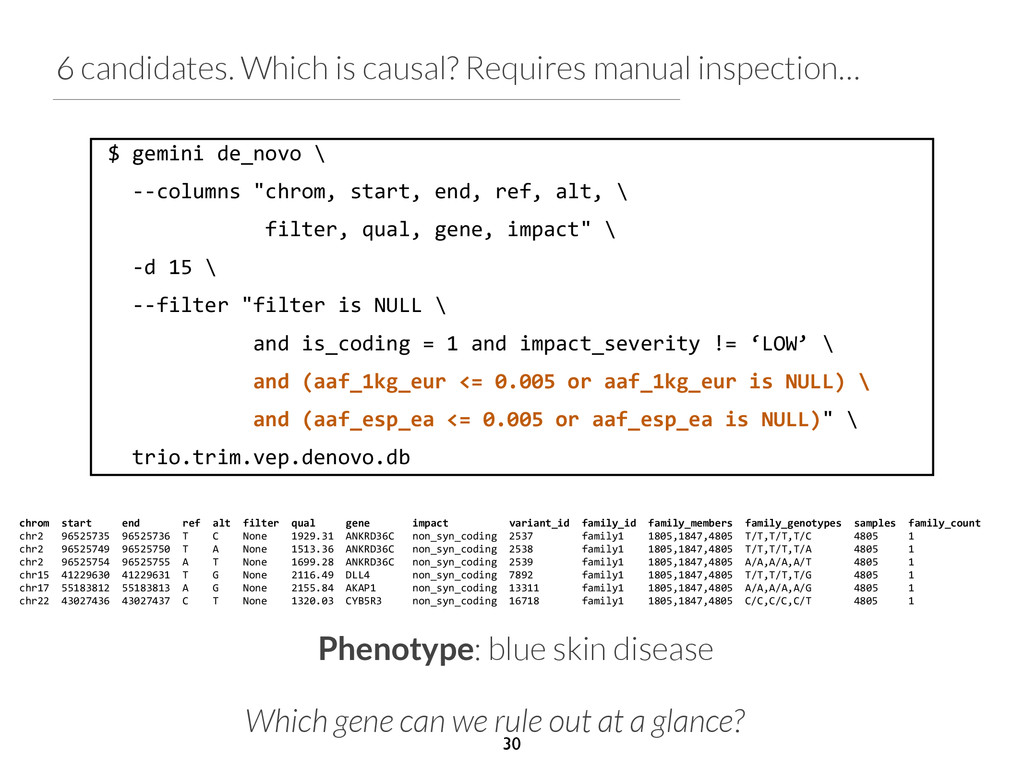
6 candidates. Which is causal? Requires manual inspection… chrom
start end ref alt filter qual gene impact variant_id family_id family_members family_genotypes samples family_count chr2 96525735 96525736 T C None 1929.31 ANKRD36C non_syn_coding 2537 family1 1805,1847,4805 T/T,T/T,T/C 4805 1 chr2 96525749 96525750 T A None 1513.36 ANKRD36C non_syn_coding 2538 family1 1805,1847,4805 T/T,T/T,T/A 4805 1 chr2 96525754 96525755 A T None 1699.28 ANKRD36C non_syn_coding 2539 family1 1805,1847,4805 A/A,A/A,A/T 4805 1 chr15 41229630 41229631 T G None 2116.49 DLL4 non_syn_coding 7892 family1 1805,1847,4805 T/T,T/T,T/G 4805 1 chr17 55183812 55183813 A G None 2155.84 AKAP1 non_syn_coding 13311 family1 1805,1847,4805 A/A,A/A,A/G 4805 1 chr22 43027436 43027437 C T None 1320.03 CYB5R3 non_syn_coding 16718 family1 1805,1847,4805 C/C,C/C,C/T 4805 1 Phenotype: blue skin disease $ gemini de_novo \ -‐-‐columns "chrom, start, end, ref, alt, \ filter, qual, gene, impact" \ -‐d 15 \ -‐-‐filter "filter is NULL \ and is_coding = 1 and impact_severity != ‘LOW’ \ and (aaf_1kg_eur <= 0.005 or aaf_1kg_eur is NULL) \ and (aaf_esp_ea <= 0.005 or aaf_esp_ea is NULL)" \ trio.trim.vep.denovo.db Which gene can we rule out at a glance? 30 -
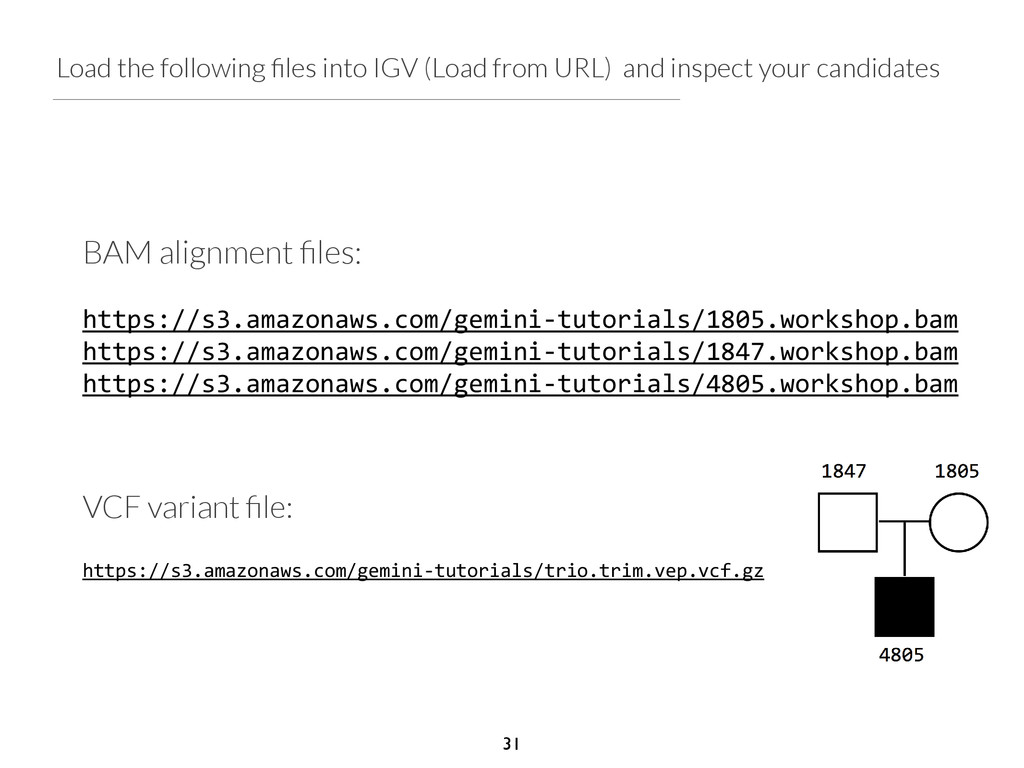
Load the following files into IGV (Load from URL) and
inspect your candidates BAM alignment files: ! https://s3.amazonaws.com/gemini-‐tutorials/1805.workshop.bam https://s3.amazonaws.com/gemini-‐tutorials/1847.workshop.bam https://s3.amazonaws.com/gemini-‐tutorials/4805.workshop.bam VCF variant file: ! https://s3.amazonaws.com/gemini-‐tutorials/trio.trim.vep.vcf.gz ! 31