Share
Embed
Making queries of the genome less difficult.
Presented on 30-Oct-2105 in the "Sequencing pipelines and assembly" session at the CSHL Genome Informatics Meeting.
Aaron Quinlan
October 31, 2015
Transcript
43 slides
-
Aaron Quinlan University of Utah quinlanlab.org @aaronquinlan ! ! !
! Making queries of the genome less difficult. -
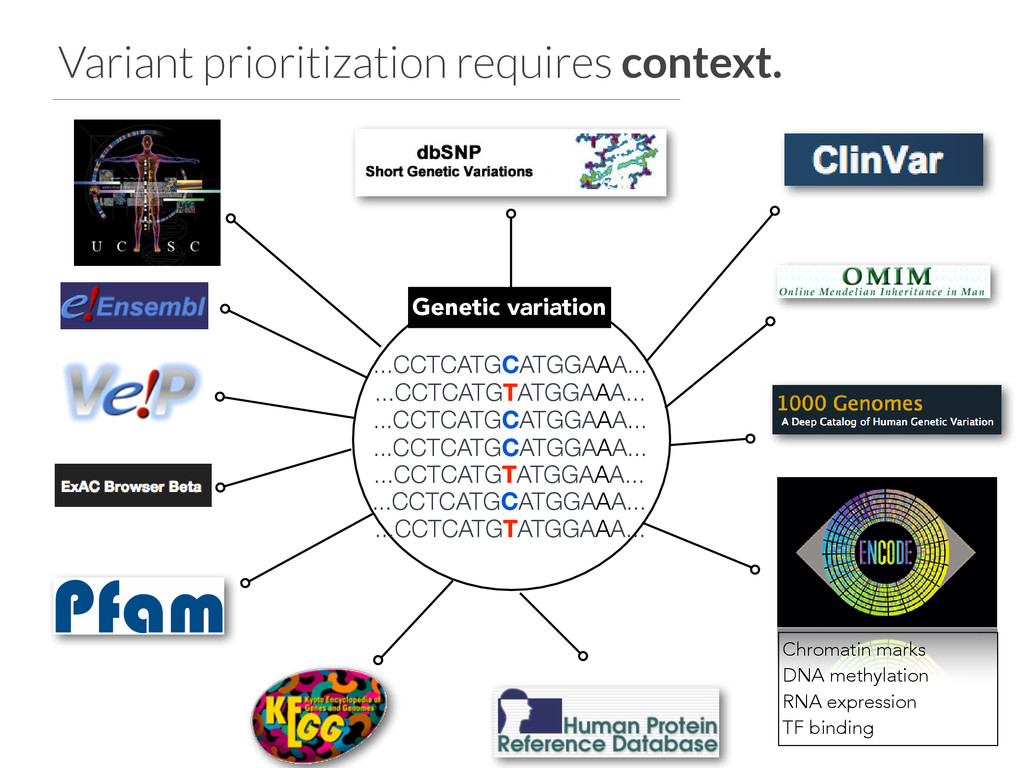
...CCTCATGCATGGAAA... Genetic variation ...CCTCATGTATGGAAA... ...CCTCATGCATGGAAA... ...CCTCATGCATGGAAA... ...CCTCATGTATGGAAA... ...CCTCATGCATGGAAA... ...CCTCATGTATGGAAA... Chromatin
marks DNA methylation RNA expression TF binding Variant prioritization requires context. -
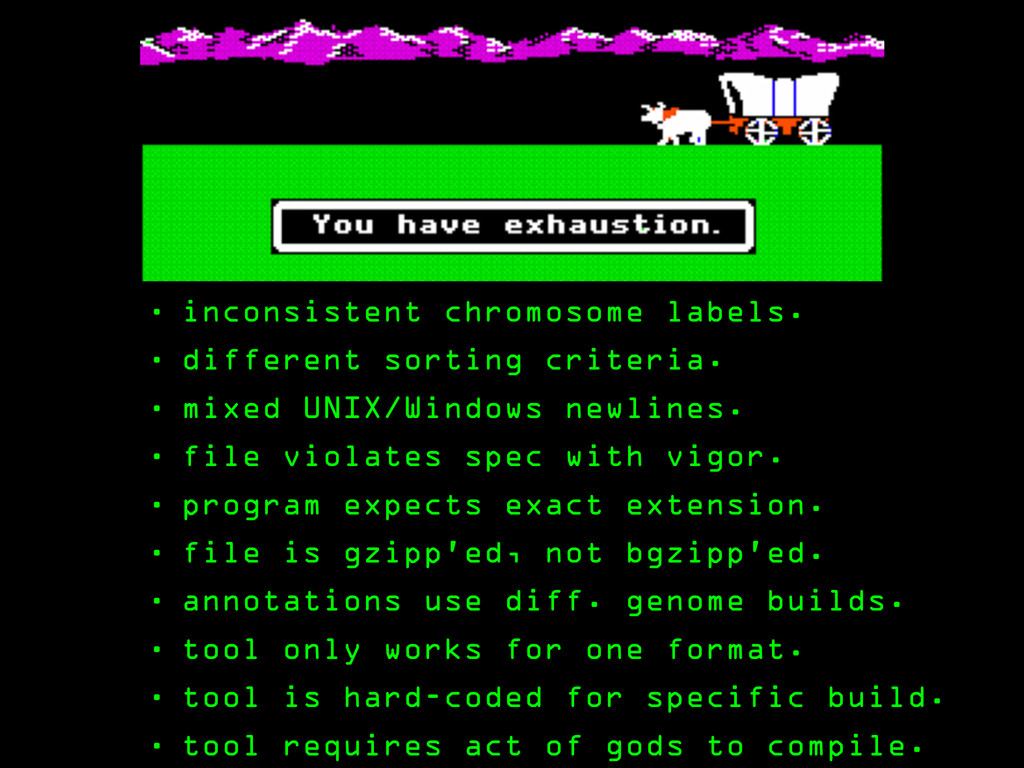
• inconsistent chromosome labels. • different sorting criteria. • mixed
UNIX/Windows newlines. • file violates spec with vigor. • program expects exact extension. • file is gzipp’ed, not bgzipp’ed. • annotations use diff. genome builds. • tool only works for one format. • tool is hard-coded for specific build. • tool requires act of gods to compile. -
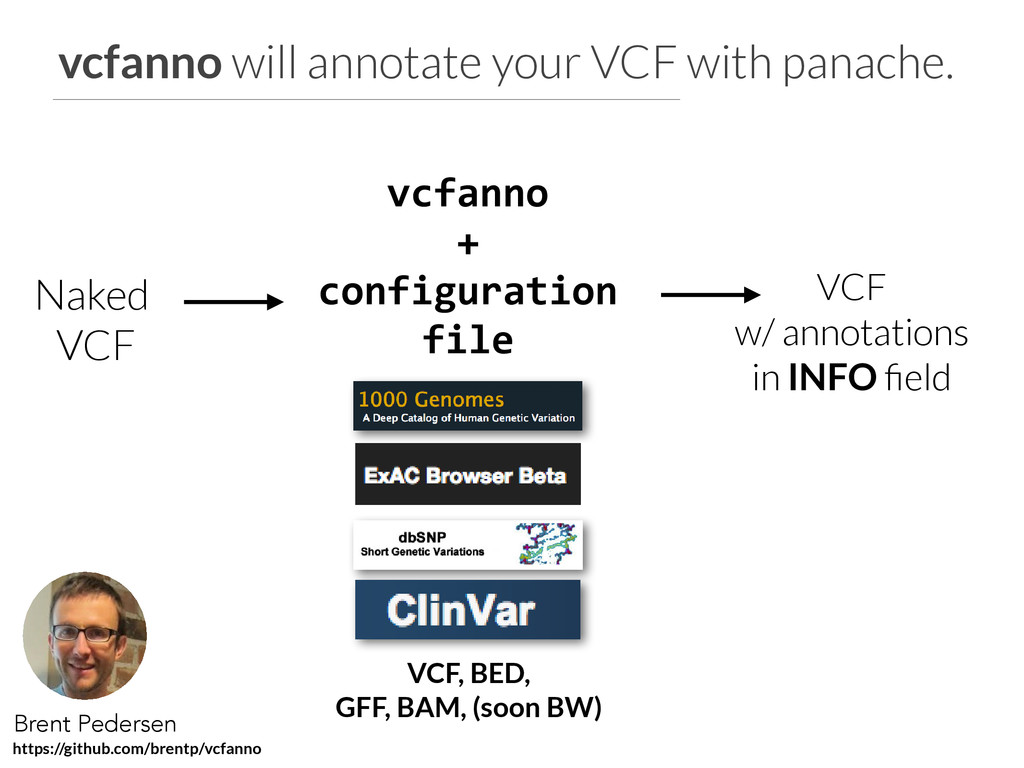
vcfanno will annotate your VCF with panache. Naked VCF vcfanno
+ configuration file VCF w/ annotations in INFO field Brent Pedersen https://github.com/brentp/vcfanno VCF, BED, GFF, BAM, (soon BW) -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. Match on POS+REF+ALT for VCF annotations. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. Allows multiple annotations from each file Match on POS+REF+ALT for VCF annotations. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. Allows multiple annotations from each file Can rename the annotations in the resulting VCF Match on POS+REF+ALT for VCF annotations. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. Allows multiple annotations from each file Can rename the annotations in the resulting VCF Multiple operations to summarize the results of multiple hits in annot. file: mean, max, min concat, count, uniq first, flag Match on POS+REF+ALT for VCF annotations. -
[[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”]
ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] vcfanno configuration file. Allows multiple annotations from each file Can rename the annotations in the resulting VCF Multiple operations to summarize the results of multiple hits in annot. file: mean, max, min concat, count, uniq first, flag Match on POS+REF+ALT for VCF annotations. Javascript for custom computations. variance() defined in custom.js -
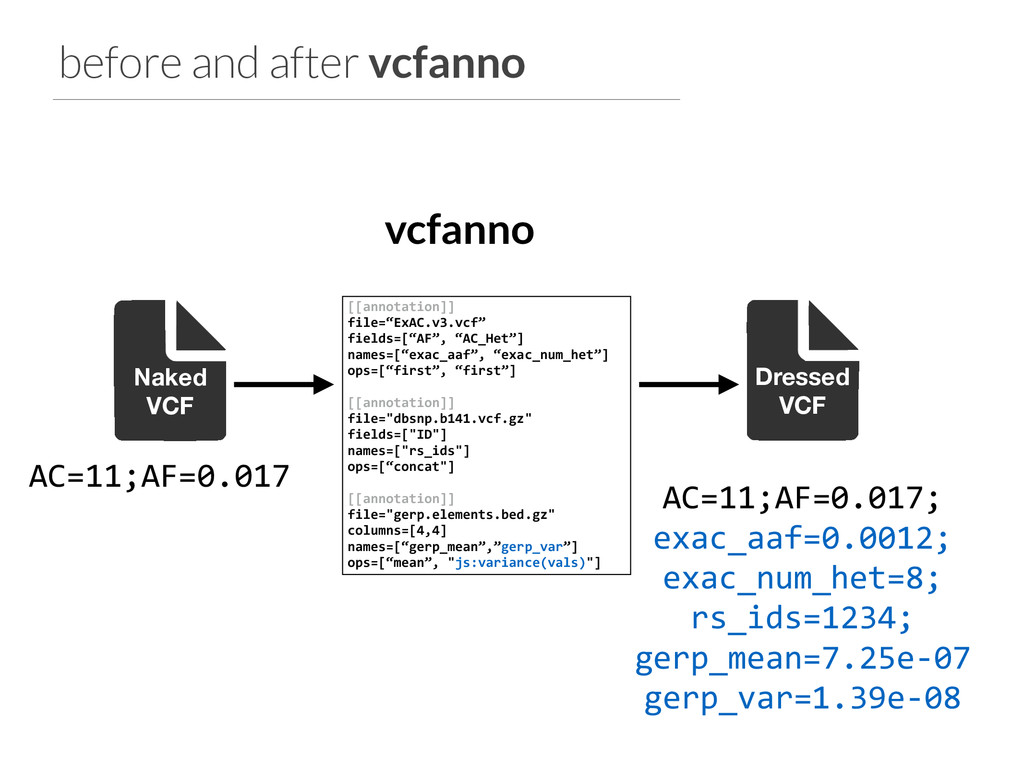
before and after vcfanno vcfanno AC=11;AF=0.017 AC=11;AF=0.017; exac_aaf=0.0012;
exac_num_het=8; rs_ids=1234; gerp_mean=7.25e-‐07 gerp_var=1.39e-‐08 Naked VCF Dressed VCF [[annotation]] file=“ExAC.v3.vcf” fields=[“AF”, “AC_Het”] names=[“exac_aaf”, “exac_num_het”] ops=[“first”, “first”] ! [[annotation]] file="dbsnp.b141.vcf.gz" fields=["ID"] names=["rs_ids"] ops=[“concat"] ! [[annotation]] file="gerp.elements.bed.gz" columns=[4,4] names=[“gerp_mean”,”gerp_var”] ops=[“mean”, "js:variance(vals)"] -
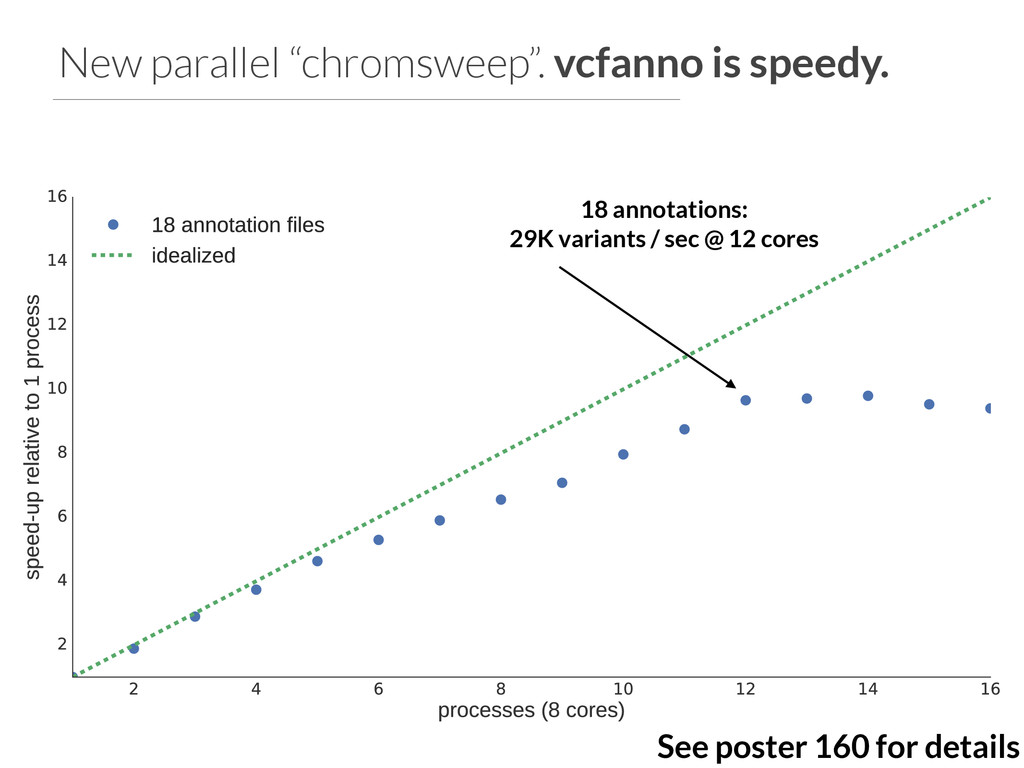
New parallel “chromsweep”. vcfanno is speedy. 18 annotations: 29K variants
/ sec @ 12 cores See poster 160 for details -
Individual-centric queries with Genotype Query Tools (GQT) github.com/ryanlayer/gqt In press.
Ryan Layer http://biorxiv.org/content/early/2015/04/20/018259 -
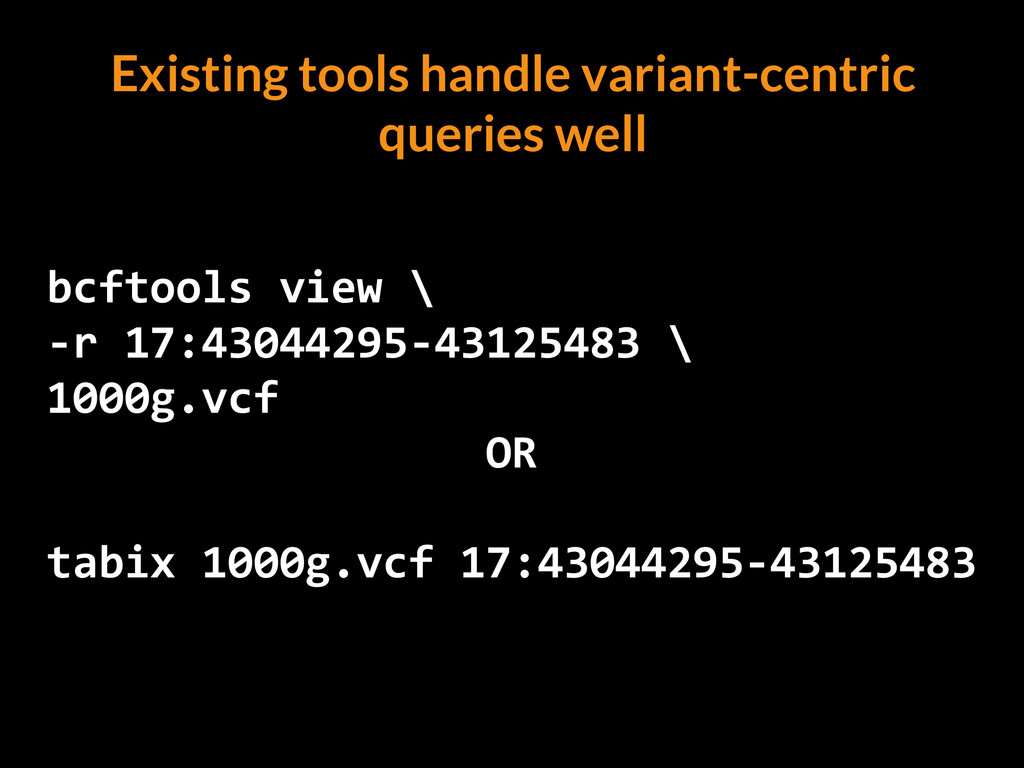
bcftools view \ -‐r 17:43044295-‐43125483 \ 1000g.vcf
OR ! tabix 1000g.vcf 17:43044295-‐43125483 ! Existing tools handle variant-centric queries well -
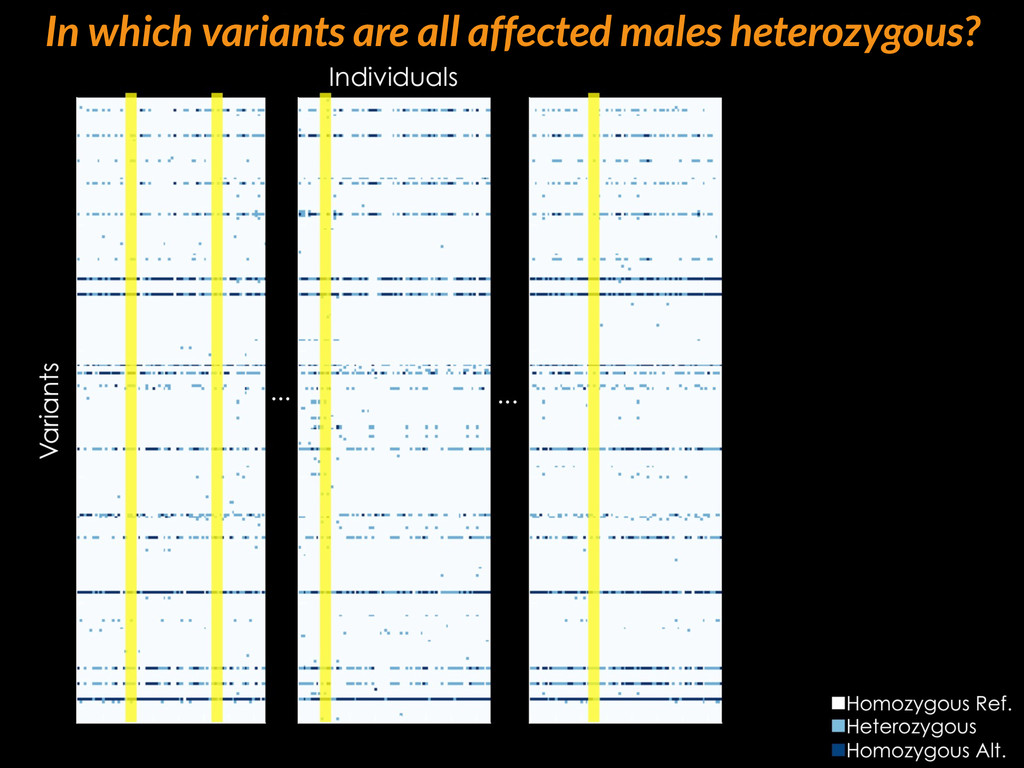
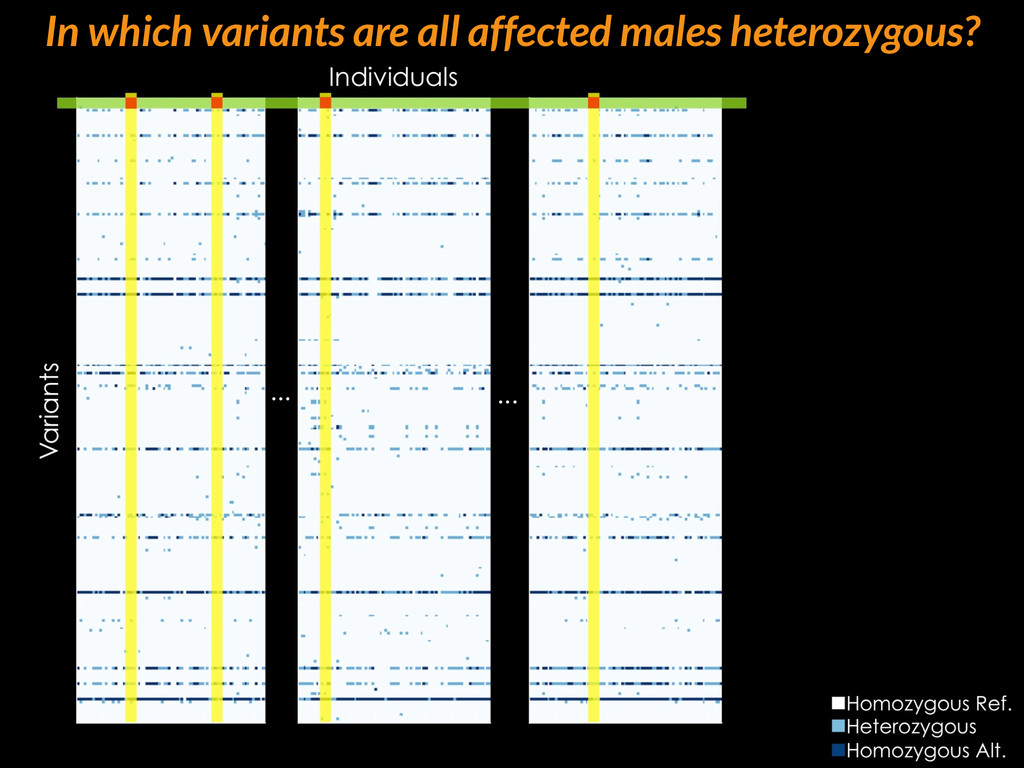
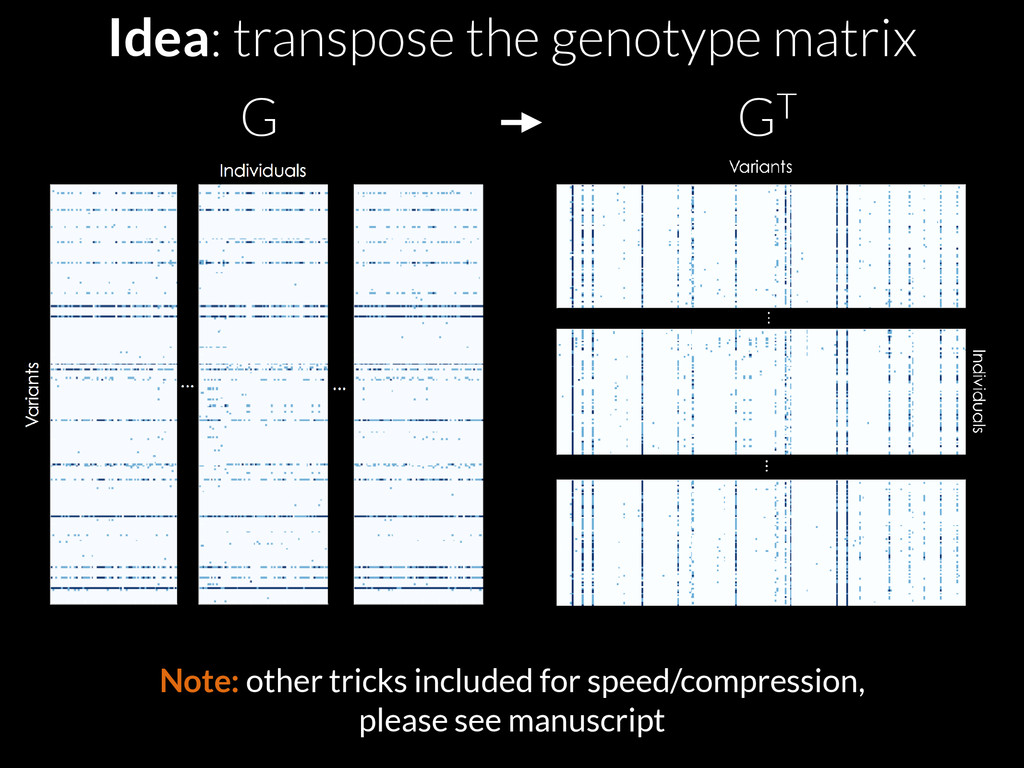
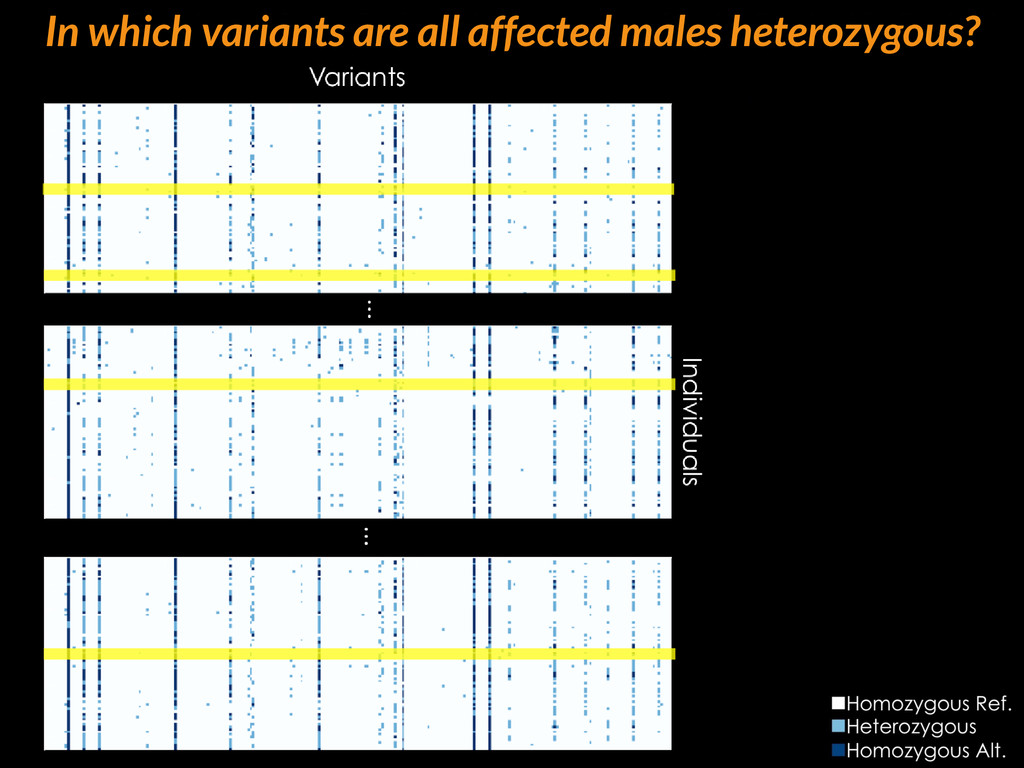
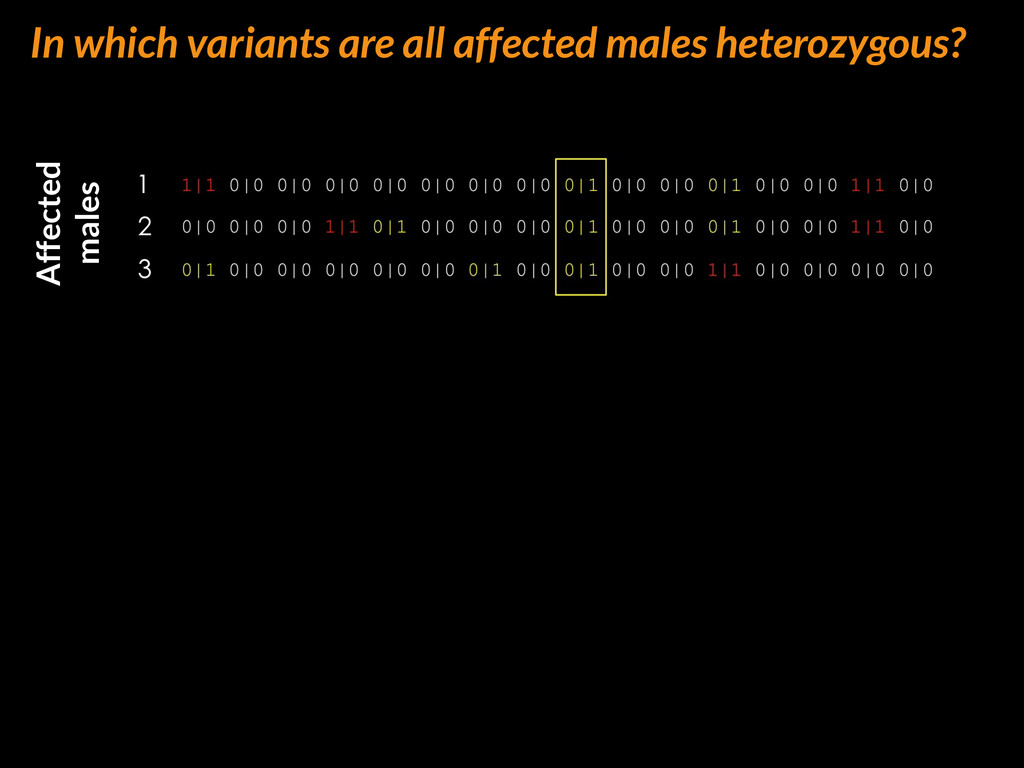
Idea: transpose the genotype matrix G GT Note: other tricks
included for speed/compression, please see manuscript -
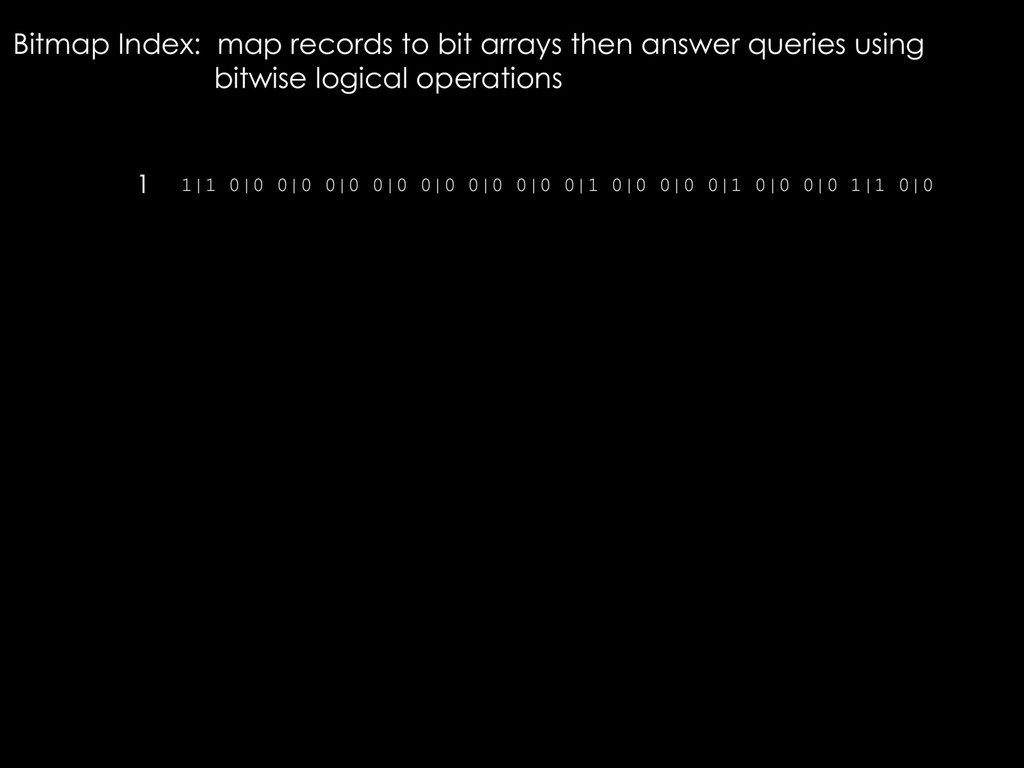
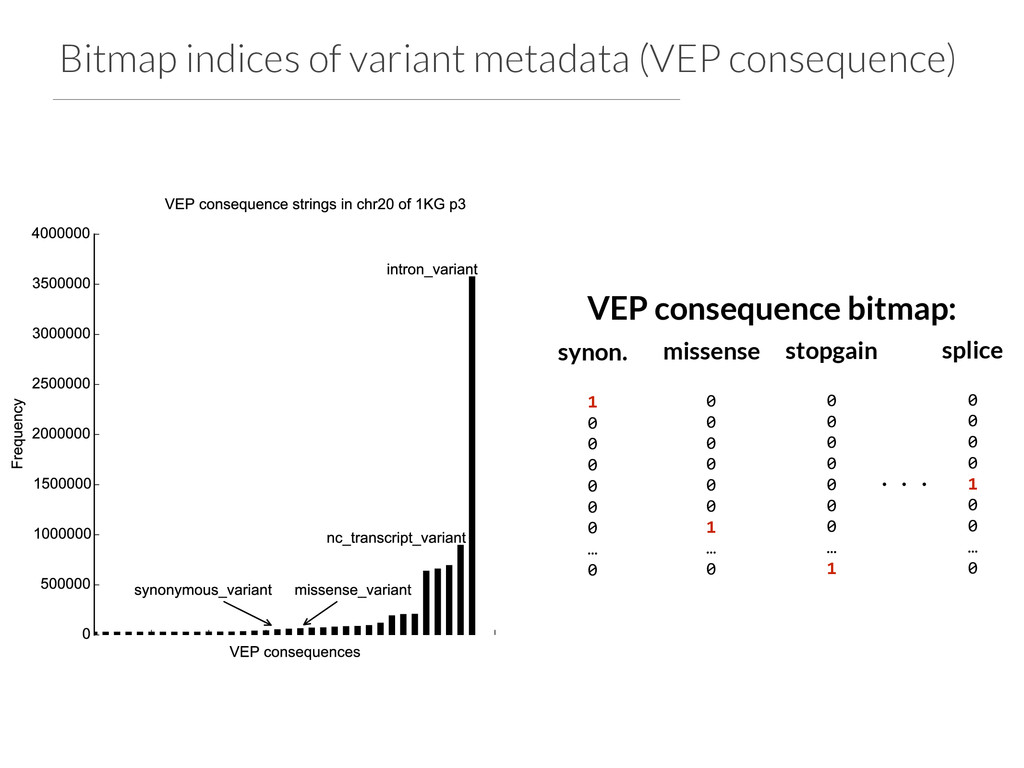
Bitmap indices of variant metadata (VEP consequence) VEP consequence bitmap:
1 0 0 0 0 0 0 … 0 synon. missense 0 0 0 0 0 0 1 … 0 stopgain 0 0 0 0 0 0 0 … 1 splice 0 0 0 0 1 0 0 … 0 . . . -
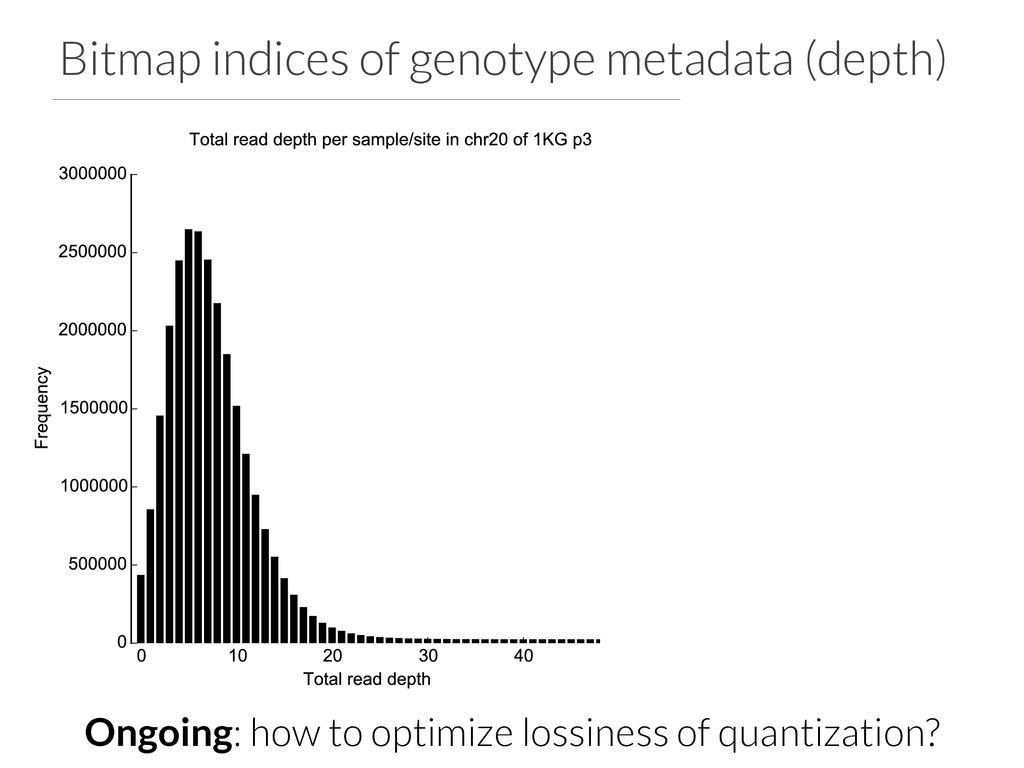
Bitmap indices of genotype metadata (depth) Ongoing: how to optimize
lossiness of quantization? -
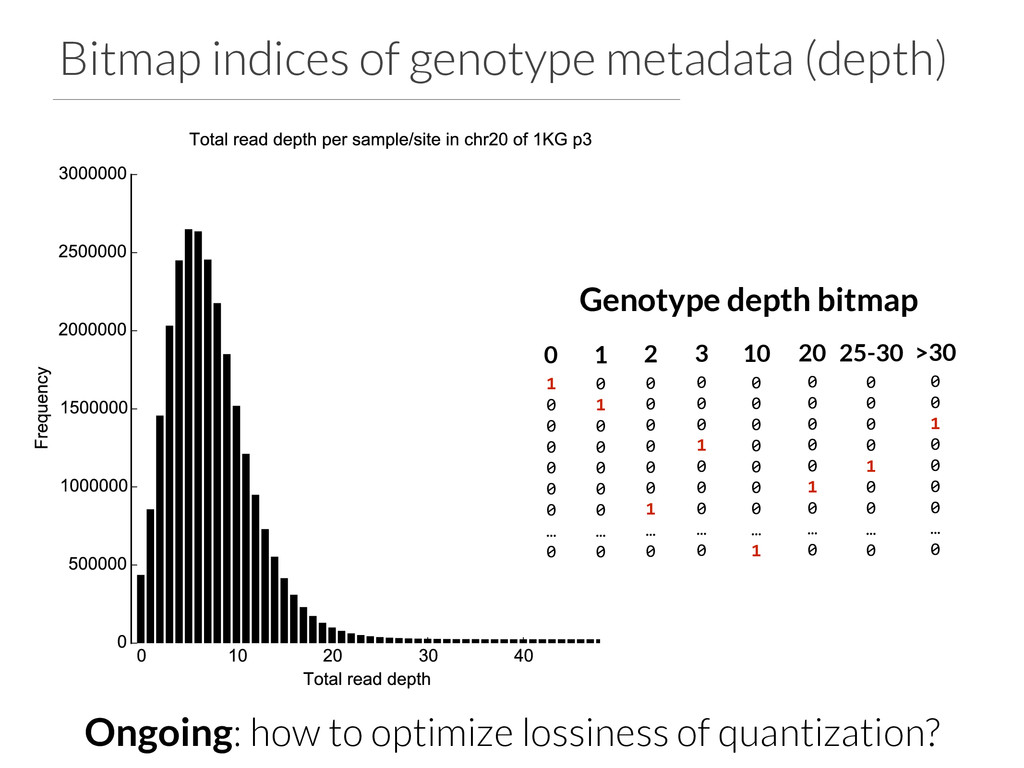
Bitmap indices of genotype metadata (depth) Genotype depth bitmap 1
0 0 0 0 0 0 … 0 0 1 0 1 0 0 0 0 0 … 0 2 0 0 0 0 0 0 1 … 0 3 0 0 0 1 0 0 0 … 0 10 0 0 0 0 0 0 0 … 1 20 0 0 0 0 0 1 0 … 0 25-30 0 0 0 0 1 0 0 … 0 >30 0 0 1 0 0 0 0 … 0 Ongoing: how to optimize lossiness of quantization? -
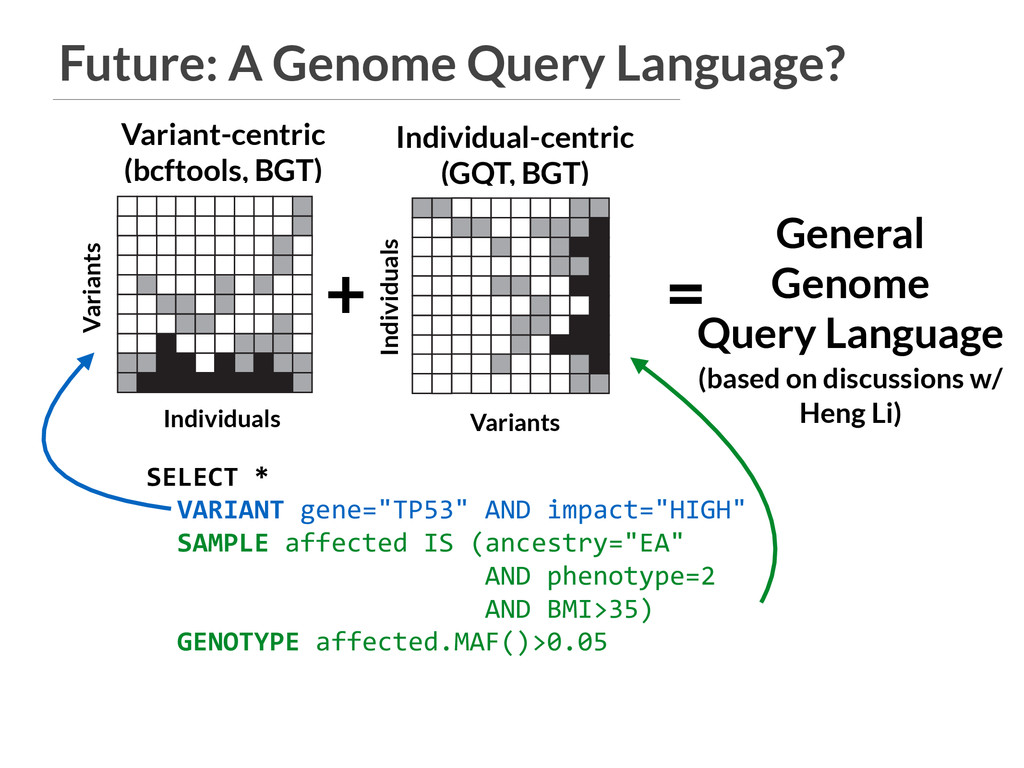
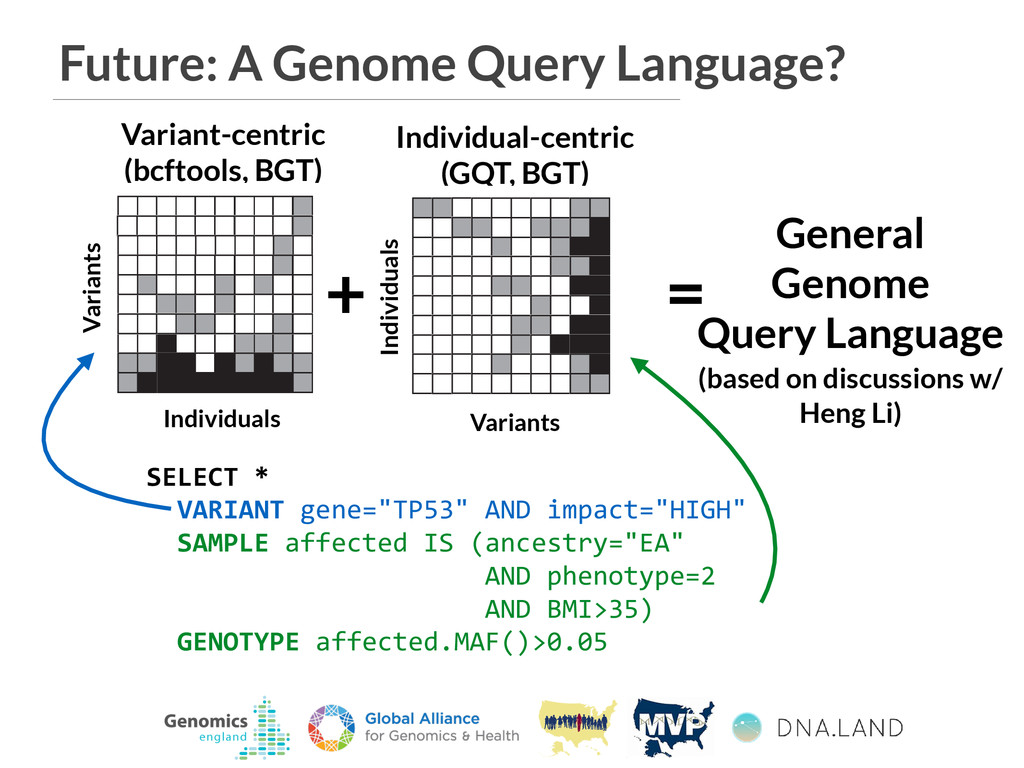
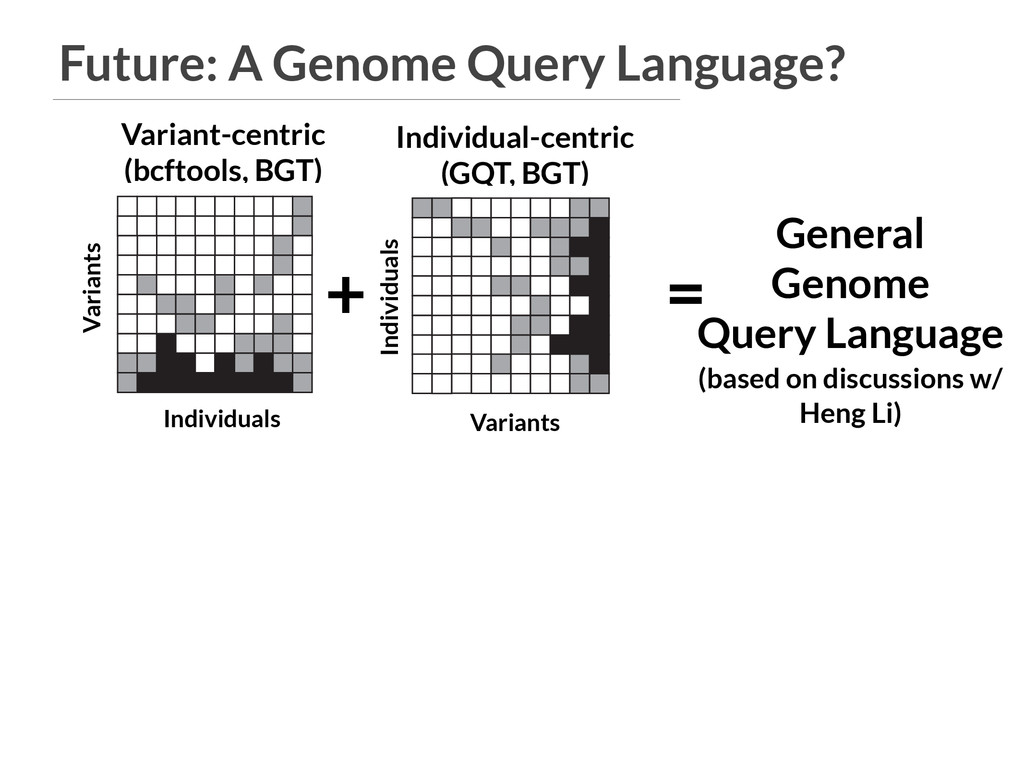
Future: A Genome Query Language? Variant-centric (bcftools, BGT) + =
General Genome Query Language (based on discussions w/ Heng Li) VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in cases and rare in controls. b gqt query study.gqt study.db -p "phenotype == 2" -g "maf() > 0.05" -p "phenotype == 1" -g "maf() < 0.05" gqt -p -g b VCF In F V In VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in g Individual-centric (GQT, BGT) Individuals Variants Variants Individuals -
Future: A Genome Query Language? Variant-centric (bcftools, BGT) + =
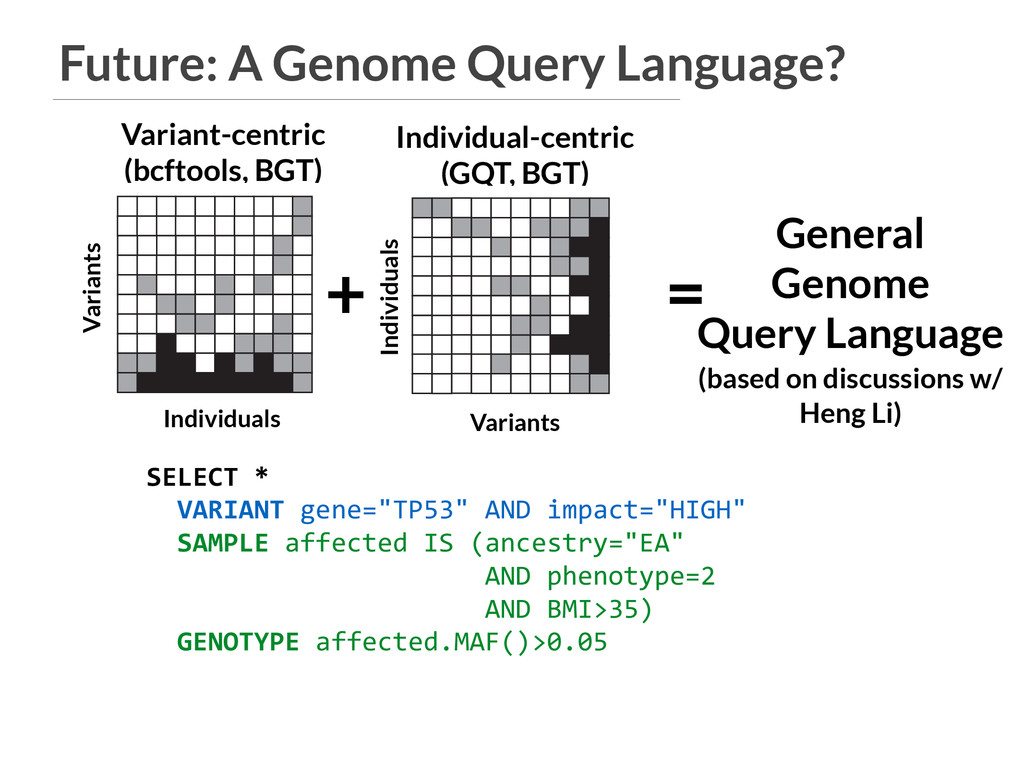
General Genome Query Language (based on discussions w/ Heng Li) VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in cases and rare in controls. b gqt query study.gqt study.db -p "phenotype == 2" -g "maf() > 0.05" -p "phenotype == 1" -g "maf() < 0.05" gqt -p -g b VCF In F V In VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in g Individual-centric (GQT, BGT) Individuals Variants Variants Individuals SELECT * VARIANT gene="TP53" AND impact="HIGH" SAMPLE affected IS (ancestry="EA" AND phenotype=2 AND BMI>35) GENOTYPE affected.MAF()>0.05 -
Future: A Genome Query Language? Variant-centric (bcftools, BGT) + =
General Genome Query Language (based on discussions w/ Heng Li) VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in cases and rare in controls. b gqt query study.gqt study.db -p "phenotype == 2" -g "maf() > 0.05" -p "phenotype == 1" -g "maf() < 0.05" gqt -p -g b VCF In F V In VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in g Individual-centric (GQT, BGT) Individuals Variants Variants Individuals SELECT * VARIANT gene="TP53" AND impact="HIGH" SAMPLE affected IS (ancestry="EA" AND phenotype=2 AND BMI>35) GENOTYPE affected.MAF()>0.05 -
Future: A Genome Query Language? Variant-centric (bcftools, BGT) + =
General Genome Query Language (based on discussions w/ Heng Li) VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in cases and rare in controls. b gqt query study.gqt study.db -p "phenotype == 2" -g "maf() > 0.05" -p "phenotype == 1" -g "maf() < 0.05" gqt -p -g b VCF In F V In VCF A B PED SQL database GQT index Individuals Variants 3 4 5 6 9 gqt convert ped gqt convert vcf D C Find variants that are common in g Individual-centric (GQT, BGT) Individuals Variants Variants Individuals SELECT * VARIANT gene="TP53" AND impact="HIGH" SAMPLE affected IS (ancestry="EA" AND phenotype=2 AND BMI>35) GENOTYPE affected.MAF()>0.05