Share
Embed
Kamil S. Jaron polyploidy webinar
Kamil S Jaroň
April 20, 2020
Transcript
49 slides
-
Characterising features of polyploid genomes directly from sequencing reads Kamil
S. Jaron @KamilSJaron April 20, 2020 1 -
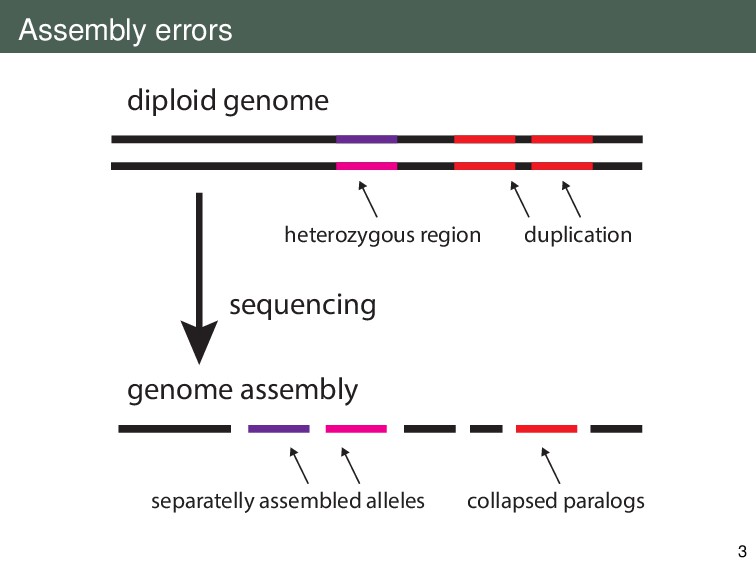
Assembly errors diploid genome duplication heterozygous region sequencing genome assembly
separatelly assembled alleles collapsed paralogs 3 -
Genome assembly is even more difficult for polyploids! More prior
information the better! 4 -
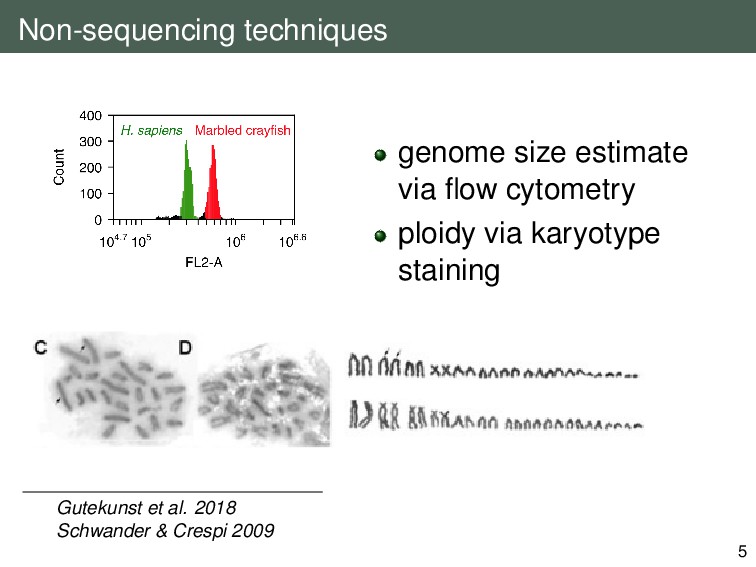
Non-sequencing techniques genome size estimate via flow cytometry ploidy via
karyotype staining Gutekunst et al. 2018 Schwander & Crespi 2009 5 -
Features from raw sequencing reads An individual genome genome size
repetitivness transposable element loads heterozygosity ploidy Population genomics divergence of two individuals GWAS references in supplementary slides 6 -
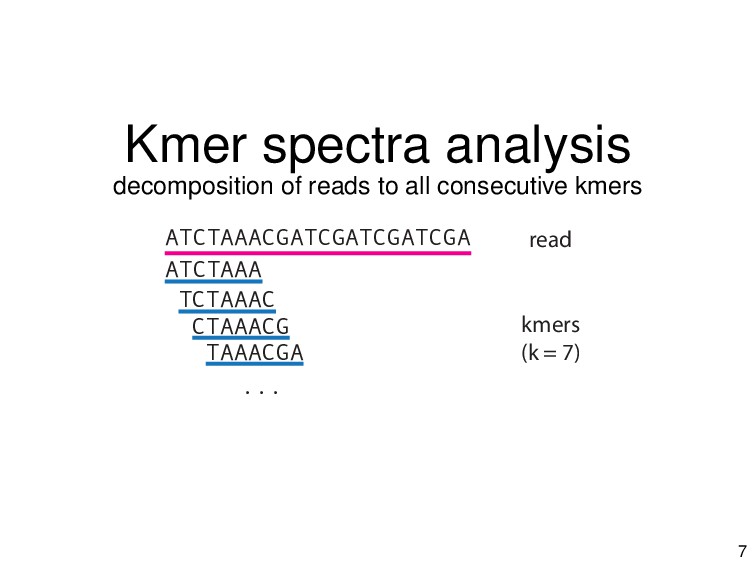
Kmer spectra analysis decomposition of reads to all consecutive kmers
read kmers (k = 7) ATCTAAACGATCGATCGATCGA ATCTAAA TCTAAAC CTAAACG TAAACGA ... 7 -
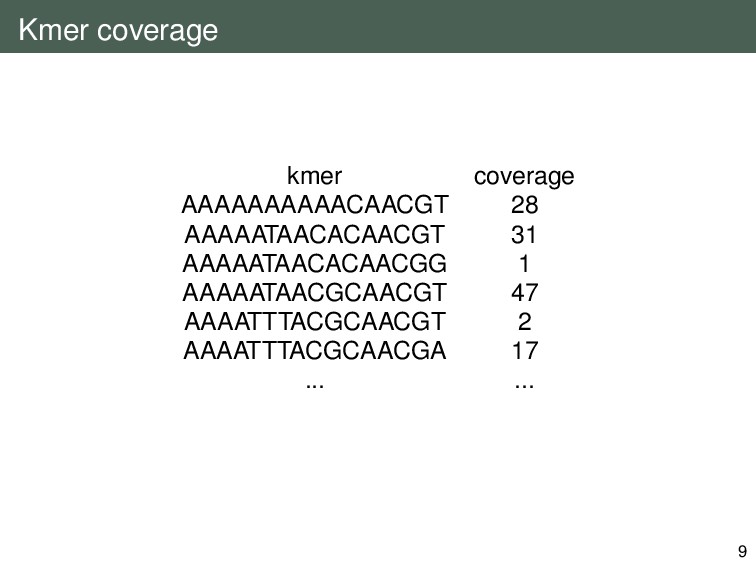
Kmer coverage kmer coverage AAAAAAAAAACAACGT 28 AAAAATAACACAACGT 31 AAAAATAACACAACGG 1
AAAAATAACGCAACGT 47 AAAATTTACGCAACGT 2 AAAATTTACGCAACGA 17 ... ... 9 -
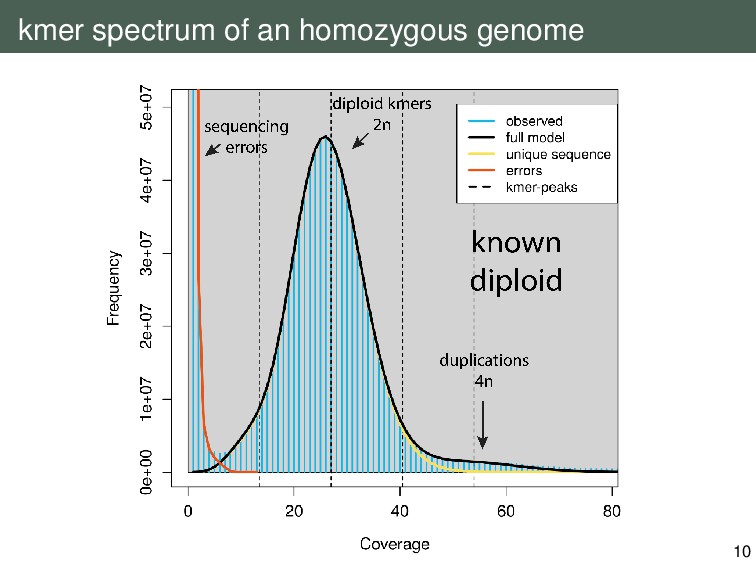
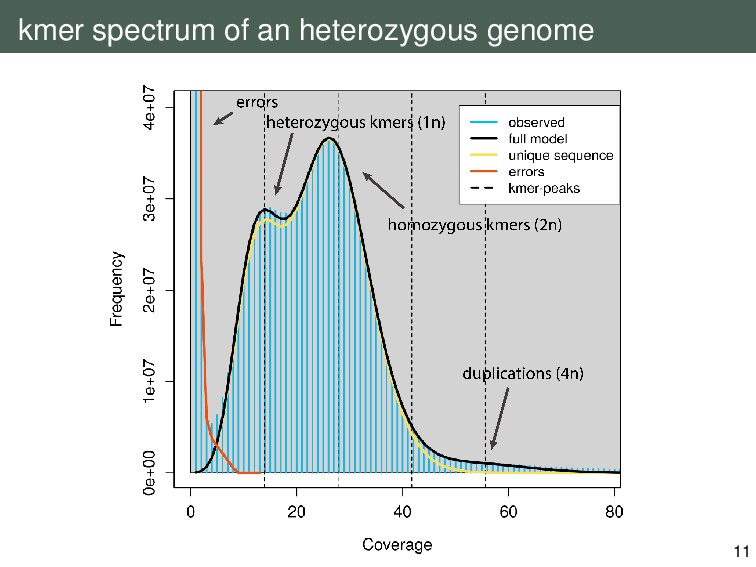
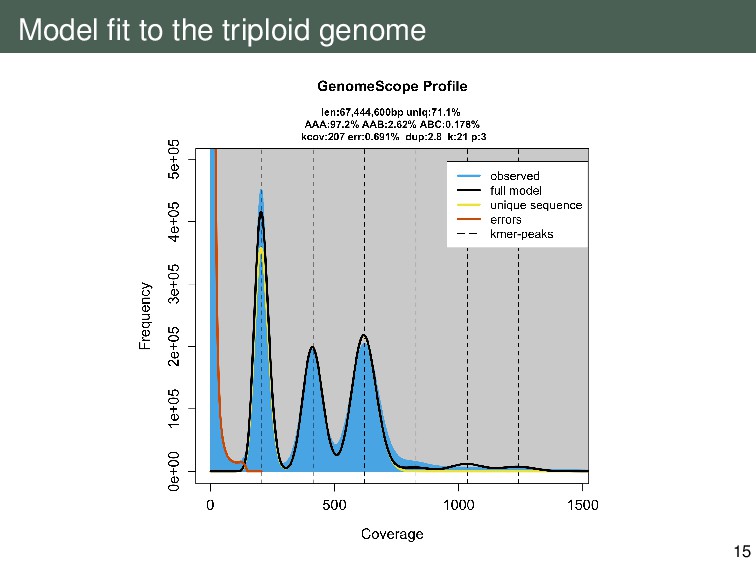
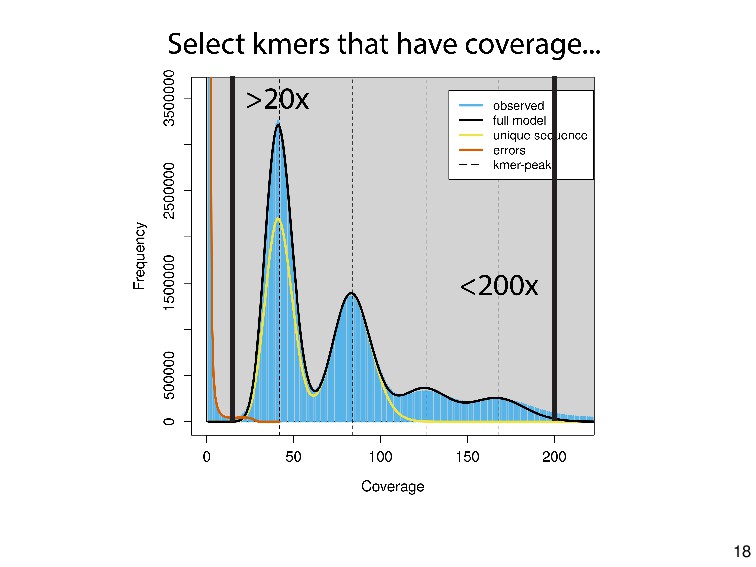
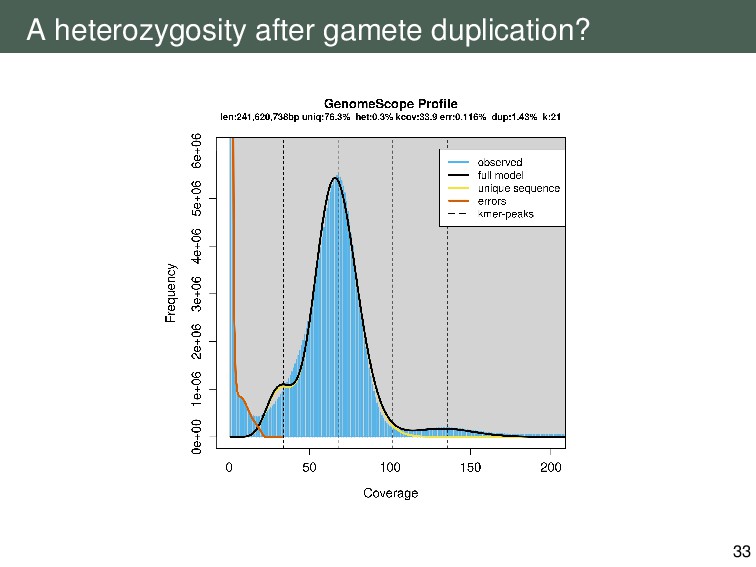
GenomeScope: given kmer spectra, what is... 1n coverage the genome
length heterozygosity repetitivness (% of the genome that is unique) Vurture et al. 2017 12 -
GenomeScope 2.0 given ploidy, for up to hexaploids ... Rhyker
Ranallo-Benavidez 13 -
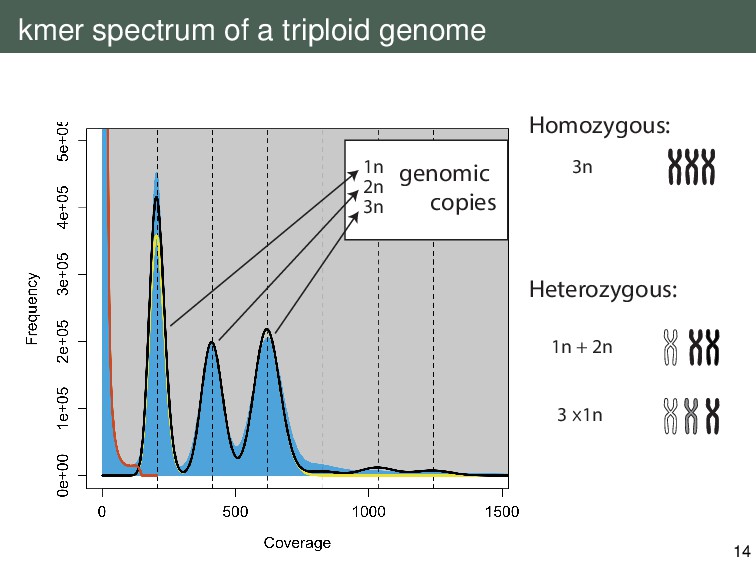
kmer spectrum of a triploid genome Homozygous: Heterozygous: 3n 1n
+ 2n 3 x1n 3n 2n 1n genomic copies 14 -
Is there more information in the kmer spectra to utilise?
Every heterozygous kmers has the sister kmers sequenced as well. 17 -
Search for kmer pairs! kmers that are different in a
single SNP kmers that form a unique pair 19 -
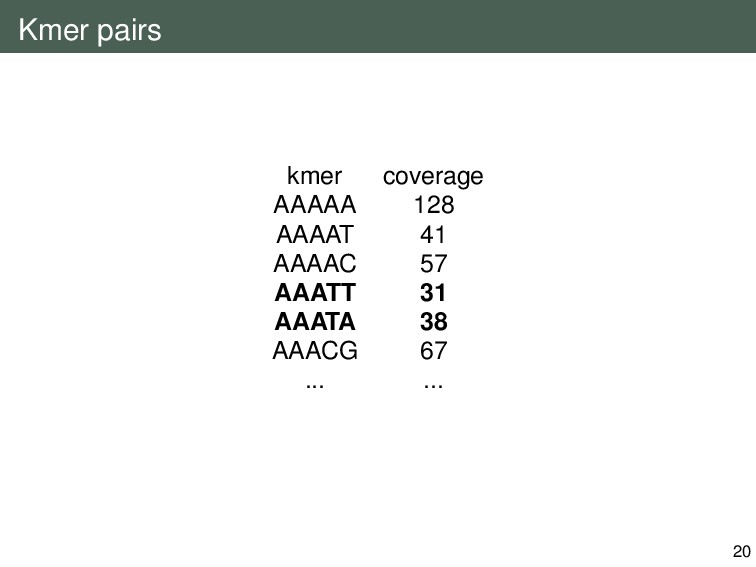
Kmer pairs kmer coverage AAAAA 128 AAAAT 41 AAAAC 57
AAATT 31 AAATA 38 AAACG 67 ... ... 20 -
The idea kmer pairs are mostly two alleles of the
same locus. 21 -
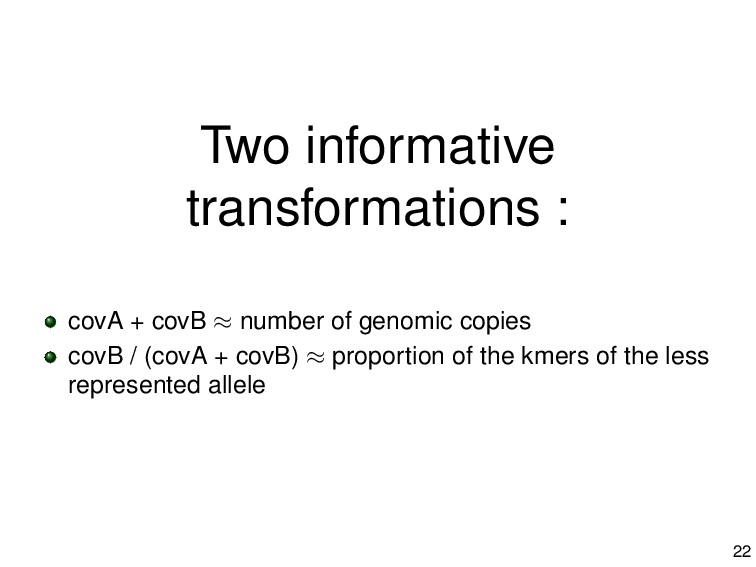
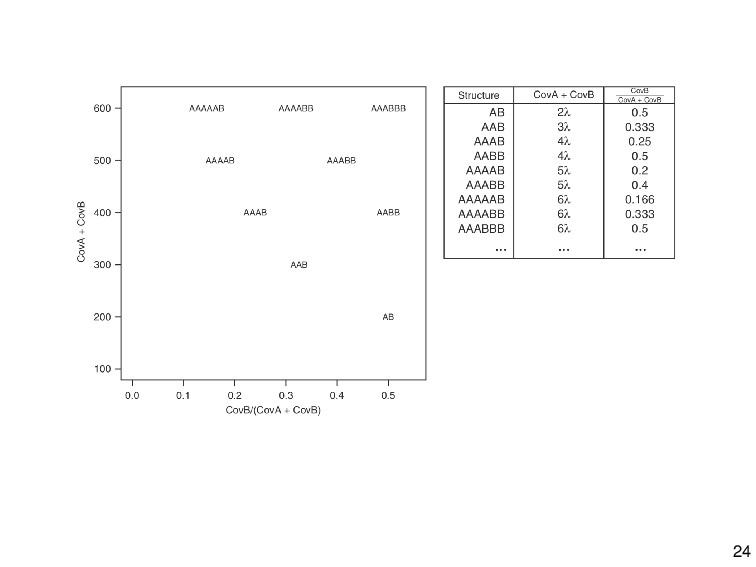
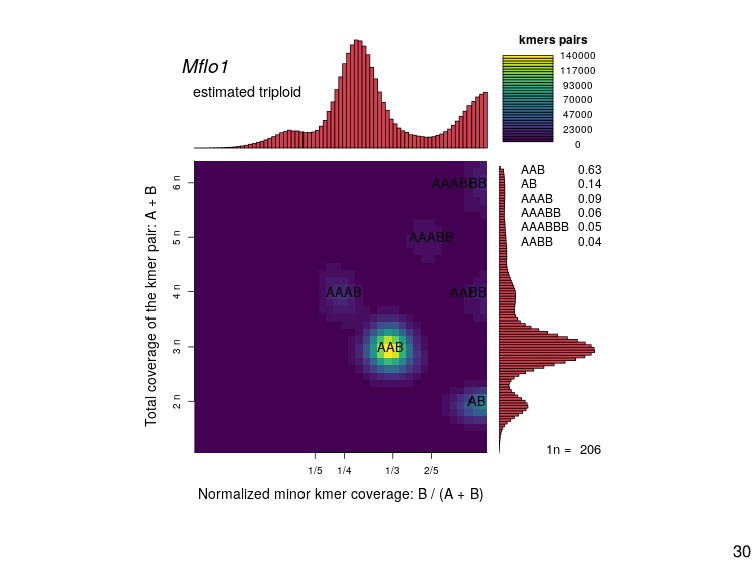
Two informative transformations : covA + covB ≈ number of
genomic copies covB / (covA + covB) ≈ proportion of the kmers of the less represented allele 22 -
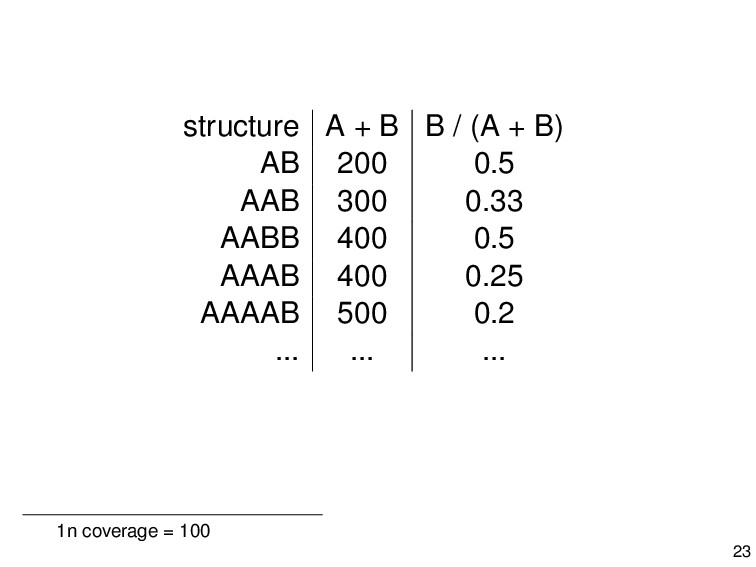
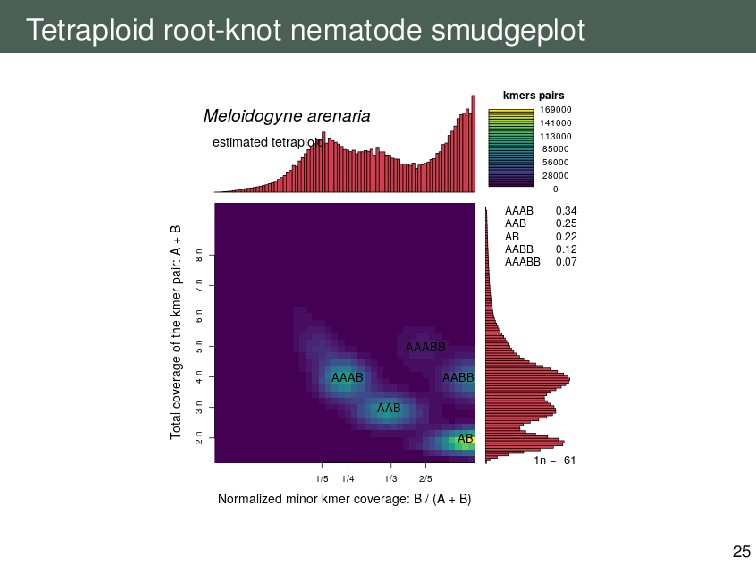
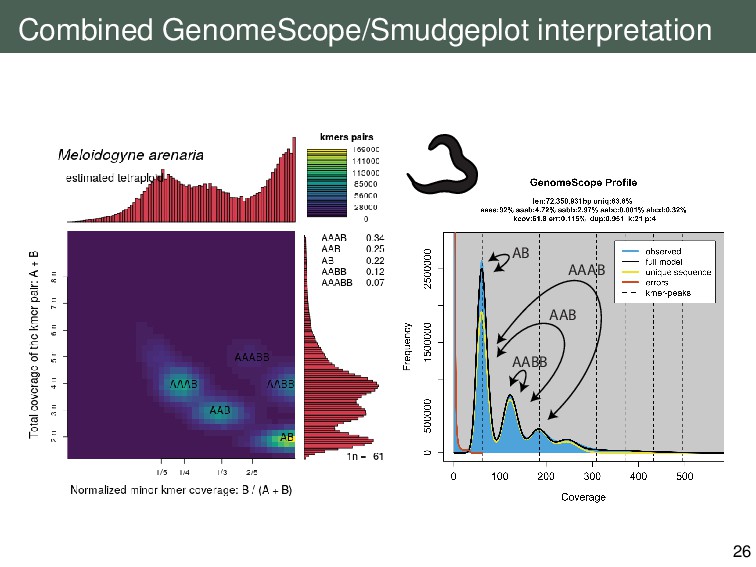
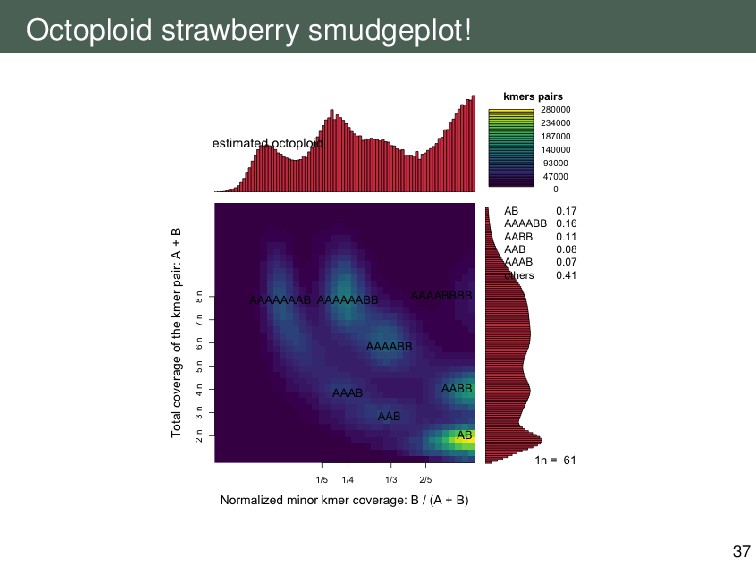
structure A + B B / (A + B) AB
200 0.5 AAB 300 0.33 AABB 400 0.5 AAAB 400 0.25 AAAAB 500 0.2 ... ... ... 1n coverage = 100 23 -
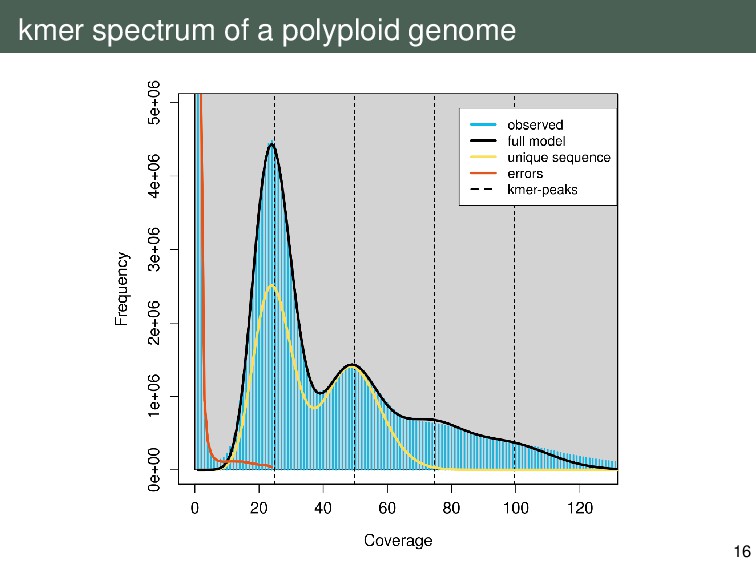
GenomeScope assumptions: uniform distribution of heterozygous loci and duplications the
coverage estimate is right heterozygosity is ”reasonable” (< 12%) GenomeScope strengths: negligible computational time, scales to any dataset Works for any coverage the allows a fit of the model Unbelivebly accurate genome size estimate* *There are conditions to it, check the resources in the supplement 40 -
GenomeScope assumptions: uniform distribution of heterozygous loci and duplications the
coverage estimate is right heterozygosity is ”reasonable” (< 12%) GenomeScope strengths: negligible computational time, scales to any dataset Works for any coverage the allows a fit of the model Unbelivebly accurate genome size estimate* *There are conditions to it, check the resources in the supplement 40 -
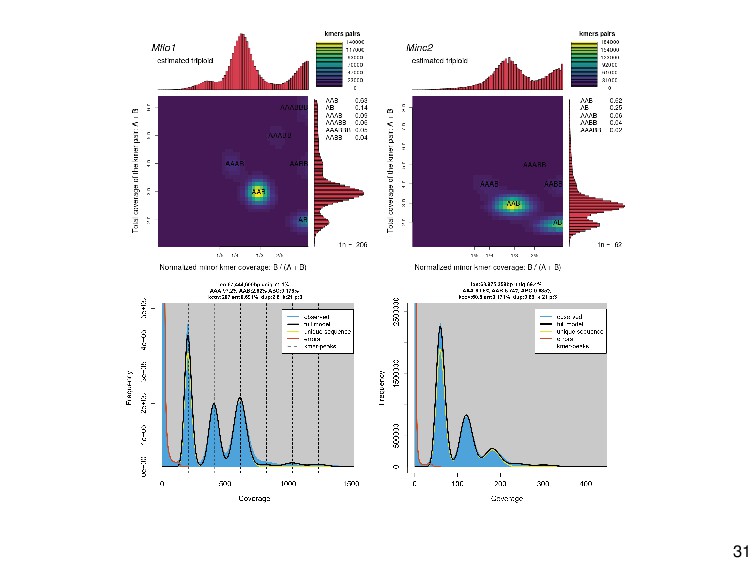
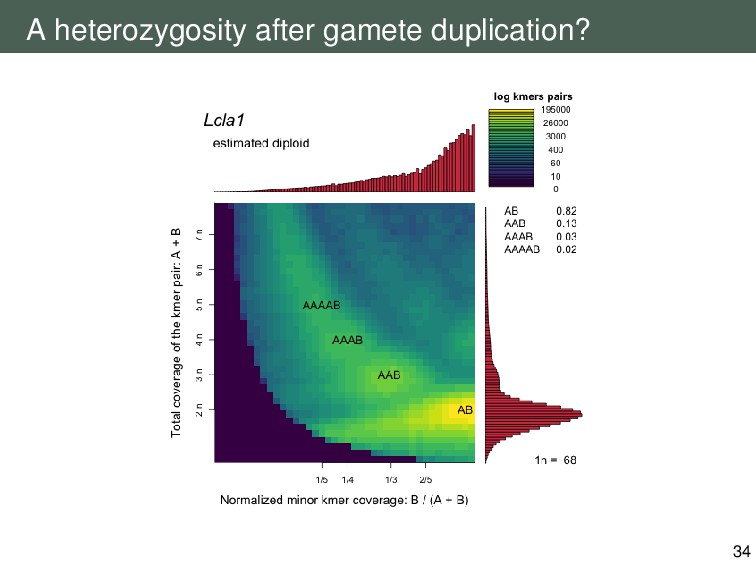
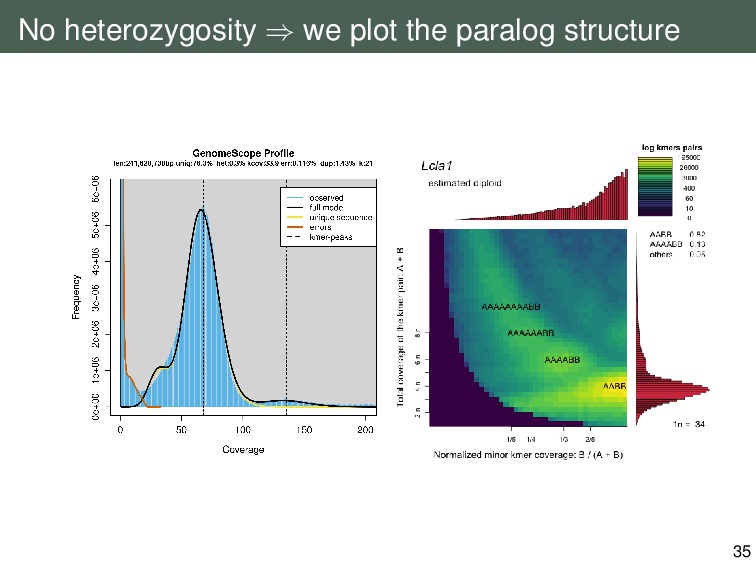
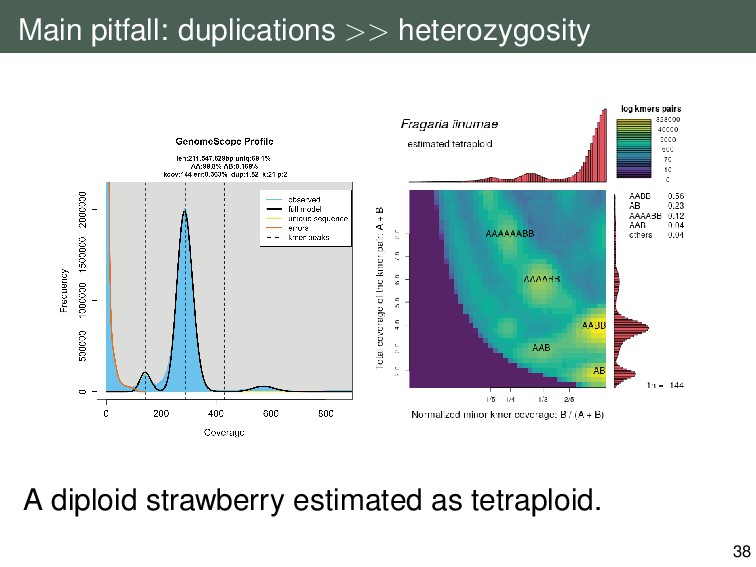
Smudgeplot assumptions: heterozygosity >> pralogy only single genome with decent
coverage computationally it’s still rather exhaustive (Biggest so far 3.5Gbp triploid; sorry plant people :-/) Smudgeplot strengths: Strong descriptive visualization technique especially paired up with GenomeScope 2.0 Decent 1n estimate 41 -
Smudgeplot assumptions: heterozygosity >> pralogy only single genome with decent
coverage computationally it’s still rather exhaustive (Biggest so far 3.5Gbp triploid; sorry plant people :-/) Smudgeplot strengths: Strong descriptive visualization technique especially paired up with GenomeScope 2.0 Decent 1n estimate 41 -
Genome assembly is still useful but it’s good to have
a big scale picture before anything as complicated as genome assembly. Do you have a weird genome to share? https://kamilsjaron.github.io/ peculiar-genomic-observations/ 42 -
Genome assembly is still useful but it’s good to have
a big scale picture before anything as complicated as genome assembly. Do you have a weird genome to share? https://kamilsjaron.github.io/ peculiar-genomic-observations/ 42 -
Resources Kmer methods GenomeScope 2.0 https://github.com/tbenavi1/genomescope2.0 Smudgeplot https://github.com/KamilSJaron/smudgeplot KMC really
fast kmer counter https://github.com/refresh-bio/KMC KAT versatile kmer analysis tools; especially useful to make sense of kmer spectra and genome assembly https://github.com/TGAC/KAT kmer GWAS papers: https://doi.org/10.1038/s41588-020-0612-7, https://doi.org/10.1101/2020.04.14.040675 Other kmer resources Zen and the art of k-mers Good kmer introduction https://dib-lab.github.io/zen-khmer/index.html Peculiar genomics cases A colleciton of strange observations in genomics https://kamilsjaron.github.io/peculiar-genomic-observations/ Genome size estimate considerations: Benchmarking of flow cytometry and kmer based methods using Rooibos genome by Mgwatyu et al. 2020 https://doi.org/10.3390/plants9020270; A peculiar case of marbled crayfish genome size: https://kamilsjaron.github.io/peculiar-genomic-observations/biological/2020/01/crayfish.html More details about the strawberry case: https://github.com/KamilSJaron/smudgeplot/wiki/strawberry-tutorial 44 -
References Vurture, Gregory W., et al. GenomeScope: fast reference-free genome
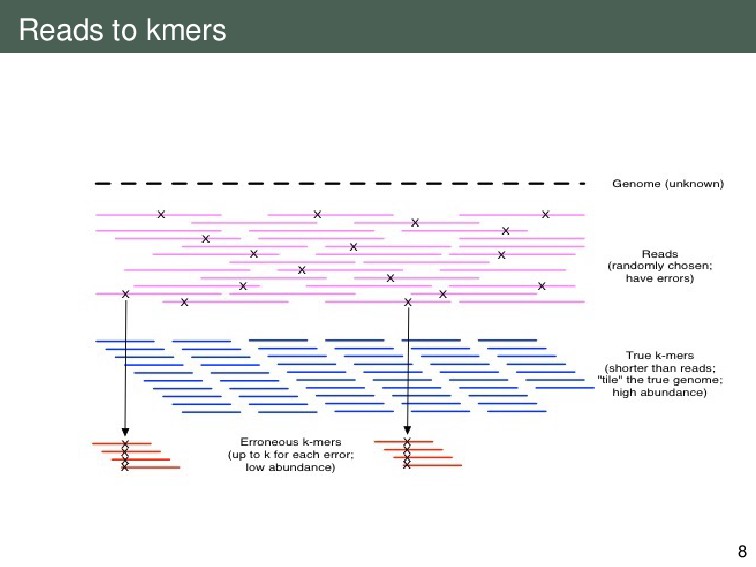
profiling from short reads. Bioinformatics 33.14 (2017): 2202-2204. Ranallo-Benavidez, T.R., Jaron, K.S. & Schatz, M.C. GenomeScope 2.0 and Smudgeplot for reference-free profiling of polyploid genomes. Nat Commun 11, 1432 (2020). https://doi.org/10.1038/s41467-020-14998-3 karyotype from Schwander, T., & Crespi, B. J. (2009). Multiple direct transitions from sexual reproduction to apomictic parthenogenesis in Timema stick insects. Evolution: International Journal of Organic Evolution, 63(1), 84-103. flow cytometry picture from Gutekunst, J., Andriantsoa, R., Falckenhayn, C., Hanna, K., Stein, W., Rasamy, J., & Lyko, F. (2018). Clonal genome evolution and rapid invasive spread of the marbled crayfish. Nature ecology & evolution, 2(3), 567. sequencing errors in kmers pic from https://dib-lab.github.io/zen-khmer/manipulating-histograms.html root knot phylogeny from Lunt, D. H., Kumar, S., Koutsovoulos, G., & Blaxter, M. L. (2014). The complex hybrid origins of the root knot nematodes revealed through comparative genomics. PeerJ, 2, e356. 45